return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 ((Z)-hex-3-ene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-235.083227
Energy at 298.15K-235.095646
HF Energy-234.194863
Nuclear repulsion energy232.238794
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3236 3023 61.46      
2 A 3230 3019 1.49      
3 A 3224 3013 43.87      
4 A 3198 2988 3.20      
5 A 3128 2923 0.14      
6 A 3128 2923 6.99      
7 A 1751 1636 0.46      
8 A 1570 1467 6.92      
9 A 1561 1458 2.69      
10 A 1553 1451 0.11      
11 A 1461 1366 0.11      
12 A 1389 1298 0.28      
13 A 1339 1251 0.02      
14 A 1309 1223 0.19      
15 A 1124 1051 0.74      
16 A 1103 1031 0.53      
17 A 1052 983 0.32      
18 A 1004 939 0.69      
19 A 874 817 1.08      
20 A 796 744 0.49      
21 A 528 493 0.30      
22 A 328 306 0.02      
23 A 221 206 0.00      
24 A 196 183 0.02      
25 A 41 38 0.01      
26 B 3231 3019 63.08      
27 B 3224 3012 1.79      
28 B 3211 3000 7.00      
29 B 3185 2976 9.02      
30 B 3128 2923 58.19      
31 B 3125 2920 37.41      
32 B 1566 1464 1.87      
33 B 1560 1458 6.76      
34 B 1536 1435 0.77      
35 B 1478 1381 2.90      
36 B 1460 1365 2.05      
37 B 1371 1281 10.84      
38 B 1328 1240 0.08      
39 B 1211 1132 0.22      
40 B 1131 1056 6.11      
41 B 1081 1010 2.01      
42 B 941 880 5.01      
43 B 831 777 3.28      
44 B 775 725 29.57      
45 B 568 531 4.76      
46 B 367 343 0.75      
47 B 252 235 0.03      
48 B 41 38 0.06      

Unscaled Zero Point Vibrational Energy (zpe) 37472.8 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 35014.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.27086 0.05247 0.04993

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.008 0.671 0.880
C2 -0.008 -0.671 0.880
C3 -0.008 1.581 -0.310
C4 0.008 -1.581 -0.310
C5 -1.324 2.351 -0.416
C6 1.324 -2.351 -0.416
H7 0.002 1.173 1.843
H8 -0.002 -1.173 1.843
H9 0.164 1.022 -1.228
H10 -0.164 -1.022 -1.228
H11 0.816 2.293 -0.223
H12 -0.816 -2.293 -0.223
H13 -1.312 3.041 -1.257
H14 -2.157 1.663 -0.549
H15 -1.510 2.926 0.489
H16 1.312 -3.041 -1.257
H17 2.157 -1.663 -0.549
H18 1.510 -2.926 0.489

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.34201.49842.54722.50543.54171.08572.07982.14322.70962.12143.26803.45422.77722.74644.47793.47943.9177
C21.34202.54721.49843.54172.50542.07981.08572.70962.14323.26802.12144.47793.47943.91773.45422.77722.7464
C31.49842.54723.16221.52854.15272.19153.49541.08872.76481.09233.95822.17512.16372.16884.89973.90724.8228
C42.54721.49843.16224.15271.52853.49542.19152.76481.08873.95821.09234.89973.90724.82282.17512.16372.1688
C52.50543.54171.52854.15275.39622.87234.38922.15433.65812.14984.67521.08871.08811.08886.06085.31466.0580
C63.54172.50544.15271.52855.39624.38922.87233.65812.15434.67522.14986.06085.31466.05801.08871.08811.0888
H71.08572.07982.19153.49542.87234.38922.34513.07933.77852.48724.11663.85133.25912.68205.39324.29004.5724
H82.07981.08573.49542.19154.38922.87232.34513.77853.07934.11662.48725.39324.29004.57243.85133.25912.6820
H92.14322.70961.08872.76482.15433.65813.07933.77852.07051.74643.60002.50162.50203.06254.22253.41194.5112
H102.70962.14322.76481.08873.65812.15433.77853.07932.07053.60001.74644.22253.41194.51122.50162.50203.0625
H112.12143.26801.09233.95822.14984.67522.48724.11661.74643.60004.86732.48193.05632.51345.45614.18945.3131
H123.26802.12143.95821.09234.67522.14984.11662.48723.60001.74644.86735.45614.18945.31312.48193.05632.5134
H133.45424.47792.17514.89971.08876.06083.85135.39322.50164.22252.48195.45611.76511.76186.62465.88786.8285
H142.77723.47942.16373.90721.08815.31463.25914.29002.50203.41193.05634.18941.76511.75895.88785.44695.9650
H152.74643.91772.16884.82281.08886.05802.68204.57243.06254.51122.51345.31311.76181.75896.82855.96506.5856
H164.47793.45424.89972.17516.06081.08875.39323.85134.22252.50165.45612.48196.62465.88786.82851.76511.7618
H173.47942.77723.90722.16375.31461.08814.29003.25913.41192.50204.18943.05635.88785.44695.96501.76511.7589
H183.91772.74644.82282.16886.05801.08884.57242.68204.51123.06255.31312.51346.82855.96506.58561.76181.7589

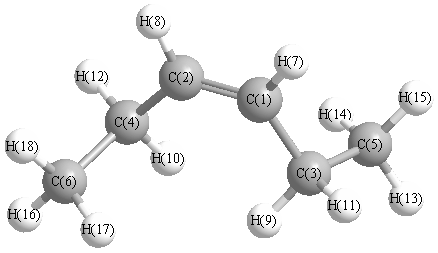
picture of (Z)-hex-3-ene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 127.388 C1 C2 H8 117.510
C1 C3 C5 111.721 C1 C3 H9 110.883
C1 C3 H11 108.930 C2 C1 C3 127.388
C2 C1 H7 117.510 C2 C4 C6 111.721
C2 C4 H10 110.883 C2 C4 H12 108.930
C3 C1 H7 115.075 C3 C5 H13 111.310
C3 C5 H14 110.430 C3 C5 H15 110.796
C4 C2 H8 115.075 C4 C6 H16 111.310
C4 C6 H17 110.430 C4 C6 H18 110.796
C5 C3 H9 109.655 C5 C3 H11 109.092
C6 C4 H10 109.655 C6 C4 H12 109.092
H9 C3 H11 106.399 H10 C4 H12 106.399
H13 C5 H14 108.371 H13 C5 H15 108.014
H14 C5 H15 107.800 H16 C6 H17 108.371
H16 C6 H18 108.014 H17 C6 H18 107.800
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability