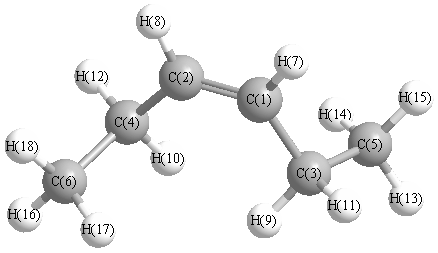
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3174 |
3025 |
55.58 |
|
|
|
| 2 |
A |
3156 |
3008 |
0.42 |
|
|
|
| 3 |
A |
3149 |
3001 |
65.97 |
|
|
|
| 4 |
A |
3122 |
2975 |
3.48 |
|
|
|
| 5 |
A |
3070 |
2926 |
0.22 |
|
|
|
| 6 |
A |
3054 |
2911 |
6.02 |
|
|
|
| 7 |
A |
1771 |
1688 |
0.86 |
|
|
|
| 8 |
A |
1536 |
1464 |
10.29 |
|
|
|
| 9 |
A |
1525 |
1454 |
3.87 |
|
|
|
| 10 |
A |
1515 |
1444 |
0.45 |
|
|
|
| 11 |
A |
1434 |
1367 |
0.15 |
|
|
|
| 12 |
A |
1365 |
1301 |
0.36 |
|
|
|
| 13 |
A |
1322 |
1260 |
0.04 |
|
|
|
| 14 |
A |
1289 |
1229 |
0.21 |
|
|
|
| 15 |
A |
1105 |
1053 |
0.81 |
|
|
|
| 16 |
A |
1086 |
1035 |
1.64 |
|
|
|
| 17 |
A |
1044 |
995 |
0.09 |
|
|
|
| 18 |
A |
1017 |
969 |
1.15 |
|
|
|
| 19 |
A |
865 |
824 |
1.94 |
|
|
|
| 20 |
A |
788 |
751 |
0.83 |
|
|
|
| 21 |
A |
519 |
494 |
0.40 |
|
|
|
| 22 |
A |
311 |
296 |
0.00 |
|
|
|
| 23 |
A |
209 |
199 |
0.00 |
|
|
|
| 24 |
A |
197 |
188 |
0.04 |
|
|
|
| 25 |
A |
61 |
58 |
0.01 |
|
|
|
| 26 |
B |
3156 |
3008 |
73.35 |
|
|
|
| 27 |
B |
3151 |
3003 |
2.33 |
|
|
|
| 28 |
B |
3146 |
2998 |
1.97 |
|
|
|
| 29 |
B |
3111 |
2965 |
7.32 |
|
|
|
| 30 |
B |
3069 |
2925 |
67.67 |
|
|
|
| 31 |
B |
3052 |
2909 |
45.44 |
|
|
|
| 32 |
B |
1534 |
1462 |
3.12 |
|
|
|
| 33 |
B |
1525 |
1454 |
9.30 |
|
|
|
| 34 |
B |
1505 |
1434 |
1.23 |
|
|
|
| 35 |
B |
1459 |
1391 |
4.44 |
|
|
|
| 36 |
B |
1433 |
1366 |
3.64 |
|
|
|
| 37 |
B |
1343 |
1280 |
11.45 |
|
|
|
| 38 |
B |
1300 |
1239 |
0.47 |
|
|
|
| 39 |
B |
1193 |
1137 |
0.20 |
|
|
|
| 40 |
B |
1110 |
1058 |
6.28 |
|
|
|
| 41 |
B |
1063 |
1013 |
3.66 |
|
|
|
| 42 |
B |
928 |
885 |
8.34 |
|
|
|
| 43 |
B |
820 |
782 |
5.42 |
|
|
|
| 44 |
B |
764 |
729 |
30.98 |
|
|
|
| 45 |
B |
569 |
543 |
6.97 |
|
|
|
| 46 |
B |
360 |
343 |
0.97 |
|
|
|
| 47 |
B |
243 |
231 |
0.04 |
|
|
|
| 48 |
B |
53 |
50 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36784.4 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 35059.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.131 |
|
|
|
| 2 |
C |
-0.131 |
|
|
|
| 3 |
C |
-0.369 |
|
|
|
| 4 |
C |
-0.369 |
|
|
|
| 5 |
C |
-0.518 |
|
|
|
| 6 |
C |
-0.518 |
|
|
|
| 7 |
H |
0.151 |
|
|
|
| 8 |
H |
0.151 |
|
|
|
| 9 |
H |
0.171 |
|
|
|
| 10 |
H |
0.171 |
|
|
|
| 11 |
H |
0.175 |
|
|
|
| 12 |
H |
0.175 |
|
|
|
| 13 |
H |
0.171 |
|
|
|
| 14 |
H |
0.178 |
|
|
|
| 15 |
H |
0.171 |
|
|
|
| 16 |
H |
0.171 |
|
|
|
| 17 |
H |
0.178 |
|
|
|
| 18 |
H |
0.171 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.197 |
0.197 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.976 |
0.456 |
0.000 |
| y |
0.456 |
-37.899 |
0.000 |
| z |
0.000 |
0.000 |
-38.510 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.772 |
0.456 |
0.000 |
| y |
0.456 |
1.844 |
0.000 |
| z |
0.000 |
0.000 |
0.928 |
|
| Polar |
|---|
| 3z2-r2 | 1.856 |
|---|
| x2-y2 | -3.077 |
|---|
| xy | 0.456 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.518 |
-0.877 |
0.000 |
| y |
-0.877 |
12.736 |
0.000 |
| z |
0.000 |
0.000 |
8.663 |
<r2> (average value of r
2) Å
2
| <r2> |
256.962 |
| (<r2>)1/2 |
16.030 |