return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7N (Cyclopropylamine)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-172.713756
Energy at 298.15K-172.722305
HF Energy-172.097639
Nuclear repulsion energy125.660548
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3573 3339 0.60      
2 A 3350 3131 12.37      
3 A 3245 3032 4.58      
4 A 3178 2970 44.33      
5 A 1698 1587 14.91      
6 A 1572 1469 13.58      
7 A 1467 1370 20.22      
8 A 1289 1205 0.26      
9 A 1217 1137 6.55      
10 A 1109 1036 15.22      
11 A 1057 988 6.32      
12 A 890 832 117.21      
13 A 852 796 36.22      
14 A 802 749 2.22      
15 A 419 391 6.56      
16 A 3675 3434 0.70      
17 A 3340 3121 0.32      
18 A 3239 3027 13.02      
19 A 1514 1415 0.95      
20 A 1304 1218 0.91      
21 A 1241 1159 0.09      
22 A 1180 1103 0.53      
23 A 1112 1039 1.27      
24 A 945 883 9.76      
25 A 875 817 2.07      
26 A 413 386 9.61      
27 A 291 272 37.54      

Unscaled Zero Point Vibrational Energy (zpe) 22423.2 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 20952.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.54937 0.22625 0.19538

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.530 0.000
H2 -0.410 1.537 0.000
N3 1.430 0.362 0.000
C4 -0.789 -0.498 0.751
C5 -0.789 -0.498 -0.751
H6 1.831 0.814 0.814
H7 1.831 0.814 -0.814
H8 -1.684 -0.185 1.266
H9 -1.684 -0.185 -1.266
H10 -0.216 -1.269 1.242
H11 -0.216 -1.269 -1.242

Atom - Atom Distances (Å)
  C1 H2 N3 C4 C5 H6 H7 H8 H9 H10 H11
C11.08701.44021.49811.49812.02392.02392.22512.22512.19702.1970
H21.08702.18342.20222.20222.49182.49182.48822.48823.07483.0748
N31.44022.18342.49582.49581.01341.01343.40623.40622.62932.6293
C41.49812.20222.49581.50282.93073.32191.07922.22951.07872.2129
C51.49812.20222.49581.50283.32192.93072.22951.07922.21291.0787
H62.02392.49181.01342.93073.32191.62783.68234.20492.95143.5714
H72.02392.49181.01343.32192.93071.62784.20493.68233.57142.9514
H82.22512.48823.40621.07922.22953.68234.20492.53251.82533.1021
H92.22512.48823.40622.22951.07924.20493.68232.53253.10211.8253
H102.19703.07482.62931.07872.21292.95143.57141.82533.10212.4843
H112.19703.07482.62932.21291.07873.57142.95143.10211.82532.4843

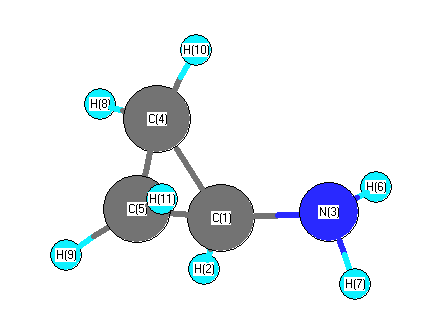
picture of Cyclopropylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 H6 109.928 C1 N3 H7 109.928
C1 C4 C5 59.897 C1 C4 H8 118.479
C1 C4 H10 116.045 C1 C5 C4 59.897
C1 C5 H9 118.479 C1 C5 H11 116.045
H2 C1 N3 118.868 H2 C1 C4 115.921
H2 C1 C5 115.921 N3 C1 C4 116.280
N3 C1 C5 116.280 C4 C1 C5 60.207
C4 C5 H9 118.494 C4 C5 H11 117.062
C5 C4 H8 118.494 C5 C4 H10 117.062
H6 N3 H7 106.857 H8 C4 H10 115.537
H9 C5 H11 115.537
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability