return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8O (Methyl cyclopropyl ketone)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-269.746274
Energy at 298.15K-269.755037
HF Energy-268.846090
Nuclear repulsion energy231.073261
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3352 3132 3.63      
2 A' 3270 3055 8.25      
3 A' 3266 3052 6.08      
4 A' 3242 3029 6.49      
5 A' 3130 2925 2.55      
6 A' 1784 1667 70.58      
7 A' 1556 1454 11.06      
8 A' 1527 1427 10.60      
9 A' 1468 1372 104.88      
10 A' 1427 1334 6.93      
11 A' 1267 1184 4.18      
12 A' 1234 1153 83.42      
13 A' 1142 1067 10.12      
14 A' 1111 1038 11.56      
15 A' 1008 942 43.52      
16 A' 960 897 9.33      
17 A' 834 779 1.75      
18 A' 779 728 0.33      
19 A' 599 560 11.85      
20 A' 379 354 3.83      
21 A' 248 232 4.15      
22 A" 3341 3122 0.49      
23 A" 3239 3026 9.06      
24 A" 3219 3008 6.82      
25 A" 1537 1436 8.67      
26 A" 1511 1412 2.32      
27 A" 1252 1170 0.14      
28 A" 1169 1092 2.30      
29 A" 1130 1056 1.84      
30 A" 1079 1008 3.06      
31 A" 921 861 0.05      
32 A" 869 812 4.79      
33 A" 612 572 1.67      
34 A" 269 252 0.40      
35 A" 146 136 0.04      
36 A" 76 71 1.75      

Unscaled Zero Point Vibrational Energy (zpe) 26977.1 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 25207.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.24043 0.08872 0.07733

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 -1.482 -0.538 0.000
C2 -0.272 -0.753 0.000
C3 0.296 -2.150 0.000
C4 0.707 0.366 0.000
C5 0.296 1.619 0.742
C6 0.296 1.619 -0.742
H7 -0.511 -2.875 0.000
H8 0.925 -2.294 -0.878
H9 0.925 -2.294 0.878
H10 1.755 0.105 0.000
H11 -0.662 1.568 1.236
H12 1.071 2.161 1.262
H13 -0.662 1.568 -1.236
H14 1.071 2.161 -1.262

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
O11.22962.40062.36812.89202.89202.53153.10653.10653.30072.57573.92342.57573.9234
C21.22961.50791.48642.54932.54932.13552.13942.13942.20112.65853.44782.65853.4478
C32.40061.50792.54913.84113.84111.08491.08941.08942.68574.03344.55834.03344.5583
C42.36811.48642.54911.51301.51303.46222.80942.80941.08062.20142.22442.20142.2244
C52.89202.54933.84111.51301.48414.62574.28123.96522.22981.07931.07902.19862.2160
C62.89202.54933.84111.51301.48414.62573.96524.28122.22982.19862.21601.07931.0790
H72.53152.13551.08493.46224.62574.62571.78041.78043.74364.61435.42744.61435.4274
H83.10652.13941.08942.80944.28123.96521.78041.75592.68594.67994.94444.19044.4738
H93.10652.13941.08942.80943.96524.28121.78041.75592.68594.19044.47384.67994.9444
H103.30072.20112.68571.08062.22982.22983.74362.68592.68593.08422.50803.08422.5080
H112.57572.65854.03342.20141.07932.19864.61434.67994.19043.08421.83152.47243.0976
H123.92343.44784.55832.22441.07902.21605.42744.94444.47382.50801.83153.09762.5243
H132.57572.65854.03342.20142.19861.07934.61434.19044.67993.08422.47243.09761.8315
H143.92343.44784.55832.22442.21601.07905.42744.47384.94442.50803.09762.52431.8315

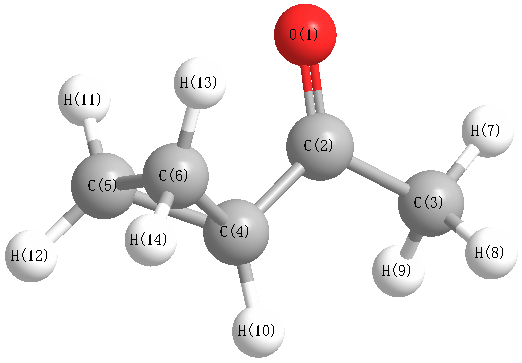
picture of Methyl cyclopropyl ketone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 122.219 O1 C2 C4 121.074
C2 C3 H7 109.824 C2 C3 H8 109.860
C2 C3 H9 109.860 C2 C4 C5 116.409
C2 C4 C6 116.409 C2 C4 H10 117.186
C3 C2 C4 116.707 C4 C5 C6 60.630
C4 C5 H11 115.239 C4 C5 H12 117.246
C4 C6 C5 60.630 C4 C6 H13 115.239
C4 C6 H14 117.246 C5 C4 C6 58.741
C5 C4 H10 117.608 C5 C6 H13 117.249
C5 C6 H14 118.818 C6 C4 H10 117.608
C6 C5 H11 117.249 C6 C5 H12 118.818
H7 C3 H8 109.936 H7 C3 H9 109.936
H8 C3 H9 107.390 H11 C5 H12 116.115
H13 C6 H14 116.115
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability