Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Geometric Data calculated at MP2=FULL/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -235.083384 |
| Energy at 298.15K | -235.095679 |
| HF Energy | -234.195411 |
| Nuclear repulsion energy | 231.232136 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3249 |
3036 |
49.43 |
|
|
|
| 2 |
A |
3238 |
3025 |
9.45 |
|
|
|
| 3 |
A |
3224 |
3012 |
30.72 |
|
|
|
| 4 |
A |
3221 |
3010 |
48.17 |
|
|
|
| 5 |
A |
3217 |
3006 |
2.99 |
|
|
|
| 6 |
A |
3200 |
2990 |
19.04 |
|
|
|
| 7 |
A |
3189 |
2980 |
23.29 |
|
|
|
| 8 |
A |
3171 |
2963 |
1.74 |
|
|
|
| 9 |
A |
3127 |
2922 |
48.19 |
|
|
|
| 10 |
A |
3125 |
2920 |
15.78 |
|
|
|
| 11 |
A |
3125 |
2920 |
17.15 |
|
|
|
| 12 |
A |
3112 |
2908 |
14.92 |
|
|
|
| 13 |
A |
1757 |
1641 |
1.23 |
|
|
|
| 14 |
A |
1569 |
1466 |
3.92 |
|
|
|
| 15 |
A |
1562 |
1459 |
5.96 |
|
|
|
| 16 |
A |
1559 |
1457 |
4.28 |
|
|
|
| 17 |
A |
1551 |
1449 |
0.06 |
|
|
|
| 18 |
A |
1544 |
1443 |
4.00 |
|
|
|
| 19 |
A |
1539 |
1438 |
0.45 |
|
|
|
| 20 |
A |
1482 |
1385 |
6.09 |
|
|
|
| 21 |
A |
1466 |
1370 |
1.60 |
|
|
|
| 22 |
A |
1452 |
1356 |
4.42 |
|
|
|
| 23 |
A |
1423 |
1330 |
0.36 |
|
|
|
| 24 |
A |
1365 |
1275 |
0.04 |
|
|
|
| 25 |
A |
1330 |
1243 |
4.10 |
|
|
|
| 26 |
A |
1329 |
1242 |
0.43 |
|
|
|
| 27 |
A |
1295 |
1210 |
0.11 |
|
|
|
| 28 |
A |
1203 |
1124 |
0.61 |
|
|
|
| 29 |
A |
1151 |
1075 |
1.35 |
|
|
|
| 30 |
A |
1100 |
1028 |
1.49 |
|
|
|
| 31 |
A |
1094 |
1022 |
0.56 |
|
|
|
| 32 |
A |
1092 |
1020 |
0.80 |
|
|
|
| 33 |
A |
1038 |
970 |
4.46 |
|
|
|
| 34 |
A |
1006 |
940 |
0.21 |
|
|
|
| 35 |
A |
947 |
885 |
2.37 |
|
|
|
| 36 |
A |
928 |
867 |
4.75 |
|
|
|
| 37 |
A |
895 |
837 |
2.83 |
|
|
|
| 38 |
A |
763 |
713 |
1.51 |
|
|
|
| 39 |
A |
735 |
687 |
28.17 |
|
|
|
| 40 |
A |
574 |
537 |
6.23 |
|
|
|
| 41 |
A |
478 |
447 |
0.71 |
|
|
|
| 42 |
A |
369 |
345 |
0.02 |
|
|
|
| 43 |
A |
299 |
280 |
0.05 |
|
|
|
| 44 |
A |
255 |
238 |
0.00 |
|
|
|
| 45 |
A |
202 |
189 |
0.01 |
|
|
|
| 46 |
A |
113 |
105 |
0.26 |
|
|
|
| 47 |
A |
94 |
88 |
0.00 |
|
|
|
| 48 |
A |
39 |
36 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37396.2 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 34943.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.410 |
0.979 |
-0.040 |
| C2 |
1.982 |
-0.450 |
-0.167 |
| C3 |
0.776 |
-0.967 |
0.111 |
| C4 |
-0.442 |
-0.235 |
0.584 |
| C5 |
-1.535 |
-0.184 |
-0.486 |
| C6 |
-2.802 |
0.504 |
0.003 |
| H7 |
1.604 |
1.627 |
0.290 |
| H8 |
3.231 |
1.075 |
0.671 |
| H9 |
2.772 |
1.356 |
-0.997 |
| H10 |
2.747 |
-1.132 |
-0.521 |
| H11 |
0.640 |
-2.032 |
-0.051 |
| H12 |
-0.196 |
0.782 |
0.891 |
| H13 |
-0.842 |
-0.735 |
1.470 |
| H14 |
-1.143 |
0.333 |
-1.363 |
| H15 |
-1.768 |
-1.202 |
-0.808 |
| H16 |
-3.564 |
0.532 |
-0.774 |
| H17 |
-3.219 |
-0.018 |
0.863 |
| H18 |
-2.594 |
1.529 |
0.305 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4973 | 2.5459 | 3.1620 | 4.1373 | 5.2343 | 1.0859 | 1.0901 | 1.0899 | 2.1918 | 3.4926 | 2.7742 | 3.9746 | 3.8457 | 4.7749 | 6.0360 | 5.7882 | 5.0464 |
C2 | 1.4973 | | 1.3417 | 2.5471 | 3.5418 | 4.8818 | 2.1604 | 2.1416 | 2.1388 | 1.0844 | 2.0777 | 2.7167 | 3.2769 | 3.4366 | 3.8780 | 5.6657 | 5.3203 | 5.0085 | C3 | 2.5459 | 1.3417 | | 1.4977 | 2.5115 | 3.8700 | 2.7291 | 3.2422 | 3.2571 | 2.0767 | 1.0854 | 2.1471 | 2.1256 | 2.7462 | 2.7143 | 4.6762 | 4.1746 | 4.1981 | C4 | 3.1620 | 2.5471 | 1.4977 | | 1.5301 | 2.5404 | 2.7819 | 3.9006 | 3.9192 | 3.4925 | 2.1915 | 1.0899 | 1.0936 | 2.1453 | 2.1513 | 3.4901 | 2.7997 | 2.7964 | C5 | 4.1373 | 3.5418 | 2.5115 | 1.5301 | | 1.5226 | 3.7061 | 5.0635 | 4.6029 | 4.3860 | 2.8866 | 2.1500 | 2.1468 | 1.0912 | 1.0918 | 2.1714 | 2.1641 | 2.1642 | C6 | 5.2343 | 4.8818 | 3.8700 | 2.5404 | 1.5226 | | 4.5561 | 6.0971 | 5.7272 | 5.8093 | 4.2756 | 2.7678 | 2.7443 | 2.1557 | 2.1528 | 1.0883 | 1.0891 | 1.0891 | H7 | 1.0859 | 2.1604 | 2.7291 | 2.7819 | 3.7061 | 4.5561 | | 1.7603 | 1.7588 | 3.0954 | 3.7990 | 2.0772 | 3.5997 | 3.4566 | 4.5358 | 5.3888 | 5.1283 | 4.1992 | H8 | 1.0901 | 2.1416 | 3.2422 | 3.9006 | 5.0635 | 6.0971 | 1.7603 | | 1.7519 | 2.5549 | 4.1091 | 3.4464 | 4.5285 | 4.8801 | 5.6882 | 6.9685 | 6.5454 | 5.8545 | H9 | 1.0899 | 2.1388 | 3.2571 | 3.9192 | 4.6029 | 5.7272 | 1.7588 | 1.7519 | | 2.5338 | 4.1131 | 3.5639 | 4.8499 | 4.0630 | 5.2143 | 6.3938 | 6.4223 | 5.5249 | H10 | 2.1918 | 1.0844 | 2.0767 | 3.4925 | 4.3860 | 5.8093 | 3.0954 | 2.5549 | 2.5338 | | 2.3387 | 3.7839 | 4.1235 | 4.2413 | 4.5245 | 6.5323 | 6.2255 | 6.0247 | H11 | 3.4926 | 2.0777 | 1.0854 | 2.1915 | 2.8866 | 4.2756 | 3.7990 | 4.1091 | 4.1131 | 2.3387 | | 3.0822 | 2.4877 | 3.2392 | 2.6570 | 4.9774 | 4.4479 | 4.8234 | H12 | 2.7742 | 2.7167 | 2.1471 | 1.0899 | 2.1500 | 2.7678 | 2.0772 | 3.4464 | 3.5639 | 3.7839 | 3.0822 | | 1.7478 | 2.4852 | 3.0478 | 3.7659 | 3.1279 | 2.5796 | H13 | 3.9746 | 3.2769 | 2.1256 | 1.0936 | 2.1468 | 2.7443 | 3.5997 | 4.5285 | 4.8499 | 4.1235 | 2.4877 | 1.7478 | | 3.0424 | 2.5022 | 3.7488 | 2.5564 | 3.0911 | H14 | 3.8457 | 3.4366 | 2.7462 | 2.1453 | 1.0912 | 2.1557 | 3.4566 | 4.8801 | 4.0630 | 4.2413 | 3.2392 | 2.4852 | 3.0424 | | 1.7477 | 2.5001 | 3.0640 | 2.5137 | H15 | 4.7749 | 3.8780 | 2.7143 | 2.1513 | 1.0918 | 2.1528 | 4.5358 | 5.6882 | 5.2143 | 4.5245 | 2.6570 | 3.0478 | 2.5022 | 1.7477 | | 2.4970 | 2.5094 | 3.0622 | H16 | 6.0360 | 5.6657 | 4.6762 | 3.4901 | 2.1714 | 1.0883 | 5.3888 | 6.9685 | 6.3938 | 6.5323 | 4.9774 | 3.7659 | 3.7488 | 2.5001 | 2.4970 | | 1.7608 | 1.7605 | H17 | 5.7882 | 5.3203 | 4.1746 | 2.7997 | 2.1641 | 1.0891 | 5.1283 | 6.5454 | 6.4223 | 6.2255 | 4.4479 | 3.1279 | 2.5564 | 3.0640 | 2.5094 | 1.7608 | | 1.7594 | H18 | 5.0464 | 5.0085 | 4.1981 | 2.7964 | 2.1642 | 1.0891 | 4.1992 | 5.8545 | 5.5249 | 6.0247 | 4.8234 | 2.5796 | 3.0911 | 2.5137 | 3.0622 | 1.7605 | 1.7594 | |
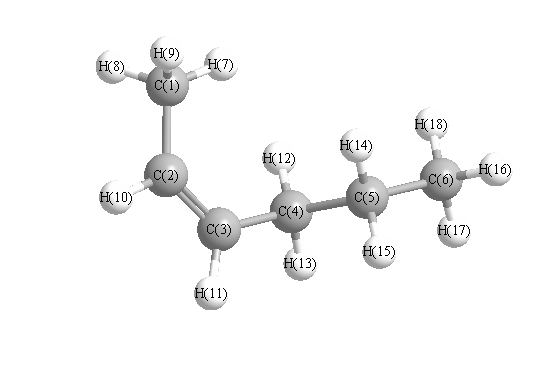 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.384 |
|
C1 |
C2 |
H10 |
115.271 |
| C2 |
C1 |
H7 |
112.538 |
|
C2 |
C1 |
H8 |
110.745 |
| C2 |
C1 |
H9 |
110.533 |
|
C2 |
C3 |
C4 |
127.465 |
| C2 |
C3 |
H11 |
117.358 |
|
C3 |
C2 |
H10 |
117.345 |
| C3 |
C4 |
C5 |
112.083 |
|
C3 |
C4 |
H12 |
111.171 |
| C3 |
C4 |
H13 |
109.236 |
|
C4 |
C3 |
H11 |
115.148 |
| C4 |
C5 |
C6 |
112.651 |
|
C4 |
C5 |
H14 |
108.696 |
| C4 |
C5 |
H15 |
109.127 |
|
C5 |
C4 |
H12 |
109.135 |
| C5 |
C4 |
H13 |
108.676 |
|
C5 |
C6 |
H16 |
111.459 |
| C5 |
C6 |
H17 |
110.826 |
|
C5 |
C6 |
H18 |
110.831 |
| C6 |
C5 |
H14 |
110.031 |
|
C6 |
C5 |
H15 |
109.761 |
| H7 |
C1 |
H8 |
107.989 |
|
H7 |
C1 |
H9 |
107.864 |
| H8 |
C1 |
H9 |
106.955 |
|
H12 |
C4 |
H13 |
106.348 |
| H14 |
C5 |
H16 |
94.170 |
|
H16 |
C6 |
H17 |
107.930 |
| H16 |
C6 |
H18 |
107.896 |
|
H17 |
C6 |
H18 |
107.750 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
