Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -235.584864 |
| Energy at 298.15K | -235.597146 |
| HF Energy | -235.584864 |
| Nuclear repulsion energy | 230.265018 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3180 |
3035 |
54.39 |
|
|
|
| 2 |
A |
3162 |
3018 |
9.56 |
|
|
|
| 3 |
A |
3150 |
3006 |
11.72 |
|
|
|
| 4 |
A |
3145 |
3001 |
32.73 |
|
|
|
| 5 |
A |
3138 |
2995 |
49.61 |
|
|
|
| 6 |
A |
3111 |
2969 |
21.28 |
|
|
|
| 7 |
A |
3110 |
2968 |
18.81 |
|
|
|
| 8 |
A |
3091 |
2950 |
5.54 |
|
|
|
| 9 |
A |
3058 |
2918 |
42.08 |
|
|
|
| 10 |
A |
3056 |
2916 |
18.98 |
|
|
|
| 11 |
A |
3053 |
2914 |
28.06 |
|
|
|
| 12 |
A |
3034 |
2895 |
21.33 |
|
|
|
| 13 |
A |
1769 |
1688 |
1.97 |
|
|
|
| 14 |
A |
1518 |
1449 |
5.44 |
|
|
|
| 15 |
A |
1509 |
1440 |
7.54 |
|
|
|
| 16 |
A |
1504 |
1435 |
5.07 |
|
|
|
| 17 |
A |
1501 |
1432 |
0.85 |
|
|
|
| 18 |
A |
1491 |
1423 |
5.57 |
|
|
|
| 19 |
A |
1488 |
1420 |
0.25 |
|
|
|
| 20 |
A |
1448 |
1381 |
10.13 |
|
|
|
| 21 |
A |
1424 |
1359 |
3.19 |
|
|
|
| 22 |
A |
1412 |
1347 |
2.97 |
|
|
|
| 23 |
A |
1386 |
1323 |
0.30 |
|
|
|
| 24 |
A |
1330 |
1269 |
0.04 |
|
|
|
| 25 |
A |
1304 |
1244 |
1.48 |
|
|
|
| 26 |
A |
1294 |
1235 |
3.27 |
|
|
|
| 27 |
A |
1259 |
1201 |
0.18 |
|
|
|
| 28 |
A |
1178 |
1125 |
0.94 |
|
|
|
| 29 |
A |
1124 |
1073 |
1.04 |
|
|
|
| 30 |
A |
1078 |
1029 |
2.00 |
|
|
|
| 31 |
A |
1069 |
1020 |
1.43 |
|
|
|
| 32 |
A |
1063 |
1015 |
0.57 |
|
|
|
| 33 |
A |
1020 |
973 |
4.93 |
|
|
|
| 34 |
A |
1014 |
968 |
2.79 |
|
|
|
| 35 |
A |
935 |
892 |
4.01 |
|
|
|
| 36 |
A |
910 |
868 |
7.24 |
|
|
|
| 37 |
A |
877 |
837 |
3.91 |
|
|
|
| 38 |
A |
752 |
717 |
2.31 |
|
|
|
| 39 |
A |
722 |
689 |
29.98 |
|
|
|
| 40 |
A |
573 |
547 |
8.42 |
|
|
|
| 41 |
A |
474 |
452 |
0.93 |
|
|
|
| 42 |
A |
367 |
351 |
0.03 |
|
|
|
| 43 |
A |
289 |
276 |
0.04 |
|
|
|
| 44 |
A |
249 |
238 |
0.01 |
|
|
|
| 45 |
A |
201 |
192 |
0.04 |
|
|
|
| 46 |
A |
139 |
133 |
0.34 |
|
|
|
| 47 |
A |
96 |
91 |
0.01 |
|
|
|
| 48 |
A |
55 |
53 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36554.1 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 34883.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.487 |
0.935 |
-0.083 |
| C2 |
1.995 |
-0.477 |
-0.133 |
| C3 |
0.766 |
-0.923 |
0.141 |
| C4 |
-0.434 |
-0.125 |
0.546 |
| C5 |
-1.594 |
-0.259 |
-0.447 |
| C6 |
-2.837 |
0.504 |
-0.010 |
| H7 |
1.707 |
1.647 |
0.194 |
| H8 |
3.306 |
1.038 |
0.639 |
| H9 |
2.891 |
1.241 |
-1.056 |
| H10 |
2.742 |
-1.215 |
-0.423 |
| H11 |
0.591 |
-1.996 |
0.049 |
| H12 |
-0.181 |
0.935 |
0.666 |
| H13 |
-0.780 |
-0.468 |
1.533 |
| H14 |
-1.262 |
0.095 |
-1.430 |
| H15 |
-1.837 |
-1.322 |
-0.574 |
| H16 |
-3.650 |
0.395 |
-0.733 |
| H17 |
-3.202 |
0.146 |
0.959 |
| H18 |
-2.628 |
1.575 |
0.094 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.4954 | 2.5425 | 3.1708 | 4.2674 | 5.3418 | 1.0922 | 1.0969 | 1.0967 | 2.1917 | 3.4931 | 2.7715 | 3.9056 | 4.0713 | 4.9026 | 6.1955 | 5.8375 | 5.1577 |
C2 | 1.4954 | | 1.3361 | 2.5470 | 3.6093 | 4.9322 | 2.1680 | 2.1464 | 2.1458 | 1.0900 | 2.0768 | 2.7140 | 3.2365 | 3.5525 | 3.9495 | 5.7441 | 5.3471 | 5.0627 | C3 | 2.5425 | 1.3361 | | 1.4975 | 2.5209 | 3.8784 | 2.7378 | 3.2473 | 3.2605 | 2.0758 | 1.0910 | 2.1504 | 2.1292 | 2.7602 | 2.7292 | 4.6912 | 4.1902 | 4.2142 | C4 | 3.1708 | 2.5470 | 1.4975 | | 1.5320 | 2.5449 | 2.8019 | 3.9181 | 3.9355 | 3.4956 | 2.1911 | 1.0959 | 1.1005 | 2.1538 | 2.1577 | 3.4999 | 2.8114 | 2.8112 | C5 | 4.2674 | 3.6093 | 2.5209 | 1.5320 | | 1.5227 | 3.8652 | 5.1835 | 4.7678 | 4.4402 | 2.8349 | 2.1581 | 2.1501 | 1.0970 | 1.0980 | 2.1771 | 2.1738 | 2.1735 | C6 | 5.3418 | 4.9322 | 3.8784 | 2.5449 | 1.5227 | | 4.6899 | 6.2001 | 5.8687 | 5.8527 | 4.2436 | 2.7738 | 2.7489 | 2.1600 | 2.1571 | 1.0942 | 1.0957 | 1.0956 | H7 | 1.0922 | 2.1680 | 2.7378 | 2.8019 | 3.8652 | 4.6899 | | 1.7681 | 1.7688 | 3.1059 | 3.8132 | 2.0727 | 3.5289 | 3.7234 | 4.6872 | 5.5797 | 5.1905 | 4.3365 | H8 | 1.0969 | 2.1464 | 3.2473 | 3.9181 | 5.1835 | 6.2001 | 1.7681 | | 1.7569 | 2.5540 | 4.1138 | 3.4889 | 4.4455 | 5.1028 | 5.7875 | 7.1198 | 6.5769 | 5.9828 | H9 | 1.0967 | 2.1458 | 3.2605 | 3.9355 | 4.7678 | 5.8687 | 1.7688 | 1.7569 | | 2.5409 | 4.1214 | 3.5348 | 4.8059 | 4.3242 | 5.3997 | 6.6037 | 6.5102 | 5.6469 | H10 | 2.1917 | 1.0900 | 2.0758 | 3.4956 | 4.4402 | 5.8527 | 3.1059 | 2.5540 | 2.5409 | | 2.3364 | 3.7888 | 4.0974 | 4.3318 | 4.5833 | 6.5997 | 6.2528 | 6.0735 | H11 | 3.4931 | 2.0768 | 1.0910 | 2.1911 | 2.8349 | 4.2436 | 3.8132 | 4.1138 | 4.1214 | 2.3364 | | 3.0930 | 2.5331 | 3.1613 | 2.5962 | 4.9315 | 4.4503 | 4.8077 | H12 | 2.7715 | 2.7140 | 2.1504 | 1.0959 | 2.1581 | 2.7738 | 2.0727 | 3.4889 | 3.5348 | 3.7888 | 3.0930 | | 1.7545 | 2.5034 | 3.0614 | 3.7795 | 3.1361 | 2.5925 | H13 | 3.9056 | 3.2365 | 2.1292 | 1.1005 | 2.1501 | 2.7489 | 3.5289 | 4.4455 | 4.8059 | 4.0974 | 2.5331 | 1.7545 | | 3.0543 | 2.5069 | 3.7575 | 2.5639 | 3.1076 | H14 | 4.0713 | 3.5525 | 2.7602 | 2.1538 | 1.0970 | 2.1600 | 3.7234 | 5.1028 | 4.3242 | 4.3318 | 3.1613 | 2.5034 | 3.0543 | | 1.7528 | 2.5063 | 3.0782 | 2.5259 | H15 | 4.9026 | 3.9495 | 2.7292 | 2.1577 | 1.0980 | 2.1571 | 4.6872 | 5.7875 | 5.3997 | 4.5833 | 2.5962 | 3.0614 | 2.5069 | 1.7528 | | 2.5020 | 2.5231 | 3.0761 | H16 | 6.1955 | 5.7441 | 4.6912 | 3.4999 | 2.1771 | 1.0942 | 5.5797 | 7.1198 | 6.6037 | 6.5997 | 4.9315 | 3.7795 | 3.7575 | 2.5063 | 2.5020 | | 1.7680 | 1.7674 | H17 | 5.8375 | 5.3471 | 4.1902 | 2.8114 | 2.1738 | 1.0957 | 5.1905 | 6.5769 | 6.5102 | 6.2528 | 4.4503 | 3.1361 | 2.5639 | 3.0782 | 2.5231 | 1.7680 | | 1.7663 | H18 | 5.1577 | 5.0627 | 4.2142 | 2.8112 | 2.1735 | 1.0956 | 4.3365 | 5.9828 | 5.6469 | 6.0735 | 4.8077 | 2.5925 | 3.1076 | 2.5259 | 3.0761 | 1.7674 | 1.7663 | |
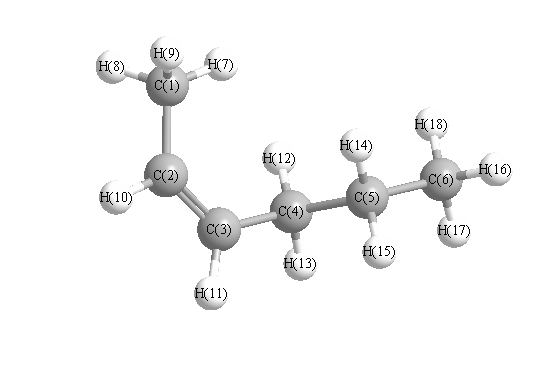 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.684 |
|
C1 |
C2 |
H10 |
115.026 |
| C2 |
C1 |
H7 |
112.902 |
|
C2 |
C1 |
H8 |
110.851 |
| C2 |
C1 |
H9 |
110.816 |
|
C2 |
C3 |
C4 |
127.928 |
| C2 |
C3 |
H11 |
117.309 |
|
C3 |
C2 |
H10 |
117.289 |
| C3 |
C4 |
C5 |
112.634 |
|
C3 |
C4 |
H12 |
111.084 |
| C3 |
C4 |
H13 |
109.128 |
|
C4 |
C3 |
H11 |
114.755 |
| C4 |
C5 |
C6 |
112.838 |
|
C4 |
C5 |
H14 |
108.895 |
| C4 |
C5 |
H15 |
109.142 |
|
C5 |
C4 |
H12 |
109.292 |
| C5 |
C4 |
H13 |
108.414 |
|
C5 |
C6 |
H16 |
111.548 |
| C5 |
C6 |
H17 |
111.191 |
|
C5 |
C6 |
H18 |
111.175 |
| C6 |
C5 |
H14 |
110.020 |
|
C6 |
C5 |
H15 |
109.728 |
| H7 |
C1 |
H8 |
107.739 |
|
H7 |
C1 |
H9 |
107.814 |
| H8 |
C1 |
H9 |
106.439 |
|
H12 |
C4 |
H13 |
106.040 |
| H14 |
C5 |
H16 |
94.059 |
|
H16 |
C6 |
H17 |
107.672 |
| H16 |
C6 |
H18 |
107.631 |
|
H17 |
C6 |
H18 |
107.428 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.426 |
|
|
|
| 2 |
C |
-0.106 |
|
|
|
| 3 |
C |
-0.092 |
|
|
|
| 4 |
C |
-0.266 |
|
|
|
| 5 |
C |
-0.236 |
|
|
|
| 6 |
C |
-0.391 |
|
|
|
| 7 |
H |
0.132 |
|
|
|
| 8 |
H |
0.142 |
|
|
|
| 9 |
H |
0.141 |
|
|
|
| 10 |
H |
0.111 |
|
|
|
| 11 |
H |
0.107 |
|
|
|
| 12 |
H |
0.119 |
|
|
|
| 13 |
H |
0.130 |
|
|
|
| 14 |
H |
0.130 |
|
|
|
| 15 |
H |
0.123 |
|
|
|
| 16 |
H |
0.128 |
|
|
|
| 17 |
H |
0.127 |
|
|
|
| 18 |
H |
0.128 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.033 |
0.190 |
-0.024 |
0.194 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.205 |
0.747 |
-0.596 |
| y |
0.747 |
-38.593 |
0.048 |
| z |
-0.596 |
0.048 |
-40.480 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.331 |
0.747 |
-0.596 |
| y |
0.747 |
0.750 |
0.048 |
| z |
-0.596 |
0.048 |
-2.081 |
|
| Polar |
|---|
| 3z2-r2 | -4.162 |
|---|
| x2-y2 | 0.388 |
|---|
| xy | 0.747 |
|---|
| xz | -0.596 |
|---|
| yz | 0.048 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.817 |
0.671 |
-0.662 |
| y |
0.671 |
9.033 |
0.025 |
| z |
-0.662 |
0.025 |
7.436 |
<r2> (average value of r
2) Å
2
| <r2> |
261.429 |
| (<r2>)1/2 |
16.169 |
