Jump to
S1C2
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -151.410384 |
| Energy at 298.15K | -151.412629 |
| HF Energy | -151.285899 |
| Nuclear repulsion energy | 36.659402 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3808 |
3808 |
7.39 |
91.28 |
0.19 |
0.32 |
| 2 |
A |
1450 |
1450 |
0.31 |
8.73 |
0.64 |
0.78 |
| 3 |
A |
937 |
937 |
1.44 |
11.26 |
0.25 |
0.40 |
| 4 |
A |
337 |
337 |
206.85 |
6.80 |
0.75 |
0.86 |
| 5 |
B |
3809 |
3809 |
39.42 |
34.94 |
0.75 |
0.86 |
| 6 |
B |
1311 |
1311 |
117.34 |
2.30 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 5825.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5825.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.730 |
-0.053 |
| O2 |
0.000 |
-0.730 |
-0.053 |
| H3 |
0.826 |
0.887 |
0.428 |
| H4 |
-0.826 |
-0.887 |
0.428 |
Atom - Atom Distances (Å)
| |
O1 |
O2 |
H3 |
H4 |
| O1 | | 1.4604 | 0.9683 | 1.8785 |
O2 | 1.4604 | | 1.8785 | 0.9683 | H3 | 0.9683 | 1.8785 | | 2.4236 | H4 | 1.8785 | 0.9683 | 2.4236 | |
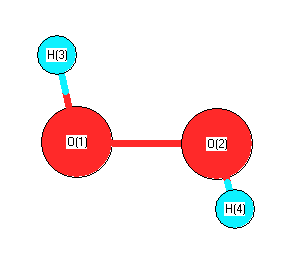 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
O2 |
H4 |
99.321 |
|
O2 |
O1 |
H3 |
99.321 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.345 |
|
|
|
| 2 |
O |
-0.345 |
|
|
|
| 3 |
H |
0.345 |
|
|
|
| 4 |
H |
0.345 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.754 |
1.754 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-9.162 |
3.023 |
0.000 |
| y |
3.023 |
-11.095 |
0.000 |
| z |
0.000 |
0.000 |
-11.614 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.192 |
3.023 |
0.000 |
| y |
3.023 |
-0.707 |
0.000 |
| z |
0.000 |
0.000 |
-1.486 |
|
| Polar |
|---|
| 3z2-r2 | -2.971 |
|---|
| x2-y2 | 1.933 |
|---|
| xy | 3.023 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.404 |
0.334 |
0.000 |
| y |
0.334 |
2.066 |
0.000 |
| z |
0.000 |
0.000 |
0.931 |
<r2> (average value of r
2) Å
2
| <r2> |
18.516 |
| (<r2>)1/2 |
4.303 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G**
| | hartrees |
|---|
| Energy at 0K | -151.409379 |
| Energy at 298.15K | |
| HF Energy | -151.284616 |
| Nuclear repulsion energy | 36.519398 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3832 |
3832 |
0.00 |
|
|
|
| 2 |
Ag |
1540 |
1540 |
0.00 |
|
|
|
| 3 |
Ag |
939 |
939 |
0.00 |
|
|
|
| 4 |
Au |
252i |
252i |
303.37 |
|
|
|
| 5 |
Bu |
3841 |
3841 |
73.49 |
|
|
|
| 6 |
Bu |
1244 |
1244 |
145.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5572.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5572.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G**
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.735 |
0.000 |
| O2 |
0.000 |
-0.735 |
0.000 |
| H3 |
0.958 |
0.868 |
0.000 |
| H4 |
-0.958 |
-0.868 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
O2 |
H3 |
H4 |
| O1 | | 1.4706 | 0.9675 | 1.8678 |
O2 | 1.4706 | | 1.8678 | 0.9675 | H3 | 0.9675 | 1.8678 | | 2.5859 | H4 | 1.8678 | 0.9675 | 2.5859 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
O2 |
H4 |
97.879 |
|
O2 |
O1 |
H3 |
97.879 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.349 |
|
|
|
| 2 |
O |
-0.349 |
|
|
|
| 3 |
H |
0.349 |
|
|
|
| 4 |
H |
0.349 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-7.934 |
3.471 |
0.000 |
| y |
3.471 |
-11.199 |
0.000 |
| z |
0.000 |
0.000 |
-12.590 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.960 |
3.471 |
0.000 |
| y |
3.471 |
-0.937 |
0.000 |
| z |
0.000 |
0.000 |
-3.023 |
|
| Polar |
|---|
| 3z2-r2 | -6.047 |
|---|
| x2-y2 | 3.265 |
|---|
| xy | 3.471 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.618 |
0.354 |
0.000 |
| y |
0.354 |
2.074 |
0.000 |
| z |
0.000 |
0.000 |
0.705 |
<r2> (average value of r
2) Å
2
| <r2> |
18.598 |
| (<r2>)1/2 |
4.313 |
