return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for Si3H8 (trisilane)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-871.722770
Energy at 298.15K-871.730603
HF Energy-871.397488
Nuclear repulsion energy193.354324
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 2338 2184 183.82      
2 A1 2323 2171 22.95      
3 A1 2301 2150 96.10      
4 A1 1002 936 85.86      
5 A1 977 912 15.24      
6 A1 942 880 221.05      
7 A1 596 557 9.31      
8 A1 409 382 0.84      
9 A1 101 94 1.69      
10 A2 2333 2180 0.00      
11 A2 991 926 0.00      
12 A2 744 695 0.00      
13 A2 435 406 0.00      
14 A2 83 77 0.00      
15 B1 2338 2185 322.57      
16 B1 2314 2162 40.18      
17 B1 995 930 101.19      
18 B1 628 587 11.19      
19 B1 322 301 25.71      
20 B1 105 98 0.08      
21 B2 2336 2183 99.95      
22 B2 2318 2166 120.90      
23 B2 992 927 42.49      
24 B2 932 871 366.93      
25 B2 765 715 338.23      
26 B2 491 459 4.94      
27 B2 457 427 26.13      

Unscaled Zero Point Vibrational Energy (zpe) 15282.4 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 14279.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.31369 0.06644 0.05810

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Si1 0.000 0.000 0.889
Si2 0.000 1.936 -0.418
Si3 0.000 -1.936 -0.418
H4 1.196 0.000 1.760
H5 -1.196 0.000 1.760
H6 0.000 3.146 0.428
H7 0.000 -3.146 0.428
H8 1.198 1.960 -1.281
H9 -1.198 1.960 -1.281
H10 -1.198 -1.960 -1.281
H11 1.198 -1.960 -1.281

Atom - Atom Distances (Å)
  Si1 Si2 Si3 H4 H5 H6 H7 H8 H9 H10 H11
Si12.33512.33511.48001.48003.17913.17913.15963.15963.15963.1596
Si22.33513.87113.14973.14971.47615.15101.47671.47674.16614.1661
Si32.33513.87113.14973.14975.15101.47614.16614.16611.47671.4767
H41.48003.14973.14972.39213.61953.61953.61814.33834.33833.6181
H51.48003.14973.14972.39213.61953.61954.33833.61813.61814.3383
H63.17911.47615.15103.61953.61956.29112.39982.39985.51565.5156
H73.17915.15101.47613.61953.61956.29115.51565.51562.39982.3998
H83.15961.47674.16613.61814.33832.39985.51562.39574.59433.9203
H93.15961.47674.16614.33833.61812.39985.51562.39573.92034.5943
H103.15964.16611.47674.33833.61815.51562.39984.59433.92032.3957
H113.15964.16611.47673.61814.33835.51562.39983.92034.59432.3957

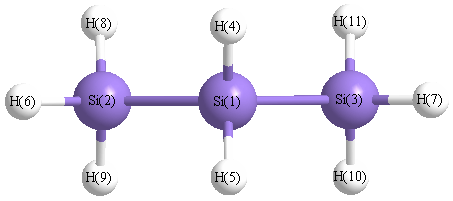
picture of trisilane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 H6 111.045 S1 S2 H8 109.924
S1 S2 H9 109.924 S1 S3 H7 111.045
S1 S3 H10 109.924 S1 S3 H11 109.924
S2 S1 S3 111.972 S2 S1 H4 109.237
S2 S1 H5 109.237 S3 S1 H4 109.237
S3 S1 H5 109.237 H4 S1 H5 107.829
H6 S2 H8 108.732 H6 S2 H9 108.732
H7 S3 H10 108.732 H7 S3 H11 108.732
H8 S2 H9 108.428 H10 S3 H11 108.428
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability