return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H6 (1-Buten-3-yne, 2-methyl-)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-193.450202
Energy at 298.15K-193.454635
HF Energy-192.754252
Nuclear repulsion energy142.681664
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3545 3312 58.99      
2 A' 3351 3131 7.77      
3 A' 3245 3032 2.98      
4 A' 3238 3026 13.73      
5 A' 3129 2923 15.24      
6 A' 2155 2013 1.53      
7 A' 1711 1599 8.01      
8 A' 1557 1455 8.22      
9 A' 1485 1388 0.43      
10 A' 1457 1362 2.08      
11 A' 1327 1240 11.74      
12 A' 1061 991 2.98      
13 A' 998 932 0.11      
14 A' 791 739 1.48      
15 A' 613 573 46.08      
16 A' 556 519 1.98      
17 A' 399 373 0.16      
18 A' 183 171 0.73      
19 A" 3218 3007 13.02      
20 A" 1536 1435 5.74      
21 A" 1102 1030 0.08      
22 A" 903 844 41.33      
23 A" 754 705 1.86      
24 A" 573 536 18.65      
25 A" 505 472 24.87      
26 A" 264 247 2.85      
27 A" 181 169 0.17      

Unscaled Zero Point Vibrational Energy (zpe) 19918.1 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 18611.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.31349 0.13296 0.09499

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.457 0.822 0.000
H2 -1.573 1.903 0.000
H3 -1.957 0.415 0.878
H4 -1.957 0.415 -0.878
C5 0.502 -2.162 0.000
H6 0.701 -3.205 0.000
C7 0.296 -0.957 0.000
C8 0.000 0.444 0.000
C9 0.990 1.354 0.000
H10 2.028 1.057 0.000
H11 0.768 2.411 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 C7 C8 C9 H10 H11
C11.08741.08881.08883.56914.56842.49761.50472.50403.49232.7344
H21.08741.76921.76924.56395.59123.41702.14522.62143.69902.3961
H31.08881.76921.75523.66844.57612.78052.14483.21504.13023.4900
H41.08881.76921.75523.66844.57612.78052.14483.21504.13023.4900
C53.56914.56393.66843.66841.06171.22222.65443.55003.56254.5811
H64.56845.59124.57614.57611.06172.28383.71614.56824.46375.6166
C72.49763.41702.78052.78051.22222.28381.43272.41332.65613.4015
C81.50472.14522.14482.14482.65443.71611.43271.34462.11822.1116
C92.50402.62143.21503.21503.55004.56822.41331.34461.07931.0802
H103.49233.69904.13024.13023.56254.46372.65612.11821.07931.8494
H112.73442.39613.49003.49004.58115.61663.40152.11161.08021.8494

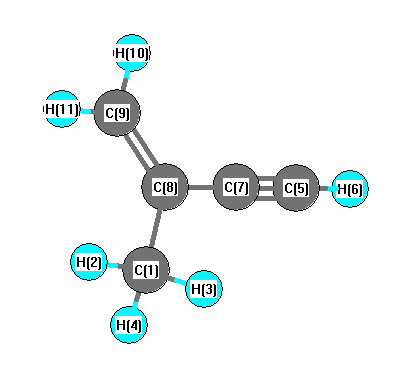
picture of 1-Buten-3-yne, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C8 C7 116.461 C1 C8 C9 122.903
H2 C1 H3 108.780 H2 C1 H4 108.780
H2 C1 C8 110.675 H3 C1 H4 107.412
H3 C1 C8 110.553 H4 C1 C8 110.553
C5 C7 C8 177.733 H6 C5 C7 178.851
C7 C8 C9 120.636 C8 C9 H10 121.447
C8 C9 H11 120.722 H10 C9 H11 117.830
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability