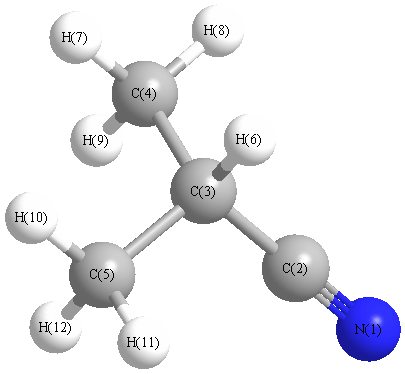
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3173 |
3028 |
16.99 |
|
|
|
| 2 |
A' |
3166 |
3021 |
30.08 |
|
|
|
| 3 |
A' |
3078 |
2937 |
15.68 |
|
|
|
| 4 |
A' |
3070 |
2930 |
3.43 |
|
|
|
| 5 |
A' |
2391 |
2281 |
9.84 |
|
|
|
| 6 |
A' |
1521 |
1451 |
10.09 |
|
|
|
| 7 |
A' |
1511 |
1441 |
8.72 |
|
|
|
| 8 |
A' |
1432 |
1366 |
3.37 |
|
|
|
| 9 |
A' |
1356 |
1294 |
6.34 |
|
|
|
| 10 |
A' |
1203 |
1148 |
3.16 |
|
|
|
| 11 |
A' |
1131 |
1079 |
5.65 |
|
|
|
| 12 |
A' |
948 |
905 |
1.31 |
|
|
|
| 13 |
A' |
786 |
750 |
0.57 |
|
|
|
| 14 |
A' |
559 |
533 |
1.44 |
|
|
|
| 15 |
A' |
354 |
337 |
0.19 |
|
|
|
| 16 |
A' |
289 |
276 |
0.91 |
|
|
|
| 17 |
A' |
222 |
212 |
2.14 |
|
|
|
| 18 |
A" |
3172 |
3027 |
11.36 |
|
|
|
| 19 |
A" |
3163 |
3018 |
0.29 |
|
|
|
| 20 |
A" |
3076 |
2935 |
14.78 |
|
|
|
| 21 |
A" |
1498 |
1430 |
3.86 |
|
|
|
| 22 |
A" |
1496 |
1428 |
0.31 |
|
|
|
| 23 |
A" |
1410 |
1346 |
4.87 |
|
|
|
| 24 |
A" |
1328 |
1267 |
0.36 |
|
|
|
| 25 |
A" |
1147 |
1094 |
2.02 |
|
|
|
| 26 |
A" |
981 |
937 |
0.40 |
|
|
|
| 27 |
A" |
942 |
899 |
2.70 |
|
|
|
| 28 |
A" |
578 |
552 |
0.01 |
|
|
|
| 29 |
A" |
230 |
220 |
0.01 |
|
|
|
| 30 |
A" |
188 |
179 |
4.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22696.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 21659.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.478 |
|
|
|
| 2 |
C |
0.354 |
|
|
|
| 3 |
C |
-0.200 |
|
|
|
| 4 |
C |
-0.378 |
|
|
|
| 5 |
C |
-0.378 |
|
|
|
| 6 |
H |
0.181 |
|
|
|
| 7 |
H |
0.142 |
|
|
|
| 8 |
H |
0.155 |
|
|
|
| 9 |
H |
0.152 |
|
|
|
| 10 |
H |
0.142 |
|
|
|
| 11 |
H |
0.155 |
|
|
|
| 12 |
H |
0.152 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.121 |
3.824 |
0.000 |
3.985 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.145 |
1.737 |
0.000 |
| y |
1.737 |
-38.684 |
0.000 |
| z |
0.000 |
0.000 |
-29.548 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.970 |
1.737 |
0.000 |
| y |
1.737 |
-8.838 |
0.000 |
| z |
0.000 |
0.000 |
4.867 |
|
| Polar |
|---|
| 3z2-r2 | 9.734 |
|---|
| x2-y2 | 8.539 |
|---|
| xy | 1.737 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.584 |
-0.649 |
0.000 |
| y |
-0.649 |
8.136 |
0.000 |
| z |
0.000 |
0.000 |
6.146 |
<r2> (average value of r
2) Å
2
| <r2> |
128.438 |
| (<r2>)1/2 |
11.333 |