return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH(CH3)CN (Propanenitrile, 2-methyl-)

using model chemistry: CCD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/6-31G**
 hartrees
Energy at 0K-210.763731
Energy at 298.15K-210.771063
HF Energy-210.006643
Nuclear repulsion energy158.827066
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3221 3020 22.84      
2 A' 3216 3015 39.72      
3 A' 3131 2935 7.72      
4 A' 3126 2931 12.52      
5 A' 2378 2230 0.53      
6 A' 1571 1473 7.58      
7 A' 1563 1465 5.45      
8 A' 1486 1393 2.18      
9 A' 1405 1317 6.85      
10 A' 1240 1163 1.85      
11 A' 1165 1092 3.75      
12 A' 967 907 0.21      
13 A' 792 742 1.08      
14 A' 554 519 0.86      
15 A' 362 339 0.25      
16 A' 296 277 0.87      
17 A' 223 209 2.54      
18 A" 3219 3018 14.36      
19 A" 3211 3011 0.11      
20 A" 3124 2929 16.19      
21 A" 1553 1456 2.31      
22 A" 1550 1453 0.37      
23 A" 1465 1373 2.08      
24 A" 1386 1300 0.53      
25 A" 1179 1105 1.70      
26 A" 1010 947 0.26      
27 A" 968 908 1.21      
28 A" 569 534 0.03      
29 A" 238 223 0.01      
30 A" 188 176 4.81      

Unscaled Zero Point Vibrational Energy (zpe) 23175.9 cm-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 21729.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/6-31G**
ABC
0.26587 0.13105 0.09613

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.405 -2.189 0.000
C2 0.027 -1.087 0.000
C3 -0.441 0.315 0.000
C4 0.027 1.032 1.272
C5 0.027 1.032 -1.272
H6 -1.533 0.276 0.000
H7 -0.360 2.051 1.281
H8 -0.325 0.519 2.166
H9 1.115 1.076 1.307
H10 -0.360 2.051 -1.281
H11 -0.325 0.519 -2.166
H12 1.115 1.076 -1.307

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12
N11.16462.64233.48373.48373.13564.49463.54333.58754.49463.54333.5875
C21.16461.47782.47212.47212.07183.41172.71932.75173.41172.71932.7517
C32.64231.47781.53381.53381.09312.15932.17842.16972.15932.17842.1697
C43.48372.47211.53382.54502.15041.08971.08881.08972.77673.49412.7997
C53.48372.47211.53382.54502.15042.77673.49412.79971.08971.08881.0897
H63.13562.07181.09312.15042.15042.48332.49163.05942.48332.49163.0594
H74.49463.41172.15931.08972.77672.48331.76911.76852.56293.77243.1345
H83.54332.71932.17841.08883.49412.49161.76911.76743.77244.33153.8004
H93.58752.75172.16971.08972.79973.05941.76851.76743.13453.80042.6132
H104.49463.41172.15932.77671.08972.48332.56293.77243.13451.76911.7685
H113.54332.71932.17843.49411.08882.49163.77244.33153.80041.76911.7674
H123.58752.75172.16972.79971.08973.05943.13453.80042.61321.76851.7674

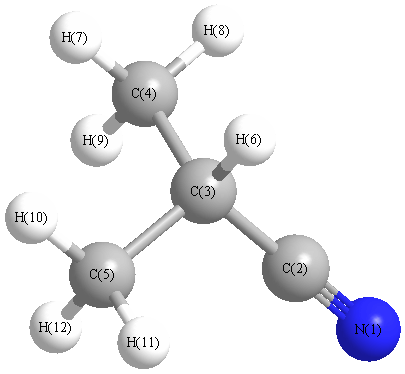
picture of Propanenitrile, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 179.491 C2 C3 C4 110.326
C2 C3 C5 110.326 C2 C3 H6 106.427
C3 C4 H7 109.622 C3 C4 H8 111.195
C3 C4 H9 110.440 C3 C5 H10 109.622
C3 C5 H11 111.195 C3 C5 H12 110.440
C4 C3 C5 112.117 C4 C3 H6 108.727
C5 C3 H6 108.727 H7 C4 H8 108.601
H7 C4 H9 108.477 H8 C4 H9 108.441
H10 C5 H11 108.601 H10 C5 H12 108.477
H11 C5 H12 108.441
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability