return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (1-Propanol, 2-methyl-)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-232.959646
Energy at 298.15K-232.971022
Nuclear repulsion energy193.695366
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3907 3651 20.35      
2 A 3224 3013 34.38      
3 A 3222 3011 16.62      
4 A 3213 3002 51.70      
5 A 3209 2999 11.42      
6 A 3186 2977 29.72      
7 A 3135 2930 18.05      
8 A 3120 2915 16.38      
9 A 3116 2912 21.46      
10 A 3073 2871 46.42      
11 A 1579 1475 10.33      
12 A 1567 1464 4.04      
13 A 1563 1460 0.21      
14 A 1556 1454 4.17      
15 A 1550 1448 0.80      
16 A 1475 1378 13.75      
17 A 1458 1363 1.72      
18 A 1451 1356 19.98      
19 A 1428 1334 4.95      
20 A 1409 1317 8.33      
21 A 1366 1276 1.41      
22 A 1287 1202 12.10      
23 A 1240 1158 0.27      
24 A 1198 1119 2.62      
25 A 1150 1074 5.38      
26 A 1106 1033 114.69      
27 A 1006 940 0.16      
28 A 985 920 7.93      
29 A 958 895 0.35      
30 A 927 867 8.70      
31 A 847 791 6.16      
32 A 502 469 5.76      
33 A 429 401 1.73      
34 A 370 346 21.65      
35 A 323 302 81.24      
36 A 267 250 7.14      
37 A 257 240 32.90      
38 A 233 218 1.88      
39 A 128 120 4.17      

Unscaled Zero Point Vibrational Energy (zpe) 31008.6 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 28974.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.25491 0.11787 0.08881

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.452 0.003 -0.350
C2 -0.766 -0.706 0.229
C3 1.722 -0.747 0.033
C4 0.512 1.456 0.103
O5 -2.003 -0.115 -0.155
H6 0.343 -0.015 -1.437
H7 1.688 -1.782 -0.303
H8 2.599 -0.279 -0.409
H9 1.855 -0.748 1.115
H10 -0.366 2.010 -0.225
H11 0.573 1.516 1.191
H12 1.386 1.958 -0.307
H13 -0.805 -1.731 -0.139
H14 -0.672 -0.754 1.321
H15 -2.092 0.706 0.340

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.52381.52391.52312.46521.09272.17192.16702.16382.17042.16312.16722.15232.15152.7279
C21.52382.49582.51491.42392.11792.73123.45202.76732.78272.76743.46681.08941.09731.9409
C31.52392.49582.51403.78262.14431.08931.08841.09053.46742.78892.74652.71762.71844.0928
C41.52312.51492.51402.97632.13663.46892.76232.77251.08851.09141.08853.45702.78682.7200
O52.46521.42393.78262.97632.67574.05234.61234.11092.68293.33313.97572.01162.08760.9629
H61.09272.11792.14432.13662.67572.49372.49353.05622.46413.05042.50152.43943.03083.0998
H72.17192.73121.08933.46894.05232.49371.76161.76314.31303.78803.75242.49883.04354.5708
H82.16703.45201.08842.76234.61232.49351.76161.76033.75053.14432.54683.71143.73124.8521
H92.16382.76731.09052.77254.11093.05621.76311.76033.78622.60253.09283.10132.53564.2777
H102.17042.78273.46741.08852.68292.46414.31303.75053.78621.76971.75503.76743.18092.2352
H112.16312.76742.78891.09143.33313.05043.78803.14432.60251.76971.76083.77002.59172.9130
H122.16723.46682.74651.08853.97572.50153.75242.54683.09281.75501.76084.29423.77343.7527
H132.15231.08942.71763.45702.01162.43942.49883.71143.10133.76743.77004.29421.76232.7976
H142.15151.09732.71842.78682.08763.03083.04353.73122.53563.18092.59173.77341.76232.2609
H152.72791.94094.09282.72000.96293.09984.57084.85214.27772.23522.91303.75272.79762.2609

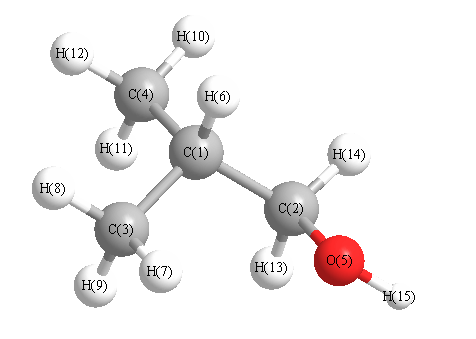
picture of 1-Propanol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O5 113.469 C1 C2 H13 109.784
C1 C2 H14 109.260 C1 C3 H7 111.349
C1 C3 H8 111.003 C1 C3 H9 110.626
C1 C4 H10 111.324 C1 C4 H11 110.562
C1 C4 H12 111.069 C2 C1 C3 109.956
C2 C1 C4 111.259 C2 C1 H6 106.932
C2 O5 H15 107.239 C3 C1 C4 111.192
C3 C1 H6 108.961 C4 C1 H6 108.408
O5 C2 H13 105.562 O5 C2 H14 111.136
H7 C3 H8 107.980 H7 C3 H9 107.968
H8 C3 H9 107.774 H10 C4 H11 108.548
H10 C4 H12 107.445 H11 C4 H12 107.754
H13 C2 H14 107.403
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability