return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H10O (1-Propanol, 2-methyl-)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-232.994765
Energy at 298.15K-233.006113
Nuclear repulsion energy193.106507
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3913 3684 15.42      
2 A 3185 2999 45.51      
3 A 3182 2996 21.29      
4 A 3174 2988 71.39      
5 A 3168 2983 14.35      
6 A 3154 2969 27.84      
7 A 3104 2922 22.44      
8 A 3095 2914 17.37      
9 A 3090 2909 26.21      
10 A 3050 2871 46.46      
11 A 1575 1483 8.39      
12 A 1564 1472 2.22      
13 A 1562 1471 1.58      
14 A 1554 1463 3.86      
15 A 1548 1458 0.60      
16 A 1479 1392 14.19      
17 A 1465 1379 2.93      
18 A 1458 1372 19.51      
19 A 1432 1349 7.61      
20 A 1416 1333 7.70      
21 A 1370 1289 2.26      
22 A 1287 1212 12.41      
23 A 1237 1165 0.25      
24 A 1194 1124 3.34      
25 A 1148 1081 8.03      
26 A 1112 1047 107.93      
27 A 1002 943 0.13      
28 A 980 923 7.46      
29 A 956 900 0.13      
30 A 925 871 6.97      
31 A 841 792 5.48      
32 A 500 471 5.84      
33 A 428 403 1.24      
34 A 368 346 19.50      
35 A 321 302 85.57      
36 A 267 251 9.86      
37 A 253 238 27.26      
38 A 228 215 1.52      
39 A 126 118 4.12      

Unscaled Zero Point Vibrational Energy (zpe) 30855.5 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 29047.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.25288 0.11717 0.08821

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.452 0.003 -0.349
C2 -0.772 -0.708 0.231
C3 1.727 -0.752 0.033
C4 0.516 1.463 0.104
O5 -2.007 -0.114 -0.157
H6 0.344 -0.014 -1.438
H7 1.691 -1.789 -0.307
H8 2.606 -0.283 -0.412
H9 1.864 -0.757 1.117
H10 -0.364 2.018 -0.223
H11 0.580 1.526 1.194
H12 1.392 1.963 -0.311
H13 -0.809 -1.736 -0.136
H14 -0.683 -0.754 1.325
H15 -2.106 0.704 0.340

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.52951.53001.52942.46931.09492.17882.17382.17272.17762.17222.17382.15812.16002.7403
C21.52952.50672.52701.42482.12422.74223.46442.78082.79362.78363.47981.09151.09911.9458
C31.53002.50672.52483.79282.14981.09181.09101.09323.48002.80232.75652.72492.73484.1116
C41.52942.52702.52482.98652.14233.48142.77132.78741.09111.09411.09093.46982.80042.7399
O52.46931.42483.79282.98652.67904.06204.62324.12572.69233.34853.98632.01642.08840.9620
H61.09492.12422.14982.14232.67902.49892.49903.06502.47133.05952.50572.44693.03993.1110
H72.17882.74221.09183.48144.06202.49891.76561.76714.32683.80463.76342.50573.06104.5876
H82.17383.46441.09102.77134.62322.49901.76561.76443.76183.15502.55453.72113.74944.8726
H92.17272.78081.09322.78744.12573.06501.76711.76443.80292.62023.10793.10962.55534.3010
H102.17762.79363.48001.09112.69232.47134.32683.76183.80291.77321.75953.78073.19122.2535
H112.17222.78362.80231.09413.34853.05953.80463.15502.62021.77321.76513.78652.61012.9369
H122.17383.47982.75651.09093.98632.50573.76342.55453.10791.75951.76514.30713.79013.7745
H132.15811.09152.72493.46982.01642.44692.50573.72113.10963.78073.78654.30711.76502.8034
H142.16001.09912.73482.80042.08843.03993.06103.74942.55533.19122.61013.79011.76502.2632
H152.74031.94584.11162.73990.96203.11104.58764.87264.30102.25352.93693.77452.80342.2632

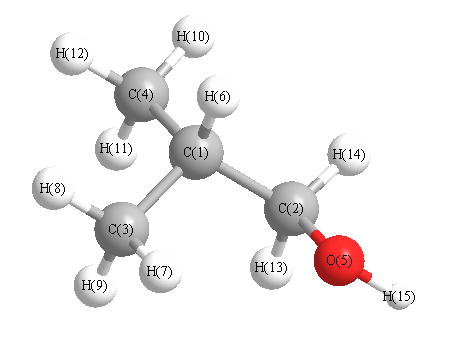
picture of 1-Propanol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O5 113.354 C1 C2 H13 109.718
C1 C2 H14 109.424 C1 C3 H7 111.309
C1 C3 H8 110.961 C1 C3 H9 110.744
C1 C4 H10 111.295 C1 C4 H11 110.680
C1 C4 H12 111.001 C2 C1 C3 110.033
C2 C1 C4 111.399 C2 C1 H6 106.903
C2 O5 H15 107.637 C3 C1 C4 111.227
C3 C1 H6 108.839 C4 C1 H6 108.296
O5 C2 H13 105.750 O5 C2 H14 111.015
H7 C3 H8 107.973 H7 C3 H9 107.944
H8 C3 H9 107.767 H10 C4 H11 108.475
H10 C4 H12 107.486 H11 C4 H12 107.763
H13 C2 H14 107.354
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability