return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H10N2 (1,2-Diaminopropane)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-229.138258
Energy at 298.15K-229.150955
HF Energy-228.326864
Nuclear repulsion energy195.963750
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3685 3443 0.40      
2 A 3669 3428 0.13      
3 A 3572 3338 0.51      
4 A 3569 3335 0.52      
5 A 3234 3022 21.69      
6 A 3218 3007 26.41      
7 A 3193 2984 24.83      
8 A 3127 2922 38.23      
9 A 3124 2919 13.29      
10 A 3020 2822 71.95      
11 A 1701 1590 1.32      
12 A 1689 1578 59.97      
13 A 1564 1461 2.63      
14 A 1553 1451 3.85      
15 A 1534 1434 0.39      
16 A 1469 1373 2.92      
17 A 1456 1361 10.75      
18 A 1441 1346 8.06      
19 A 1417 1324 5.36      
20 A 1382 1291 10.68      
21 A 1300 1214 1.00      
22 A 1249 1167 3.66      
23 A 1208 1129 5.40      
24 A 1151 1076 7.65      
25 A 1080 1009 11.42      
26 A 1062 992 1.96      
27 A 997 931 12.47      
28 A 945 883 124.82      
29 A 919 859 74.00      
30 A 876 819 82.89      
31 A 829 775 56.89      
32 A 509 475 3.67      
33 A 484 452 21.87      
34 A 384 359 49.08      
35 A 378 353 7.82      
36 A 266 249 11.69      
37 A 262 245 23.95      
38 A 238 222 32.64      
39 A 127 119 6.39      

Unscaled Zero Point Vibrational Energy (zpe) 31440.3 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 29377.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.26895 0.11793 0.09005

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.463 1.378 -0.220
H2 -0.298 1.910 0.186
H3 1.311 1.862 0.056
N4 -2.043 -0.133 0.020
H5 -2.096 0.748 -0.477
H6 -2.170 0.071 1.005
C7 -0.734 -0.738 -0.204
H8 -0.742 -1.740 0.230
H9 -0.602 -0.848 -1.281
C10 1.766 -0.663 -0.035
H11 1.777 -1.691 0.321
H12 1.875 -0.664 -1.118
H13 2.625 -0.149 0.394
C14 0.468 0.027 0.346
H15 0.386 0.031 1.444

Atom - Atom Distances (Å)
  N1 H2 H3 N4 H5 H6 C7 H8 H9 C10 H11 H12 H13 C14 H15
N11.01361.01432.93682.64823.18542.43153.37312.68602.42833.38272.63982.71761.46492.1432
H21.01361.61462.69222.24122.74942.71183.67683.13853.30564.15883.61203.58182.03882.3627
H31.01431.61463.90273.62374.02823.31804.14933.57622.56733.59382.84212.42662.03992.4771
N42.93682.69223.90271.01261.01401.45962.07812.06923.84594.13724.11464.68342.53712.8206
H52.64822.24123.62371.01261.63112.03392.91892.32924.13474.64634.26284.88392.78713.2193
H63.18542.74944.02821.01401.63112.04412.43252.92064.13654.37694.62724.83912.71942.5937
C72.43152.71183.31801.45962.03392.04411.09171.09052.50632.73702.76553.46231.52712.1358
H83.37313.67684.14932.07812.91892.43251.09171.76012.74202.52163.13453.72752.14452.4250
H92.68603.13853.57622.06922.32922.92061.09051.76012.68132.98932.48883.70212.13443.0288
C102.42833.30562.56733.84594.13474.13652.50632.74202.68131.08851.08841.08951.51862.1388
H113.38274.15883.59384.13724.64634.37692.73702.52162.98931.08851.77091.76122.16102.4831
H122.63983.61202.84214.11464.26284.62722.76553.13452.48881.08841.77091.76482.14513.0442
H132.71763.58182.42664.68344.88394.83913.46233.72753.70211.08951.76121.76482.16522.4800
C141.46492.03882.03992.53712.78712.71941.52712.14452.13441.51862.16102.14512.16521.1015
H152.14322.36272.47712.82063.21932.59372.13582.42503.02882.13882.48313.04422.48001.1015

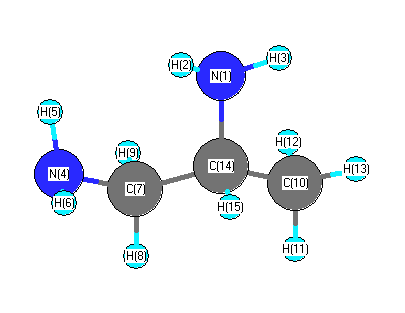
picture of 1,2-Diaminopropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C14 C7 108.700 N1 C14 C10 108.943
N1 C14 H15 112.476 H2 N1 H3 105.537
H2 N1 C14 109.333 H3 N1 C14 109.387
N4 C7 H8 108.214 N4 C7 H9 107.585
N4 C7 C14 116.294 H5 N4 H6 107.187
H5 N4 C7 109.381 H6 N4 C7 110.155
C7 C14 C10 110.758 C7 C14 H15 107.586
H8 C7 H9 107.522 H8 C7 C14 108.812
H9 C7 C14 108.095 C10 C14 H15 108.385
H11 C10 H12 108.879 H11 C10 H13 107.925
H11 C10 C14 110.894 H12 C10 H13 108.258
H12 C10 C14 109.637 H13 C10 C14 111.172
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability