Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Geometric Data calculated at MP2=FULL/6-31G**
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -231.778080 |
| Energy at 298.15K | -231.786388 |
| Nuclear repulsion energy | 176.308115 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3266 |
3052 |
7.37 |
|
|
|
| 2 |
A |
3252 |
3039 |
17.75 |
|
|
|
| 3 |
A |
3244 |
3031 |
18.14 |
|
|
|
| 4 |
A |
3221 |
3010 |
8.66 |
|
|
|
| 5 |
A |
3165 |
2958 |
8.83 |
|
|
|
| 6 |
A |
3148 |
2941 |
18.22 |
|
|
|
| 7 |
A |
3133 |
2927 |
6.47 |
|
|
|
| 8 |
A |
3117 |
2913 |
12.55 |
|
|
|
| 9 |
A |
1792 |
1674 |
74.21 |
|
|
|
| 10 |
A |
1560 |
1458 |
7.07 |
|
|
|
| 11 |
A |
1557 |
1455 |
5.47 |
|
|
|
| 12 |
A |
1539 |
1438 |
9.48 |
|
|
|
| 13 |
A |
1529 |
1429 |
14.30 |
|
|
|
| 14 |
A |
1516 |
1416 |
3.22 |
|
|
|
| 15 |
A |
1470 |
1374 |
4.47 |
|
|
|
| 16 |
A |
1444 |
1349 |
46.69 |
|
|
|
| 17 |
A |
1419 |
1325 |
8.82 |
|
|
|
| 18 |
A |
1323 |
1236 |
0.24 |
|
|
|
| 19 |
A |
1232 |
1151 |
62.42 |
|
|
|
| 20 |
A |
1165 |
1089 |
0.10 |
|
|
|
| 21 |
A |
1144 |
1069 |
0.72 |
|
|
|
| 22 |
A |
1035 |
967 |
3.25 |
|
|
|
| 23 |
A |
981 |
916 |
2.67 |
|
|
|
| 24 |
A |
978 |
913 |
6.56 |
|
|
|
| 25 |
A |
792 |
740 |
2.36 |
|
|
|
| 26 |
A |
780 |
729 |
2.62 |
|
|
|
| 27 |
A |
598 |
559 |
9.36 |
|
|
|
| 28 |
A |
476 |
445 |
0.10 |
|
|
|
| 29 |
A |
409 |
382 |
4.18 |
|
|
|
| 30 |
A |
256 |
239 |
4.75 |
|
|
|
| 31 |
A |
220 |
206 |
0.16 |
|
|
|
| 32 |
A |
110 |
103 |
0.01 |
|
|
|
| 33 |
A |
32 |
30 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25451.0 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 23781.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.884 |
-0.496 |
0.001 |
| C2 |
0.523 |
0.162 |
-0.001 |
| C3 |
-0.677 |
-0.764 |
-0.001 |
| C4 |
-2.001 |
-0.022 |
0.001 |
| O5 |
0.396 |
1.382 |
-0.000 |
| H6 |
2.656 |
0.266 |
-0.002 |
| H7 |
1.994 |
-1.129 |
0.880 |
| H8 |
1.994 |
-1.135 |
-0.875 |
| H9 |
-2.834 |
-0.721 |
-0.001 |
| H10 |
-2.082 |
0.614 |
0.878 |
| H11 |
-2.082 |
0.618 |
-0.874 |
| H12 |
-0.592 |
-1.421 |
-0.871 |
| H13 |
-0.591 |
-1.422 |
0.868 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5113 | 2.5747 | 3.9133 | 2.3954 | 1.0849 | 1.0892 | 1.0892 | 4.7230 | 4.2102 | 4.2105 | 2.7828 | 2.7812 |
C2 | 1.5113 | | 1.5155 | 2.5304 | 1.2266 | 2.1356 | 2.1462 | 2.1466 | 3.4708 | 2.7857 | 2.7846 | 2.1222 | 2.1221 | C3 | 2.5747 | 1.5155 | | 1.5178 | 2.3991 | 3.4885 | 2.8364 | 2.8343 | 2.1574 | 2.1552 | 2.1552 | 1.0930 | 1.0930 | C4 | 3.9133 | 2.5304 | 1.5178 | | 2.7773 | 4.6656 | 4.2380 | 4.2381 | 1.0875 | 1.0864 | 1.0864 | 2.1680 | 2.1680 | O5 | 2.3954 | 1.2266 | 2.3991 | 2.7773 | | 2.5202 | 3.1038 | 3.1064 | 3.8537 | 2.7384 | 2.7358 | 3.0961 | 3.0967 | H6 | 1.0849 | 2.1356 | 3.4885 | 4.6656 | 2.5202 | | 1.7782 | 1.7784 | 5.5778 | 4.8311 | 4.8299 | 3.7616 | 3.7617 | H7 | 1.0892 | 2.1462 | 2.8364 | 4.2380 | 3.1038 | 1.7782 | | 1.7544 | 4.9250 | 4.4332 | 4.7687 | 3.1368 | 2.6022 | H8 | 1.0892 | 2.1466 | 2.8343 | 4.2381 | 3.1064 | 1.7784 | 1.7544 | | 4.9235 | 4.7685 | 4.4362 | 2.6016 | 3.1304 | H9 | 4.7230 | 3.4708 | 2.1574 | 1.0875 | 3.8537 | 5.5778 | 4.9250 | 4.9235 | | 1.7661 | 1.7661 | 2.5044 | 2.5053 | H10 | 4.2102 | 2.7857 | 2.1552 | 1.0864 | 2.7384 | 4.8311 | 4.4332 | 4.7685 | 1.7661 | | 1.7517 | 3.0685 | 2.5235 | H11 | 4.2105 | 2.7846 | 2.1552 | 1.0864 | 2.7358 | 4.8299 | 4.7687 | 4.4362 | 1.7661 | 1.7517 | | 2.5245 | 3.0685 | H12 | 2.7828 | 2.1222 | 1.0930 | 2.1680 | 3.0961 | 3.7616 | 3.1368 | 2.6016 | 2.5044 | 3.0685 | 2.5245 | | 1.7383 | H13 | 2.7812 | 2.1221 | 1.0930 | 2.1680 | 3.0967 | 3.7617 | 2.6022 | 3.1304 | 2.5053 | 2.5235 | 3.0685 | 1.7383 | |
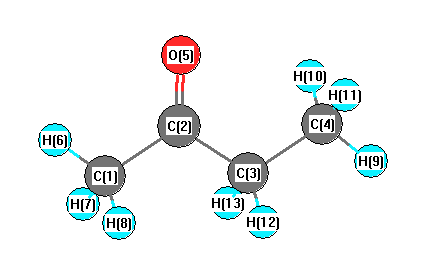 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
116.559 |
|
C1 |
C2 |
O5 |
121.729 |
| C2 |
C1 |
H6 |
109.594 |
|
C2 |
C1 |
H7 |
110.186 |
| C2 |
C1 |
H8 |
110.216 |
|
C2 |
C3 |
C4 |
113.060 |
| C2 |
C3 |
H12 |
107.795 |
|
C2 |
C3 |
H13 |
107.787 |
| C3 |
C2 |
O5 |
121.712 |
|
C3 |
C4 |
H9 |
110.721 |
| C3 |
C4 |
H10 |
110.612 |
|
C3 |
C4 |
H11 |
110.608 |
| C4 |
C3 |
H12 |
111.238 |
|
C4 |
C3 |
H13 |
111.231 |
| H6 |
C1 |
H7 |
109.748 |
|
H6 |
C1 |
H8 |
109.773 |
| H7 |
C1 |
H8 |
107.294 |
|
H9 |
C4 |
H10 |
108.674 |
| H9 |
C4 |
H11 |
108.674 |
|
H10 |
C4 |
H11 |
107.458 |
| H12 |
C3 |
H13 |
105.351 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
