Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -1071.441205 |
| Energy at 298.15K | -1071.443768 |
| HF Energy | -1070.702054 |
| Nuclear repulsion energy | 263.861497 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3246 |
3033 |
0.87 |
|
|
|
| 2 |
A' |
3074 |
2873 |
43.98 |
|
|
|
| 3 |
A' |
1796 |
1678 |
89.68 |
|
|
|
| 4 |
A' |
1442 |
1347 |
2.66 |
|
|
|
| 5 |
A' |
1266 |
1183 |
8.48 |
|
|
|
| 6 |
A' |
1109 |
1037 |
23.16 |
|
|
|
| 7 |
A' |
823 |
769 |
20.86 |
|
|
|
| 8 |
A' |
449 |
419 |
4.09 |
|
|
|
| 9 |
A' |
333 |
311 |
23.78 |
|
|
|
| 10 |
A' |
267 |
250 |
2.19 |
|
|
|
| 11 |
A" |
1314 |
1227 |
28.83 |
|
|
|
| 12 |
A" |
1040 |
972 |
22.03 |
|
|
|
| 13 |
A" |
793 |
741 |
100.40 |
|
|
|
| 14 |
A" |
289 |
270 |
2.36 |
|
|
|
| 15 |
A" |
73 |
69 |
7.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8656.8 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 8088.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.379 |
-0.024 |
0.000 |
| C2 |
-0.172 |
1.391 |
0.000 |
| H3 |
1.463 |
0.003 |
0.000 |
| Cl4 |
-0.172 |
-0.841 |
1.469 |
| Cl5 |
-0.172 |
-0.841 |
-1.469 |
| O6 |
0.552 |
2.368 |
0.000 |
| H7 |
-1.271 |
1.448 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5188 | 1.0842 | 1.7691 | 1.7691 | 2.3984 | 2.2114 |
C2 | 1.5188 | | 2.1449 | 2.6721 | 2.6721 | 1.2159 | 1.1004 | H3 | 1.0842 | 2.1449 | | 2.3549 | 2.3549 | 2.5344 | 3.0925 | Cl4 | 1.7691 | 2.6721 | 2.3549 | | 2.9383 | 3.6028 | 2.9333 | Cl5 | 1.7691 | 2.6721 | 2.3549 | 2.9383 | | 3.6028 | 2.9333 | O6 | 2.3984 | 1.2159 | 2.5344 | 3.6028 | 3.6028 | | 2.0420 | H7 | 2.2114 | 1.1004 | 3.0925 | 2.9333 | 2.9333 | 2.0420 | |
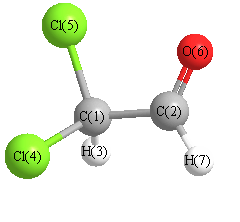 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
122.176 |
|
C1 |
C2 |
H7 |
114.247 |
| C2 |
C1 |
H3 |
109.848 |
|
C2 |
C1 |
Cl4 |
108.480 |
| C2 |
C1 |
Cl5 |
108.480 |
|
H3 |
C1 |
Cl4 |
108.864 |
| H3 |
C1 |
Cl5 |
108.864 |
|
Cl4 |
C1 |
Cl5 |
112.286 |
| O6 |
C2 |
H7 |
123.577 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -1071.439079 |
| Energy at 298.15K | |
| HF Energy | -1070.699333 |
| Nuclear repulsion energy | 266.580306 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3193 |
2984 |
4.58 |
76.23 |
0.25 |
0.40 |
| 2 |
A |
3064 |
2863 |
52.65 |
120.16 |
0.30 |
0.46 |
| 3 |
A |
1802 |
1684 |
76.78 |
7.34 |
0.52 |
0.68 |
| 4 |
A |
1437 |
1343 |
11.98 |
5.24 |
0.60 |
0.75 |
| 5 |
A |
1315 |
1229 |
17.44 |
6.35 |
0.67 |
0.80 |
| 6 |
A |
1288 |
1204 |
11.49 |
9.12 |
0.75 |
0.86 |
| 7 |
A |
1065 |
996 |
13.36 |
1.99 |
0.40 |
0.57 |
| 8 |
A |
971 |
907 |
7.41 |
7.18 |
0.64 |
0.78 |
| 9 |
A |
864 |
808 |
73.09 |
6.51 |
0.71 |
0.83 |
| 10 |
A |
676 |
632 |
31.94 |
7.85 |
0.10 |
0.18 |
| 11 |
A |
630 |
588 |
25.50 |
5.80 |
0.49 |
0.66 |
| 12 |
A |
358 |
335 |
0.86 |
3.08 |
0.26 |
0.41 |
| 13 |
A |
287 |
268 |
3.18 |
4.92 |
0.66 |
0.80 |
| 14 |
A |
229 |
214 |
2.97 |
1.24 |
0.70 |
0.82 |
| 15 |
A |
83 |
77 |
9.90 |
2.13 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 8631.7 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 8065.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.107 |
-0.029 |
0.510 |
| C2 |
-0.725 |
-1.268 |
0.181 |
| H3 |
0.224 |
0.040 |
1.589 |
| Cl4 |
1.732 |
-0.279 |
-0.165 |
| Cl5 |
-0.650 |
1.447 |
-0.057 |
| O6 |
-1.838 |
-1.239 |
-0.302 |
| H7 |
-0.212 |
-2.205 |
0.445 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5279 | 1.0880 | 1.7777 | 1.7525 | 2.4300 | 2.1999 |
C2 | 1.5279 | | 2.1435 | 2.6713 | 2.7260 | 1.2133 | 1.1009 | H3 | 1.0880 | 2.1435 | | 2.3356 | 2.3347 | 3.0763 | 2.5571 | Cl4 | 1.7777 | 2.6713 | 2.3356 | | 2.9434 | 3.6996 | 2.8035 | Cl5 | 1.7525 | 2.7260 | 2.3347 | 2.9434 | | 2.9475 | 3.7120 | O6 | 2.4300 | 1.2133 | 3.0763 | 3.6996 | 2.9475 | | 2.0337 | H7 | 2.1999 | 1.1009 | 2.5571 | 2.8035 | 3.7120 | 2.0337 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.470 |
|
C1 |
C2 |
H7 |
112.608 |
| C2 |
C1 |
H3 |
108.890 |
|
C2 |
C1 |
Cl4 |
107.587 |
| C2 |
C1 |
Cl5 |
112.223 |
|
H3 |
C1 |
Cl4 |
106.689 |
| H3 |
C1 |
Cl5 |
108.275 |
|
Cl4 |
C1 |
Cl5 |
112.972 |
| O6 |
C2 |
H7 |
122.915 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
