Jump to
S1C2
Vibrational Frequencies calculated at PBE1PBE/6-31G**
Geometric Data calculated at PBE1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -1072.505729 |
| Energy at 298.15K | |
| HF Energy | -1072.505729 |
| Nuclear repulsion energy | 266.324823 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3142 |
2994 |
4.25 |
83.42 |
0.24 |
0.38 |
| 2 |
A |
2999 |
2858 |
52.82 |
129.57 |
0.32 |
0.48 |
| 3 |
A |
1896 |
1807 |
133.34 |
14.37 |
0.44 |
0.61 |
| 4 |
A |
1413 |
1346 |
14.11 |
4.67 |
0.67 |
0.80 |
| 5 |
A |
1288 |
1227 |
15.18 |
5.83 |
0.73 |
0.84 |
| 6 |
A |
1251 |
1193 |
11.63 |
10.78 |
0.74 |
0.85 |
| 7 |
A |
1045 |
996 |
13.47 |
2.17 |
0.52 |
0.69 |
| 8 |
A |
951 |
906 |
10.13 |
6.87 |
0.60 |
0.75 |
| 9 |
A |
832 |
793 |
88.17 |
6.38 |
0.67 |
0.80 |
| 10 |
A |
662 |
631 |
40.11 |
8.59 |
0.11 |
0.20 |
| 11 |
A |
619 |
590 |
31.15 |
6.61 |
0.43 |
0.60 |
| 12 |
A |
350 |
333 |
1.41 |
3.37 |
0.24 |
0.39 |
| 13 |
A |
279 |
266 |
3.73 |
5.52 |
0.66 |
0.79 |
| 14 |
A |
221 |
211 |
2.73 |
1.45 |
0.71 |
0.83 |
| 15 |
A |
88 |
84 |
10.22 |
2.06 |
0.72 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 8517.5 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 8118.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.101 |
-0.025 |
0.510 |
| C2 |
-0.713 |
-1.280 |
0.181 |
| H3 |
0.222 |
0.051 |
1.594 |
| Cl4 |
1.738 |
-0.260 |
-0.164 |
| Cl5 |
-0.674 |
1.445 |
-0.057 |
| O6 |
-1.808 |
-1.269 |
-0.303 |
| H7 |
-0.181 |
-2.213 |
0.454 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5310 | 1.0937 | 1.7858 | 1.7560 | 2.4187 | 2.2064 |
C2 | 1.5310 | | 2.1547 | 2.6772 | 2.7350 | 1.1970 | 1.1087 | H3 | 1.0937 | 2.1547 | | 2.3421 | 2.3395 | 3.0762 | 2.5671 | Cl4 | 1.7858 | 2.6772 | 2.3421 | | 2.9552 | 3.6891 | 2.8069 | Cl5 | 1.7560 | 2.7350 | 2.3395 | 2.9552 | | 2.9511 | 3.7261 | O6 | 2.4187 | 1.1970 | 3.0762 | 3.6891 | 2.9511 | | 2.0277 | H7 | 2.2064 | 1.1087 | 2.5671 | 2.8069 | 3.7261 | 2.0277 | |
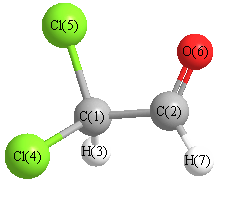 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.457 |
|
C1 |
C2 |
H7 |
112.432 |
| C2 |
C1 |
H3 |
109.224 |
|
C2 |
C1 |
Cl4 |
107.395 |
| C2 |
C1 |
Cl5 |
112.445 |
|
H3 |
C1 |
Cl4 |
106.360 |
| H3 |
C1 |
Cl5 |
108.101 |
|
Cl4 |
C1 |
Cl5 |
113.099 |
| O6 |
C2 |
H7 |
123.107 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.475 |
|
|
|
| 2 |
C |
0.265 |
|
|
|
| 3 |
H |
0.284 |
|
|
|
| 4 |
Cl |
0.019 |
|
|
|
| 5 |
Cl |
0.054 |
|
|
|
| 6 |
O |
-0.325 |
|
|
|
| 7 |
H |
0.179 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.149 |
-0.906 |
1.959 |
2.445 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.329 |
-1.915 |
-1.081 |
| y |
-1.915 |
-41.288 |
-1.424 |
| z |
-1.081 |
-1.424 |
-39.925 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.722 |
-1.915 |
-1.081 |
| y |
-1.915 |
2.338 |
-1.424 |
| z |
-1.081 |
-1.424 |
4.384 |
|
| Polar |
|---|
| 3z2-r2 | 8.768 |
|---|
| x2-y2 | -6.041 |
|---|
| xy | -1.915 |
|---|
| xz | -1.081 |
|---|
| yz | -1.424 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.632 |
-0.720 |
0.175 |
| y |
-0.720 |
6.615 |
-0.314 |
| z |
0.175 |
-0.314 |
4.252 |
<r2> (average value of r
2) Å
2
| <r2> |
185.158 |
| (<r2>)1/2 |
13.607 |
