Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -1072.558462 |
| Energy at 298.15K | -1072.560956 |
| HF Energy | -1072.558462 |
| Nuclear repulsion energy | 263.532294 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3192 |
3046 |
0.74 |
|
|
|
| 2 |
A' |
2999 |
2862 |
44.09 |
|
|
|
| 3 |
A' |
1878 |
1792 |
159.60 |
|
|
|
| 4 |
A' |
1410 |
1345 |
3.79 |
|
|
|
| 5 |
A' |
1223 |
1167 |
7.48 |
|
|
|
| 6 |
A' |
1079 |
1030 |
27.15 |
|
|
|
| 7 |
A' |
797 |
761 |
25.77 |
|
|
|
| 8 |
A' |
444 |
424 |
4.39 |
|
|
|
| 9 |
A' |
324 |
309 |
25.55 |
|
|
|
| 10 |
A' |
259 |
247 |
2.78 |
|
|
|
| 11 |
A" |
1262 |
1204 |
24.77 |
|
|
|
| 12 |
A" |
1025 |
978 |
20.00 |
|
|
|
| 13 |
A" |
746 |
712 |
130.81 |
|
|
|
| 14 |
A" |
275 |
262 |
3.24 |
|
|
|
| 15 |
A" |
81 |
77 |
7.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8496.3 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 8108.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.384 |
-0.021 |
0.000 |
| C2 |
-0.172 |
1.397 |
0.000 |
| H3 |
1.473 |
-0.002 |
0.000 |
| Cl4 |
-0.172 |
-0.841 |
1.475 |
| Cl5 |
-0.172 |
-0.841 |
-1.475 |
| O6 |
0.545 |
2.360 |
0.000 |
| H7 |
-1.278 |
1.451 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5231 | 1.0892 | 1.7767 | 1.7767 | 2.3863 | 2.2203 |
C2 | 1.5231 | | 2.1594 | 2.6801 | 2.6801 | 1.2003 | 1.1076 | H3 | 1.0892 | 2.1594 | | 2.3631 | 2.3631 | 2.5376 | 3.1113 | Cl4 | 1.7767 | 2.6801 | 2.3631 | | 2.9508 | 3.5960 | 2.9414 | Cl5 | 1.7767 | 2.6801 | 2.3631 | 2.9508 | | 3.5960 | 2.9414 | O6 | 2.3863 | 1.2003 | 2.5376 | 3.5960 | 3.5960 | | 2.0372 | H7 | 2.2203 | 1.1076 | 3.1113 | 2.9414 | 2.9414 | 2.0372 | |
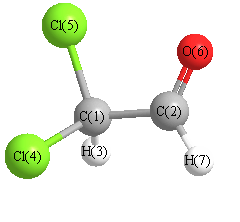 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
121.924 |
|
C1 |
C2 |
H7 |
114.192 |
| C2 |
C1 |
H3 |
110.403 |
|
C2 |
C1 |
Cl4 |
108.377 |
| C2 |
C1 |
Cl5 |
108.377 |
|
H3 |
C1 |
Cl4 |
108.702 |
| H3 |
C1 |
Cl5 |
108.702 |
|
Cl4 |
C1 |
Cl5 |
112.283 |
| O6 |
C2 |
H7 |
123.884 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.418 |
|
|
|
| 2 |
C |
0.326 |
|
|
|
| 3 |
H |
0.251 |
|
|
|
| 4 |
Cl |
0.029 |
|
|
|
| 5 |
Cl |
0.029 |
|
|
|
| 6 |
O |
-0.355 |
|
|
|
| 7 |
H |
0.139 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.262 |
-0.569 |
0.000 |
0.626 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.439 |
-2.827 |
0.000 |
| y |
-2.827 |
-47.929 |
0.000 |
| z |
0.000 |
0.000 |
-42.653 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.852 |
-2.827 |
0.000 |
| y |
-2.827 |
-6.383 |
0.000 |
| z |
0.000 |
0.000 |
1.531 |
|
| Polar |
|---|
| 3z2-r2 | 3.062 |
|---|
| x2-y2 | 7.489 |
|---|
| xy | -2.827 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.988 |
0.700 |
0.000 |
| y |
0.700 |
6.832 |
0.000 |
| z |
0.000 |
0.000 |
7.078 |
<r2> (average value of r
2) Å
2
| <r2> |
191.914 |
| (<r2>)1/2 |
13.853 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -1072.556274 |
| Energy at 298.15K | |
| HF Energy | -1072.556274 |
| Nuclear repulsion energy | 266.232711 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3130 |
2987 |
4.26 |
85.03 |
0.23 |
0.37 |
| 2 |
A |
2982 |
2846 |
51.71 |
131.44 |
0.31 |
0.48 |
| 3 |
A |
1891 |
1805 |
133.00 |
14.51 |
0.44 |
0.61 |
| 4 |
A |
1405 |
1341 |
13.69 |
4.61 |
0.67 |
0.80 |
| 5 |
A |
1274 |
1216 |
14.29 |
5.43 |
0.73 |
0.85 |
| 6 |
A |
1235 |
1179 |
11.94 |
10.75 |
0.74 |
0.85 |
| 7 |
A |
1044 |
996 |
13.11 |
2.20 |
0.53 |
0.69 |
| 8 |
A |
947 |
904 |
10.55 |
6.87 |
0.60 |
0.75 |
| 9 |
A |
827 |
789 |
87.52 |
6.43 |
0.67 |
0.80 |
| 10 |
A |
659 |
629 |
42.03 |
8.59 |
0.12 |
0.21 |
| 11 |
A |
616 |
588 |
30.75 |
6.86 |
0.41 |
0.58 |
| 12 |
A |
349 |
333 |
1.43 |
3.45 |
0.24 |
0.39 |
| 13 |
A |
279 |
266 |
3.71 |
5.55 |
0.66 |
0.79 |
| 14 |
A |
221 |
211 |
2.75 |
1.47 |
0.71 |
0.83 |
| 15 |
A |
88 |
84 |
10.25 |
2.06 |
0.72 |
0.84 |
Unscaled Zero Point Vibrational Energy (zpe) 8473.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 8086.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.100 |
-0.026 |
0.509 |
| C2 |
-0.711 |
-1.281 |
0.179 |
| H3 |
0.221 |
0.050 |
1.593 |
| Cl4 |
1.740 |
-0.257 |
-0.164 |
| Cl5 |
-0.677 |
1.444 |
-0.057 |
| O6 |
-1.807 |
-1.271 |
-0.302 |
| H7 |
-0.173 |
-2.213 |
0.449 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5302 | 1.0934 | 1.7872 | 1.7569 | 2.4175 | 2.2048 |
C2 | 1.5302 | | 2.1537 | 2.6783 | 2.7359 | 1.1970 | 1.1088 | H3 | 1.0934 | 2.1537 | | 2.3423 | 2.3398 | 3.0738 | 2.5661 | Cl4 | 1.7872 | 2.6783 | 2.3423 | | 2.9574 | 3.6915 | 2.8041 | Cl5 | 1.7569 | 2.7359 | 2.3398 | 2.9574 | | 2.9512 | 3.7263 | O6 | 2.4175 | 1.1970 | 3.0738 | 3.6915 | 2.9512 | | 2.0294 | H7 | 2.2048 | 1.1088 | 2.5661 | 2.8041 | 3.7263 | 2.0294 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
124.399 |
|
C1 |
C2 |
H7 |
112.337 |
| C2 |
C1 |
H3 |
109.213 |
|
C2 |
C1 |
Cl4 |
107.419 |
| C2 |
C1 |
Cl5 |
112.493 |
|
H3 |
C1 |
Cl4 |
106.292 |
| H3 |
C1 |
Cl5 |
108.082 |
|
Cl4 |
C1 |
Cl5 |
113.118 |
| O6 |
C2 |
H7 |
123.259 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.429 |
|
|
|
| 2 |
C |
0.314 |
|
|
|
| 3 |
H |
0.245 |
|
|
|
| 4 |
Cl |
0.015 |
|
|
|
| 5 |
Cl |
0.051 |
|
|
|
| 6 |
O |
-0.335 |
|
|
|
| 7 |
H |
0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.153 |
-0.902 |
1.950 |
2.438 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.330 |
-1.924 |
-1.076 |
| y |
-1.924 |
-41.278 |
-1.413 |
| z |
-1.076 |
-1.413 |
-39.917 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.733 |
-1.924 |
-1.076 |
| y |
-1.924 |
2.346 |
-1.413 |
| z |
-1.076 |
-1.413 |
4.387 |
|
| Polar |
|---|
| 3z2-r2 | 8.774 |
|---|
| x2-y2 | -6.052 |
|---|
| xy | -1.924 |
|---|
| xz | -1.076 |
|---|
| yz | -1.413 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.678 |
-0.719 |
0.173 |
| y |
-0.719 |
6.655 |
-0.316 |
| z |
0.173 |
-0.316 |
4.276 |
<r2> (average value of r
2) Å
2
| <r2> |
185.288 |
| (<r2>)1/2 |
13.612 |
