return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H7NO (Propanamide)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-247.816823
Energy at 298.15K-247.825041
HF Energy-247.025490
Nuclear repulsion energy178.915268
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3830 3579 40.94      
2 A 3683 3441 44.21      
3 A 3259 3045 14.66      
4 A 3243 3030 20.59      
5 A 3179 2970 12.92      
6 A 3148 2942 18.81      
7 A 3124 2919 17.10      
8 A 1824 1704 230.87      
9 A 1667 1557 102.47      
10 A 1563 1461 7.94      
11 A 1556 1454 5.21      
12 A 1530 1429 4.28      
13 A 1471 1374 6.31      
14 A 1453 1358 69.21      
15 A 1328 1241 5.53      
16 A 1322 1235 92.69      
17 A 1158 1082 1.20      
18 A 1133 1059 0.21      
19 A 1096 1024 2.94      
20 A 1037 969 3.16      
21 A 838 783 8.21      
22 A 820 766 3.60      
23 A 643 601 9.96      
24 A 611 571 19.92      
25 A 505 472 12.37      
26 A 449 419 3.32      
27 A 276 258 248.54      
28 A 264 247 9.60      
29 A 219 204 0.88      
30 A 25 23 2.38      

Unscaled Zero Point Vibrational Energy (zpe) 23125.1 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 21608.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.32506 0.12775 0.09552

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
H1 -2.582 -0.006 -0.017
H2 -1.788 -1.531 0.058
N3 -1.738 -0.544 -0.114
O4 -0.504 1.361 0.052
C5 -0.555 0.135 0.023
H6 0.671 -1.223 1.100
H7 0.588 -1.557 -0.614
C8 0.676 -0.750 0.115
H9 2.826 -0.610 0.023
H10 1.982 0.476 -1.084
H11 2.024 0.845 0.629
C12 1.959 0.036 -0.091

Atom - Atom Distances (Å)
  H1 H2 N3 O4 C5 H6 H7 C8 H9 H10 H11 C12
H11.72071.00552.48862.03233.64873.57903.34425.44194.71244.72854.5414
H21.72071.00363.16452.07282.68872.46952.58534.70534.42174.52844.0639
N31.00551.00362.27551.37022.78192.58552.43304.56623.97754.07823.7414
O42.48863.16452.27551.22793.02663.18592.41943.87002.87342.64382.8003
C52.03232.07281.37021.22792.12312.13861.51863.46222.78962.74292.5182
H63.64872.68872.78193.02662.12311.74841.09262.48583.06232.51632.1589
H73.57902.46952.58553.18592.13861.74841.09112.51212.50953.06242.1653
C83.34422.58532.43302.41941.51861.09261.09112.15692.15632.15131.5185
H95.44194.70534.56623.87003.46222.48582.51212.15691.76551.76921.0876
H104.71244.42173.97752.87342.78963.06232.50952.15631.76551.75291.0867
H114.72854.52844.07822.64382.74292.51633.06242.15131.76921.75291.0855
C124.54144.06393.74142.80032.51822.15892.16531.51851.08761.08671.0855

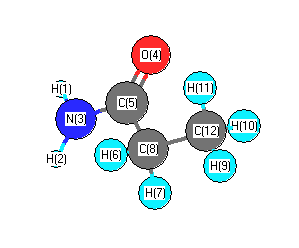
picture of Propanamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
H1 N3 H2 117.834 H1 N3 C5 116.781
H2 N3 C5 120.886 N3 C5 O4 122.186
N3 C5 C8 114.652 O4 C5 C8 123.155
C5 C8 H6 107.675 C5 C8 H7 108.970
C5 C8 C12 112.021 H6 C8 H7 106.389
H6 C8 C12 110.481 H7 C8 C12 111.088
C8 C12 H9 110.626 C8 C12 H10 110.633
C8 C12 H11 110.304 H9 C12 H10 108.588
H9 C12 H11 109.007 H10 C12 H11 107.607
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability