return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3COOCH3 (methyl acetate)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-267.641835
Energy at 298.15K-267.648679
HF Energy-266.843290
Nuclear repulsion energy180.855659
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3285 3069 9.73      
2 A' 3284 3069 9.29      
3 A' 3155 2948 3.83      
4 A' 3151 2945 24.79      
5 A' 1841 1720 178.08      
6 A' 1561 1459 6.77      
7 A' 1535 1434 12.70      
8 A' 1521 1421 9.34      
9 A' 1451 1356 58.94      
10 A' 1321 1234 331.63      
11 A' 1237 1156 2.68      
12 A' 1112 1039 66.54      
13 A' 1017 950 2.08      
14 A' 877 819 20.78      
15 A' 654 611 7.21      
16 A' 429 401 6.07      
17 A' 292 273 11.87      
18 A" 3253 3039 20.10      
19 A" 3249 3036 3.69      
20 A" 1551 1449 5.34      
21 A" 1539 1438 6.59      
22 A" 1207 1128 1.68      
23 A" 1094 1022 5.75      
24 A" 608 568 6.56      
25 A" 185 173 6.78      
26 A" 152 142 0.48      
27 A" 69 65 0.52      

Unscaled Zero Point Vibrational Energy (zpe) 20313.9 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 18981.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.33707 0.14011 0.10275

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.119 1.495 0.000
C2 0.000 0.492 0.000
O3 -1.191 0.746 0.000
O4 0.496 -0.771 0.000
C5 -0.515 -1.794 0.000
H6 0.703 2.495 0.000
H7 1.742 1.348 0.878
H8 1.742 1.348 -0.878
H9 0.028 -2.731 0.000
H10 -1.140 -1.711 0.884
H11 -1.140 -1.711 -0.884

Atom - Atom Distances (Å)
  C1 C2 O3 O4 C5 H6 H7 H8 H9 H10 H11
C11.50232.42782.34993.67161.08351.08681.08684.36414.01964.0196
C21.50231.21761.35692.34292.12302.13062.13063.22322.63312.6331
O32.42781.21762.26852.62852.57793.12033.12033.68482.61172.6117
O42.34991.35692.26851.43733.27292.61072.61072.01492.08312.0831
C53.67162.34292.62851.43734.45843.96663.96661.08321.08611.0861
H61.08352.12302.57793.27294.45841.77971.77975.26964.67634.6763
H71.08682.13063.12032.61073.96661.77971.75594.51084.20274.5572
H81.08682.13063.12032.61073.96661.77971.75594.51084.55724.2027
H94.36413.22323.68482.01491.08325.26964.51084.51081.78531.7853
H104.01962.63312.61172.08311.08614.67634.20274.55721.78531.7682
H114.01962.63312.61172.08311.08614.67634.55724.20271.78531.7682

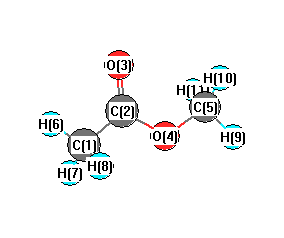
picture of methyl acetate state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 126.082 C1 C2 O4 110.446
C2 C1 H6 109.306 C2 C1 H7 109.708
C2 C1 H8 109.708 C2 O4 C5 113.933
O3 C2 O4 123.473 O4 C5 H9 105.279
O4 C5 H10 110.504 O4 C5 H11 110.504
H6 C1 H7 110.171 H6 C1 H8 110.171
H7 C1 H8 107.760 H9 C5 H10 110.772
H9 C5 H11 110.772 H10 C5 H11 108.981
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability