Jump to
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -216.524771 |
| Energy at 298.15K | |
| HF Energy | -215.925392 |
| Nuclear repulsion energy | 116.672375 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3368 |
3147 |
4.97 |
44.80 |
0.74 |
0.85 |
| 2 |
A' |
3267 |
3053 |
5.37 |
135.54 |
0.12 |
0.22 |
| 3 |
A' |
3250 |
3037 |
7.37 |
27.87 |
0.68 |
0.81 |
| 4 |
A' |
3119 |
2914 |
39.32 |
132.41 |
0.11 |
0.19 |
| 5 |
A' |
1751 |
1636 |
1.61 |
3.98 |
0.11 |
0.20 |
| 6 |
A' |
1571 |
1468 |
1.02 |
17.25 |
0.64 |
0.78 |
| 7 |
A' |
1494 |
1396 |
7.94 |
8.82 |
0.51 |
0.68 |
| 8 |
A' |
1466 |
1370 |
17.07 |
6.72 |
0.74 |
0.85 |
| 9 |
A' |
1343 |
1255 |
0.28 |
13.08 |
0.39 |
0.56 |
| 10 |
A' |
1167 |
1091 |
41.96 |
2.42 |
0.71 |
0.83 |
| 11 |
A' |
1038 |
970 |
29.61 |
4.69 |
0.75 |
0.86 |
| 12 |
A' |
945 |
883 |
2.74 |
4.54 |
0.16 |
0.27 |
| 13 |
A' |
615 |
574 |
5.09 |
1.31 |
0.74 |
0.85 |
| 14 |
A' |
283 |
264 |
3.00 |
0.67 |
0.51 |
0.67 |
| 15 |
A" |
3173 |
2965 |
38.08 |
81.10 |
0.75 |
0.86 |
| 16 |
A" |
1291 |
1207 |
0.04 |
10.49 |
0.75 |
0.86 |
| 17 |
A" |
1082 |
1011 |
12.19 |
0.19 |
0.75 |
0.86 |
| 18 |
A" |
1044 |
976 |
14.84 |
0.00 |
0.75 |
0.86 |
| 19 |
A" |
946 |
884 |
33.48 |
0.09 |
0.75 |
0.86 |
| 20 |
A" |
565 |
528 |
7.04 |
11.49 |
0.75 |
0.86 |
| 21 |
A" |
170 |
158 |
2.97 |
5.08 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16473.2 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 15392.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.950 |
-0.193 |
0.000 |
| C2 |
0.000 |
0.959 |
0.000 |
| C3 |
1.326 |
0.819 |
0.000 |
| F4 |
-0.264 |
-1.404 |
0.000 |
| H5 |
1.979 |
1.678 |
0.000 |
| H6 |
1.778 |
-0.160 |
0.000 |
| H7 |
-0.456 |
1.942 |
0.000 |
| H8 |
-1.591 |
-0.167 |
0.883 |
| H9 |
-1.591 |
-0.167 |
-0.883 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4931 | 2.4907 | 1.3915 | 3.4750 | 2.7279 | 2.1910 | 1.0918 | 1.0918 |
C2 | 1.4931 | | 1.3335 | 2.3780 | 2.1053 | 2.1008 | 1.0834 | 2.1400 | 2.1400 | C3 | 2.4907 | 1.3335 | | 2.7335 | 1.0786 | 1.0781 | 2.1066 | 3.2034 | 3.2034 | F4 | 1.3915 | 2.3780 | 2.7335 | | 3.8119 | 2.3917 | 3.3514 | 2.0172 | 2.0172 | H5 | 3.4750 | 2.1053 | 1.0786 | 3.8119 | | 1.8485 | 2.4495 | 4.1142 | 4.1142 | H6 | 2.7279 | 2.1008 | 1.0781 | 2.3917 | 1.8485 | | 3.0674 | 3.4827 | 3.4827 | H7 | 2.1910 | 1.0834 | 2.1066 | 3.3514 | 2.4495 | 3.0674 | | 2.5525 | 2.5525 | H8 | 1.0918 | 2.1400 | 3.2034 | 2.0172 | 4.1142 | 3.4827 | 2.5525 | | 1.7666 | H9 | 1.0918 | 2.1400 | 3.2034 | 2.0172 | 4.1142 | 3.4827 | 2.5525 | 1.7666 | |
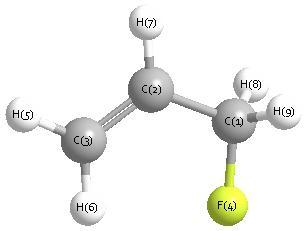 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.462 |
|
C1 |
C2 |
H7 |
115.591 |
| C2 |
C1 |
F4 |
111.006 |
|
C2 |
C1 |
H8 |
110.813 |
| C2 |
C1 |
H9 |
110.813 |
|
C2 |
C3 |
H5 |
121.203 |
| C2 |
C3 |
H6 |
120.817 |
|
C3 |
C2 |
H7 |
120.947 |
| F4 |
C1 |
H8 |
108.040 |
|
F4 |
C1 |
H9 |
108.040 |
| H5 |
C3 |
H6 |
117.980 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -216.523647 |
| Energy at 298.15K | |
| HF Energy | -215.924485 |
| Nuclear repulsion energy | 114.184489 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3347 |
3127 |
10.91 |
52.24 |
0.63 |
0.77 |
| 2 |
A |
3267 |
3053 |
5.54 |
99.48 |
0.23 |
0.37 |
| 3 |
A |
3243 |
3030 |
6.84 |
57.09 |
0.12 |
0.22 |
| 4 |
A |
3192 |
2983 |
33.25 |
59.89 |
0.75 |
0.86 |
| 5 |
A |
3127 |
2922 |
37.65 |
99.99 |
0.10 |
0.18 |
| 6 |
A |
1747 |
1632 |
0.45 |
4.40 |
0.12 |
0.22 |
| 7 |
A |
1573 |
1470 |
0.52 |
9.23 |
0.70 |
0.83 |
| 8 |
A |
1513 |
1414 |
18.09 |
9.70 |
0.56 |
0.72 |
| 9 |
A |
1439 |
1345 |
21.90 |
4.31 |
0.68 |
0.81 |
| 10 |
A |
1342 |
1254 |
0.09 |
10.55 |
0.47 |
0.64 |
| 11 |
A |
1298 |
1213 |
3.24 |
13.58 |
0.74 |
0.85 |
| 12 |
A |
1217 |
1137 |
2.72 |
2.10 |
0.68 |
0.81 |
| 13 |
A |
1096 |
1025 |
93.57 |
5.87 |
0.50 |
0.67 |
| 14 |
A |
1047 |
979 |
33.10 |
0.64 |
0.54 |
0.70 |
| 15 |
A |
1019 |
952 |
4.97 |
1.46 |
0.17 |
0.29 |
| 16 |
A |
960 |
897 |
30.43 |
0.61 |
0.19 |
0.31 |
| 17 |
A |
955 |
892 |
3.66 |
1.99 |
0.19 |
0.33 |
| 18 |
A |
662 |
619 |
5.87 |
6.45 |
0.69 |
0.82 |
| 19 |
A |
442 |
413 |
1.91 |
3.78 |
0.53 |
0.69 |
| 20 |
A |
333 |
311 |
6.61 |
4.20 |
0.75 |
0.86 |
| 21 |
A |
112 |
105 |
1.03 |
5.44 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16465.6 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 15385.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.602 |
0.439 |
0.310 |
| C2 |
0.640 |
-0.380 |
0.248 |
| C3 |
1.802 |
0.088 |
-0.214 |
| F4 |
-1.645 |
-0.231 |
-0.338 |
| H5 |
2.695 |
-0.518 |
-0.219 |
| H6 |
1.889 |
1.094 |
-0.598 |
| H7 |
0.563 |
-1.393 |
0.623 |
| H8 |
-0.916 |
0.607 |
1.342 |
| H9 |
-0.458 |
1.405 |
-0.175 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4897 | 2.4856 | 1.3989 | 3.4740 | 2.7313 | 2.1942 | 1.0911 | 1.0901 |
C2 | 1.4897 | | 1.3352 | 2.3640 | 2.1121 | 2.1099 | 1.0825 | 2.1426 | 2.1380 | C3 | 2.4856 | 1.3352 | | 3.4639 | 1.0791 | 1.0811 | 2.1042 | 3.1741 | 2.6156 | F4 | 1.3989 | 2.3640 | 3.4639 | | 4.3515 | 3.7834 | 2.6744 | 2.0139 | 2.0280 | H5 | 3.4740 | 2.1121 | 1.0791 | 4.3515 | | 1.8420 | 2.4538 | 4.0914 | 3.6930 | H6 | 2.7313 | 2.1099 | 1.0811 | 3.7834 | 1.8420 | | 3.0720 | 3.4446 | 2.4044 | H7 | 2.1942 | 1.0825 | 2.1042 | 2.6744 | 2.4538 | 3.0720 | | 2.5893 | 3.0835 | H8 | 1.0911 | 2.1426 | 3.1741 | 2.0139 | 4.0914 | 3.4446 | 2.5893 | | 1.7735 | H9 | 1.0901 | 2.1380 | 2.6156 | 2.0280 | 3.6930 | 2.4044 | 3.0835 | 1.7735 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.163 |
|
C1 |
C2 |
H7 |
116.189 |
| C2 |
C1 |
F4 |
109.808 |
|
C2 |
C1 |
H8 |
111.310 |
| C2 |
C1 |
H9 |
110.997 |
|
C2 |
C3 |
H5 |
121.686 |
| C2 |
C3 |
H6 |
121.306 |
|
C3 |
C2 |
H7 |
120.636 |
| F4 |
C1 |
H8 |
107.317 |
|
F4 |
C1 |
H9 |
108.494 |
| H5 |
C3 |
H6 |
117.007 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
