Jump to
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -216.887190 |
| Energy at 298.15K | |
| HF Energy | -216.887190 |
| Nuclear repulsion energy | 116.763124 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3292 |
3137 |
6.09 |
52.11 |
0.72 |
0.84 |
| 2 |
A' |
3206 |
3056 |
4.29 |
139.95 |
0.12 |
0.22 |
| 3 |
A' |
3188 |
3038 |
10.92 |
37.14 |
0.60 |
0.75 |
| 4 |
A' |
3044 |
2901 |
42.52 |
158.96 |
0.11 |
0.20 |
| 5 |
A' |
1761 |
1678 |
3.65 |
16.76 |
0.15 |
0.26 |
| 6 |
A' |
1531 |
1459 |
3.00 |
19.39 |
0.60 |
0.75 |
| 7 |
A' |
1464 |
1395 |
9.75 |
15.75 |
0.51 |
0.67 |
| 8 |
A' |
1440 |
1373 |
14.68 |
8.77 |
0.73 |
0.85 |
| 9 |
A' |
1326 |
1264 |
0.26 |
17.44 |
0.39 |
0.56 |
| 10 |
A' |
1169 |
1114 |
53.97 |
2.29 |
0.75 |
0.85 |
| 11 |
A' |
1030 |
982 |
28.46 |
4.98 |
0.74 |
0.85 |
| 12 |
A' |
933 |
890 |
0.90 |
3.99 |
0.16 |
0.27 |
| 13 |
A' |
614 |
585 |
4.30 |
1.51 |
0.72 |
0.84 |
| 14 |
A' |
270 |
258 |
2.24 |
0.93 |
0.54 |
0.70 |
| 15 |
A" |
3080 |
2935 |
40.56 |
99.73 |
0.75 |
0.86 |
| 16 |
A" |
1275 |
1215 |
0.01 |
11.37 |
0.75 |
0.86 |
| 17 |
A" |
1059 |
1009 |
9.94 |
0.63 |
0.75 |
0.86 |
| 18 |
A" |
1033 |
985 |
12.47 |
0.02 |
0.75 |
0.86 |
| 19 |
A" |
954 |
909 |
43.82 |
0.53 |
0.75 |
0.86 |
| 20 |
A" |
563 |
536 |
9.42 |
11.72 |
0.75 |
0.86 |
| 21 |
A" |
180 |
172 |
2.84 |
5.67 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16205.0 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 15445.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.941 |
-0.205 |
0.000 |
| C2 |
0.000 |
0.953 |
0.000 |
| C3 |
1.324 |
0.836 |
0.000 |
| F4 |
-0.267 |
-1.406 |
0.000 |
| H5 |
1.971 |
1.707 |
0.000 |
| H6 |
1.797 |
-0.141 |
0.000 |
| H7 |
-0.476 |
1.934 |
0.000 |
| H8 |
-1.592 |
-0.174 |
0.886 |
| H9 |
-1.592 |
-0.174 |
-0.886 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4926 | 2.4926 | 1.3766 | 3.4838 | 2.7391 | 2.1890 | 1.0998 | 1.0998 |
C2 | 1.4926 | | 1.3288 | 2.3738 | 2.1100 | 2.1041 | 1.0900 | 2.1422 | 2.1422 | C3 | 2.4926 | 1.3288 | | 2.7484 | 1.0856 | 1.0856 | 2.1080 | 3.2100 | 3.2100 | F4 | 1.3766 | 2.3738 | 2.7484 | | 3.8338 | 2.4207 | 3.3458 | 2.0145 | 2.0145 | H5 | 3.4838 | 2.1100 | 1.0856 | 3.8338 | | 1.8567 | 2.4569 | 4.1249 | 4.1249 | H6 | 2.7391 | 2.1041 | 1.0856 | 2.4207 | 1.8567 | | 3.0777 | 3.5032 | 3.5032 | H7 | 2.1890 | 1.0900 | 2.1080 | 3.3458 | 2.4569 | 3.0777 | | 2.5439 | 2.5439 | H8 | 1.0998 | 2.1422 | 3.2100 | 2.0145 | 4.1249 | 3.5032 | 2.5439 | | 1.7722 | H9 | 1.0998 | 2.1422 | 3.2100 | 2.0145 | 4.1249 | 3.5032 | 2.5439 | 1.7722 | |
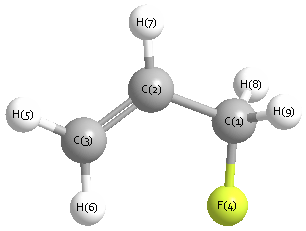 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.023 |
|
C1 |
C2 |
H7 |
115.017 |
| C2 |
C1 |
F4 |
111.586 |
|
C2 |
C1 |
H8 |
110.534 |
| C2 |
C1 |
H9 |
110.534 |
|
C2 |
C3 |
H5 |
121.511 |
| C2 |
C3 |
H6 |
120.936 |
|
C3 |
C2 |
H7 |
120.959 |
| F4 |
C1 |
H8 |
108.351 |
|
F4 |
C1 |
H9 |
108.351 |
| H5 |
C3 |
H6 |
117.553 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.020 |
|
|
|
| 2 |
C |
-0.161 |
|
|
|
| 3 |
C |
-0.377 |
|
|
|
| 4 |
F |
-0.308 |
|
|
|
| 5 |
H |
0.172 |
|
|
|
| 6 |
H |
0.188 |
|
|
|
| 7 |
H |
0.169 |
|
|
|
| 8 |
H |
0.168 |
|
|
|
| 9 |
H |
0.168 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.832 |
1.356 |
0.000 |
1.591 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.753 |
-0.370 |
0.000 |
| y |
-0.370 |
-24.489 |
0.000 |
| z |
0.000 |
0.000 |
-24.742 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.863 |
-0.370 |
0.000 |
| y |
-0.370 |
-1.742 |
0.000 |
| z |
0.000 |
0.000 |
-2.122 |
|
| Polar |
|---|
| 3z2-r2 | -4.243 |
|---|
| x2-y2 | 3.737 |
|---|
| xy | -0.370 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.559 |
0.263 |
0.000 |
| y |
0.263 |
4.532 |
0.000 |
| z |
0.000 |
0.000 |
2.882 |
<r2> (average value of r
2) Å
2
| <r2> |
79.426 |
| (<r2>)1/2 |
8.912 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -216.885787 |
| Energy at 298.15K | |
| HF Energy | -216.885787 |
| Nuclear repulsion energy | 114.274725 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3271 |
3118 |
12.09 |
61.78 |
0.60 |
0.75 |
| 2 |
A |
3200 |
3050 |
5.94 |
116.96 |
0.24 |
0.39 |
| 3 |
A |
3182 |
3033 |
10.22 |
52.45 |
0.12 |
0.22 |
| 4 |
A |
3110 |
2965 |
33.99 |
66.06 |
0.71 |
0.83 |
| 5 |
A |
3056 |
2912 |
40.44 |
120.96 |
0.12 |
0.22 |
| 6 |
A |
1758 |
1676 |
0.44 |
16.75 |
0.15 |
0.26 |
| 7 |
A |
1534 |
1462 |
0.82 |
10.54 |
0.67 |
0.80 |
| 8 |
A |
1483 |
1413 |
22.91 |
11.52 |
0.55 |
0.71 |
| 9 |
A |
1416 |
1350 |
19.26 |
6.13 |
0.65 |
0.79 |
| 10 |
A |
1325 |
1263 |
0.10 |
14.19 |
0.47 |
0.64 |
| 11 |
A |
1280 |
1220 |
3.42 |
14.86 |
0.73 |
0.84 |
| 12 |
A |
1196 |
1140 |
1.76 |
1.89 |
0.71 |
0.83 |
| 13 |
A |
1106 |
1054 |
107.15 |
5.59 |
0.53 |
0.69 |
| 14 |
A |
1043 |
994 |
24.00 |
0.98 |
0.71 |
0.83 |
| 15 |
A |
1004 |
957 |
2.07 |
1.23 |
0.19 |
0.32 |
| 16 |
A |
965 |
920 |
46.81 |
0.55 |
0.48 |
0.65 |
| 17 |
A |
938 |
894 |
2.75 |
2.41 |
0.20 |
0.34 |
| 18 |
A |
655 |
624 |
7.19 |
6.97 |
0.70 |
0.82 |
| 19 |
A |
441 |
420 |
2.47 |
4.71 |
0.53 |
0.69 |
| 20 |
A |
322 |
307 |
5.93 |
3.96 |
0.75 |
0.86 |
| 21 |
A |
116 |
111 |
0.96 |
6.54 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16199.7 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 15440.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.606 |
0.441 |
0.289 |
| C2 |
0.639 |
-0.378 |
0.229 |
| C3 |
1.811 |
0.084 |
-0.199 |
| F4 |
-1.652 |
-0.232 |
-0.318 |
| H5 |
2.706 |
-0.531 |
-0.192 |
| H6 |
1.922 |
1.101 |
-0.570 |
| H7 |
0.545 |
-1.401 |
0.590 |
| H8 |
-0.898 |
0.630 |
1.332 |
| H9 |
-0.468 |
1.407 |
-0.211 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4917 | 2.4919 | 1.3843 | 3.4857 | 2.7507 | 2.1931 | 1.0990 | 1.0970 |
C2 | 1.4917 | | 1.3300 | 2.3607 | 2.1149 | 2.1141 | 1.0895 | 2.1434 | 2.1458 | C3 | 2.4919 | 1.3300 | | 3.4798 | 1.0863 | 1.0884 | 2.1047 | 3.1591 | 2.6351 | F4 | 1.3843 | 2.3607 | 3.4798 | | 4.3707 | 3.8230 | 2.6500 | 2.0089 | 2.0252 | H5 | 3.4857 | 2.1149 | 1.0863 | 4.3707 | | 1.8501 | 2.4571 | 4.0817 | 3.7192 | H6 | 2.7507 | 2.1141 | 1.0884 | 3.8230 | 1.8501 | | 3.0825 | 3.4338 | 2.4357 | H7 | 2.1931 | 1.0895 | 2.1047 | 2.6500 | 2.4571 | 3.0825 | | 2.5994 | 3.0910 | H8 | 1.0990 | 2.1434 | 3.1591 | 2.0089 | 4.0817 | 3.4338 | 2.5994 | | 1.7807 | H9 | 1.0970 | 2.1458 | 2.6351 | 2.0252 | 3.7192 | 2.4357 | 3.0910 | 1.7807 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.940 |
|
C1 |
C2 |
H7 |
115.464 |
| C2 |
C1 |
F4 |
110.275 |
|
C2 |
C1 |
H8 |
110.745 |
| C2 |
C1 |
H9 |
111.057 |
|
C2 |
C3 |
H5 |
121.828 |
| C2 |
C3 |
H6 |
121.580 |
|
C3 |
C2 |
H7 |
120.577 |
| F4 |
C1 |
H8 |
107.434 |
|
F4 |
C1 |
H9 |
108.858 |
| H5 |
C3 |
H6 |
116.592 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.031 |
|
|
|
| 2 |
C |
-0.128 |
|
|
|
| 3 |
C |
-0.385 |
|
|
|
| 4 |
F |
-0.316 |
|
|
|
| 5 |
H |
0.180 |
|
|
|
| 6 |
H |
0.173 |
|
|
|
| 7 |
H |
0.181 |
|
|
|
| 8 |
H |
0.165 |
|
|
|
| 9 |
H |
0.161 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.376 |
0.685 |
0.766 |
1.718 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.919 |
-0.974 |
-1.556 |
| y |
-0.974 |
-21.967 |
-0.893 |
| z |
-1.556 |
-0.893 |
-24.320 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.776 |
-0.974 |
-1.556 |
| y |
-0.974 |
2.652 |
-0.893 |
| z |
-1.556 |
-0.893 |
-0.877 |
|
| Polar |
|---|
| 3z2-r2 | -1.753 |
|---|
| x2-y2 | -2.952 |
|---|
| xy | -0.974 |
|---|
| xz | -1.556 |
|---|
| yz | -0.893 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.295 |
0.176 |
-0.865 |
| y |
0.176 |
4.438 |
-0.545 |
| z |
-0.865 |
-0.545 |
3.259 |
<r2> (average value of r
2) Å
2
| <r2> |
88.953 |
| (<r2>)1/2 |
9.431 |
