Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -216.913359 |
| Energy at 298.15K | |
| HF Energy | -216.913359 |
| Nuclear repulsion energy | 116.794492 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3287 |
3136 |
5.52 |
52.90 |
0.72 |
0.84 |
| 2 |
A' |
3196 |
3050 |
4.52 |
148.69 |
0.12 |
0.22 |
| 3 |
A' |
3180 |
3034 |
9.65 |
31.66 |
0.68 |
0.81 |
| 4 |
A' |
3033 |
2894 |
41.99 |
163.50 |
0.11 |
0.20 |
| 5 |
A' |
1755 |
1674 |
3.78 |
17.62 |
0.15 |
0.26 |
| 6 |
A' |
1511 |
1442 |
3.49 |
18.30 |
0.61 |
0.76 |
| 7 |
A' |
1450 |
1384 |
10.15 |
16.32 |
0.51 |
0.68 |
| 8 |
A' |
1426 |
1361 |
12.13 |
8.12 |
0.74 |
0.85 |
| 9 |
A' |
1318 |
1258 |
0.31 |
18.14 |
0.39 |
0.56 |
| 10 |
A' |
1162 |
1109 |
54.42 |
2.19 |
0.75 |
0.86 |
| 11 |
A' |
1023 |
977 |
28.04 |
5.15 |
0.74 |
0.85 |
| 12 |
A' |
930 |
888 |
0.90 |
4.02 |
0.16 |
0.27 |
| 13 |
A' |
613 |
585 |
4.48 |
1.50 |
0.72 |
0.84 |
| 14 |
A' |
269 |
256 |
2.33 |
0.94 |
0.53 |
0.69 |
| 15 |
A" |
3071 |
2931 |
39.17 |
100.38 |
0.75 |
0.86 |
| 16 |
A" |
1268 |
1210 |
0.01 |
11.04 |
0.75 |
0.86 |
| 17 |
A" |
1052 |
1004 |
10.69 |
0.66 |
0.75 |
0.86 |
| 18 |
A" |
1031 |
984 |
10.26 |
0.00 |
0.75 |
0.86 |
| 19 |
A" |
954 |
910 |
41.75 |
0.42 |
0.75 |
0.86 |
| 20 |
A" |
564 |
539 |
9.03 |
11.66 |
0.75 |
0.86 |
| 21 |
A" |
184 |
176 |
2.84 |
5.63 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16137.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 15400.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.942 |
-0.203 |
0.000 |
| C2 |
0.000 |
0.954 |
0.000 |
| C3 |
1.323 |
0.832 |
0.000 |
| F4 |
-0.266 |
-1.405 |
0.000 |
| H5 |
1.971 |
1.702 |
0.000 |
| H6 |
1.789 |
-0.147 |
0.000 |
| H7 |
-0.473 |
1.935 |
0.000 |
| H8 |
-1.592 |
-0.171 |
0.886 |
| H9 |
-1.592 |
-0.171 |
-0.886 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4917 | 2.4899 | 1.3787 | 3.4806 | 2.7307 | 2.1893 | 1.0995 | 1.0995 |
C2 | 1.4917 | | 1.3283 | 2.3737 | 2.1084 | 2.1003 | 1.0895 | 2.1414 | 2.1414 | C3 | 2.4899 | 1.3283 | | 2.7439 | 1.0848 | 1.0849 | 2.1073 | 3.2077 | 3.2077 | F4 | 1.3787 | 2.3737 | 2.7439 | | 3.8285 | 2.4086 | 3.3467 | 2.0165 | 2.0165 | H5 | 3.4806 | 2.1084 | 1.0848 | 3.8285 | | 1.8586 | 2.4551 | 4.1221 | 4.1221 | H6 | 2.7307 | 2.1003 | 1.0849 | 2.4086 | 1.8586 | | 3.0743 | 3.4949 | 3.4949 | H7 | 2.1893 | 1.0895 | 2.1073 | 3.3467 | 2.4551 | 3.0743 | | 2.5446 | 2.5446 | H8 | 1.0995 | 2.1414 | 3.2077 | 2.0165 | 4.1221 | 3.4949 | 2.5446 | | 1.7715 | H9 | 1.0995 | 2.1414 | 3.2077 | 2.0165 | 4.1221 | 3.4949 | 2.5446 | 1.7715 | |
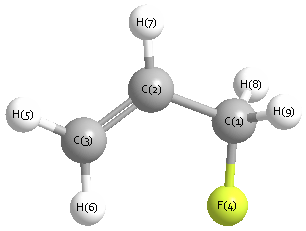 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.897 |
|
C1 |
C2 |
H7 |
115.137 |
| C2 |
C1 |
F4 |
111.506 |
|
C2 |
C1 |
H8 |
110.547 |
| C2 |
C1 |
H9 |
110.547 |
|
C2 |
C3 |
H5 |
121.460 |
| C2 |
C3 |
H6 |
120.667 |
|
C3 |
C2 |
H7 |
120.967 |
| F4 |
C1 |
H8 |
108.386 |
|
F4 |
C1 |
H9 |
108.386 |
| H5 |
C3 |
H6 |
117.873 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.076 |
|
|
|
| 2 |
C |
-0.134 |
|
|
|
| 3 |
C |
-0.277 |
|
|
|
| 4 |
F |
-0.314 |
|
|
|
| 5 |
H |
0.128 |
|
|
|
| 6 |
H |
0.143 |
|
|
|
| 7 |
H |
0.125 |
|
|
|
| 8 |
H |
0.127 |
|
|
|
| 9 |
H |
0.127 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.843 |
1.348 |
0.000 |
1.590 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.793 |
-0.355 |
0.000 |
| y |
-0.355 |
-24.508 |
0.000 |
| z |
0.000 |
0.000 |
-24.693 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.808 |
-0.355 |
0.000 |
| y |
-0.355 |
-1.765 |
0.000 |
| z |
0.000 |
0.000 |
-2.042 |
|
| Polar |
|---|
| 3z2-r2 | -4.085 |
|---|
| x2-y2 | 3.715 |
|---|
| xy | -0.355 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.626 |
0.262 |
0.000 |
| y |
0.262 |
4.601 |
0.000 |
| z |
0.000 |
0.000 |
2.939 |
<r2> (average value of r
2) Å
2
| <r2> |
79.325 |
| (<r2>)1/2 |
8.906 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -216.911819 |
| Energy at 298.15K | |
| HF Energy | -216.911819 |
| Nuclear repulsion energy | 114.282376 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3267 |
3117 |
11.59 |
62.49 |
0.60 |
0.75 |
| 2 |
A |
3193 |
3047 |
6.06 |
115.28 |
0.24 |
0.39 |
| 3 |
A |
3172 |
3027 |
8.98 |
58.32 |
0.11 |
0.20 |
| 4 |
A |
3100 |
2958 |
33.22 |
66.55 |
0.71 |
0.83 |
| 5 |
A |
3043 |
2904 |
40.08 |
124.08 |
0.12 |
0.21 |
| 6 |
A |
1753 |
1673 |
0.47 |
17.34 |
0.15 |
0.26 |
| 7 |
A |
1515 |
1446 |
0.97 |
10.35 |
0.67 |
0.80 |
| 8 |
A |
1471 |
1404 |
22.43 |
11.57 |
0.55 |
0.71 |
| 9 |
A |
1404 |
1340 |
17.24 |
6.03 |
0.65 |
0.79 |
| 10 |
A |
1317 |
1257 |
0.10 |
14.39 |
0.47 |
0.64 |
| 11 |
A |
1272 |
1214 |
3.50 |
14.51 |
0.73 |
0.84 |
| 12 |
A |
1189 |
1135 |
1.84 |
1.93 |
0.70 |
0.82 |
| 13 |
A |
1100 |
1050 |
106.96 |
5.54 |
0.53 |
0.70 |
| 14 |
A |
1040 |
993 |
23.06 |
0.94 |
0.72 |
0.83 |
| 15 |
A |
999 |
953 |
2.11 |
1.33 |
0.20 |
0.33 |
| 16 |
A |
965 |
921 |
45.02 |
0.48 |
0.42 |
0.60 |
| 17 |
A |
934 |
891 |
2.51 |
2.43 |
0.21 |
0.34 |
| 18 |
A |
654 |
624 |
6.92 |
6.94 |
0.69 |
0.82 |
| 19 |
A |
441 |
421 |
2.41 |
4.72 |
0.52 |
0.69 |
| 20 |
A |
322 |
307 |
5.95 |
3.95 |
0.75 |
0.86 |
| 21 |
A |
116 |
111 |
0.97 |
6.53 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16133.2 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 15395.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.605 |
0.441 |
0.289 |
| C2 |
0.639 |
-0.378 |
0.228 |
| C3 |
1.810 |
0.084 |
-0.200 |
| F4 |
-1.654 |
-0.232 |
-0.317 |
| H5 |
2.705 |
-0.531 |
-0.191 |
| H6 |
1.919 |
1.100 |
-0.571 |
| H7 |
0.546 |
-1.400 |
0.591 |
| H8 |
-0.893 |
0.631 |
1.332 |
| H9 |
-0.465 |
1.407 |
-0.211 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4908 | 2.4901 | 1.3857 | 3.4830 | 2.7468 | 2.1925 | 1.0988 | 1.0967 |
C2 | 1.4908 | | 1.3294 | 2.3617 | 2.1132 | 2.1120 | 1.0889 | 2.1413 | 2.1440 | C3 | 2.4901 | 1.3294 | | 3.4804 | 1.0854 | 1.0876 | 2.1035 | 3.1552 | 2.6319 | F4 | 1.3857 | 2.3617 | 3.4804 | | 4.3706 | 3.8214 | 2.6515 | 2.0111 | 2.0274 | H5 | 3.4830 | 2.1132 | 1.0854 | 4.3706 | | 1.8500 | 2.4549 | 4.0765 | 3.7151 | H6 | 2.7468 | 2.1120 | 1.0876 | 3.8214 | 1.8500 | | 3.0800 | 3.4279 | 2.4301 | H7 | 2.1925 | 1.0889 | 2.1035 | 2.6515 | 2.4549 | 3.0800 | | 2.5978 | 3.0896 | H8 | 1.0988 | 2.1413 | 3.1552 | 2.0111 | 4.0765 | 3.4279 | 2.5978 | | 1.7797 | H9 | 1.0967 | 2.1440 | 2.6319 | 2.0274 | 3.7151 | 2.4301 | 3.0896 | 1.7797 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.899 |
|
C1 |
C2 |
H7 |
115.522 |
| C2 |
C1 |
F4 |
110.315 |
|
C2 |
C1 |
H8 |
110.645 |
| C2 |
C1 |
H9 |
110.994 |
|
C2 |
C3 |
H5 |
121.794 |
| C2 |
C3 |
H6 |
121.488 |
|
C3 |
C2 |
H7 |
120.556 |
| F4 |
C1 |
H8 |
107.522 |
|
F4 |
C1 |
H9 |
108.957 |
| H5 |
C3 |
H6 |
116.717 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.065 |
|
|
|
| 2 |
C |
-0.099 |
|
|
|
| 3 |
C |
-0.285 |
|
|
|
| 4 |
F |
-0.321 |
|
|
|
| 5 |
H |
0.136 |
|
|
|
| 6 |
H |
0.128 |
|
|
|
| 7 |
H |
0.136 |
|
|
|
| 8 |
H |
0.124 |
|
|
|
| 9 |
H |
0.118 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.372 |
0.687 |
0.764 |
1.714 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.949 |
-0.981 |
-1.538 |
| y |
-0.981 |
-21.987 |
-0.866 |
| z |
-1.538 |
-0.866 |
-24.284 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.814 |
-0.981 |
-1.538 |
| y |
-0.981 |
2.629 |
-0.866 |
| z |
-1.538 |
-0.866 |
-0.816 |
|
| Polar |
|---|
| 3z2-r2 | -1.632 |
|---|
| x2-y2 | -2.962 |
|---|
| xy | -0.981 |
|---|
| xz | -1.538 |
|---|
| yz | -0.866 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.366 |
0.174 |
-0.863 |
| y |
0.174 |
4.500 |
-0.546 |
| z |
-0.863 |
-0.546 |
3.322 |
<r2> (average value of r
2) Å
2
| <r2> |
88.936 |
| (<r2>)1/2 |
9.431 |
