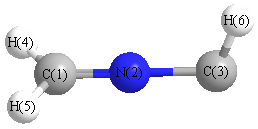
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3183 |
3037 |
1.90 |
|
|
|
| 2 |
A' |
3121 |
2979 |
13.54 |
|
|
|
| 3 |
A' |
2057 |
1963 |
202.92 |
|
|
|
| 4 |
A' |
1513 |
1443 |
1.69 |
|
|
|
| 5 |
A' |
1256 |
1199 |
9.74 |
|
|
|
| 6 |
A' |
922 |
880 |
333.37 |
|
|
|
| 7 |
A' |
728 |
695 |
81.95 |
|
|
|
| 8 |
A' |
501 |
479 |
25.26 |
|
|
|
| 9 |
A" |
3292 |
3141 |
2.64 |
|
|
|
| 10 |
A" |
1146 |
1093 |
5.63 |
|
|
|
| 11 |
A" |
972 |
928 |
8.52 |
|
|
|
| 12 |
A" |
359 |
343 |
3.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9525.3 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 9090.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.202 |
|
|
|
| 2 |
N |
-0.140 |
|
|
|
| 3 |
C |
-0.112 |
|
|
|
| 4 |
H |
0.154 |
|
|
|
| 5 |
H |
0.154 |
|
|
|
| 6 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.399 |
-0.916 |
0.000 |
1.672 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.033 |
2.266 |
0.000 |
| y |
2.266 |
-16.108 |
0.000 |
| z |
0.000 |
0.000 |
-16.821 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.568 |
2.266 |
0.000 |
| y |
2.266 |
2.318 |
0.000 |
| z |
0.000 |
0.000 |
1.250 |
|
| Polar |
|---|
| 3z2-r2 | 2.499 |
|---|
| x2-y2 | -3.924 |
|---|
| xy | 2.266 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.358 |
-0.795 |
0.000 |
| y |
-0.795 |
8.619 |
0.000 |
| z |
0.000 |
0.000 |
2.622 |
<r2> (average value of r
2) Å
2
| <r2> |
41.742 |
| (<r2>)1/2 |
6.461 |