return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O (allenol)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-191.329218
Energy at 298.15K-191.332639
HF Energy-190.718809
Nuclear repulsion energy101.648375
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3890 3662 30.71      
2 A' 3279 3087 6.66      
3 A' 3192 3005 16.17      
4 A' 2085 1963 34.80      
5 A' 1555 1464 35.16      
6 A' 1460 1375 67.27      
7 A' 1320 1242 1.60      
8 A' 1222 1150 117.26      
9 A' 1005 946 110.98      
10 A' 931 876 29.39      
11 A' 618 582 19.50      
12 A' 206 194 0.74      
13 A" 3274 3082 10.61      
14 A" 1067 1004 1.70      
15 A" 906 853 20.86      
16 A" 628 591 0.59      
17 A" 401 377 130.93      
18 A" 256 241 3.04      

Unscaled Zero Point Vibrational Energy (zpe) 13647.6 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 12847.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
1.44675 0.14408 0.13462

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.666 -0.490 0.000
C2 0.000 0.646 0.000
C3 -0.655 1.788 0.000
O4 0.114 -1.755 0.000
H5 1.746 -0.528 0.000
H6 -0.937 2.282 0.923
H7 -0.937 2.282 -0.923
H8 -0.845 -1.661 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8
C11.31662.63341.38031.08103.33263.33261.9117
C21.31661.31682.40392.10392.09952.09952.4569
C32.63341.31683.62593.33631.08411.08413.4542
O41.38032.40393.62592.04194.27314.27310.9641
H51.08102.10393.33632.04193.99363.99362.8283
H63.33262.09951.08414.27313.99361.84564.0508
H73.33262.09951.08414.27313.99361.84564.0508
H81.91172.45693.45420.96412.82834.05084.0508

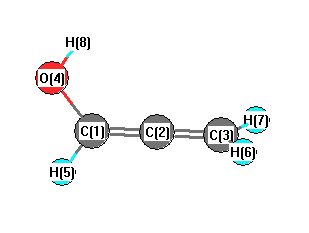
picture of allenol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 179.481 C1 O4 H8 107.938
C2 C1 O4 126.074 C2 C1 H5 122.385
C2 C3 H6 121.658 C2 C3 H7 121.658
O4 C1 H5 111.541 H6 C3 H7 116.684
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at QCISD/6-31G** Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.141      
2 C -0.008      
3 C -0.334      
4 O -0.618      
5 H 0.166      
6 H 0.149      
7 H 0.149      
8 H 0.354      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å
Primitive
 xyz
x -20.983 0.981 -0.002
y 0.981 -24.949 -0.003
z -0.002 -0.003 -24.338
Traceless
 xyz
x 3.660 0.981 -0.002
y 0.981 -2.289 -0.003
z -0.002 -0.003 -1.372
Polar
3z2-r2-2.743
x2-y23.966
xy0.981
xz-0.002
yz-0.003


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 88.416
(<r2>)1/2 9.403