return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O2 (propenalol)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-266.418512
Energy at 298.15K-266.423605
HF Energy-265.638995
Nuclear repulsion energy164.792325
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3326 3108 1.10      
2 A' 3319 3101 97.07      
3 A' 3263 3049 70.34      
4 A' 3091 2888 107.99      
5 A' 1740 1626 182.15      
6 A' 1686 1575 254.06      
7 A' 1514 1415 45.56      
8 A' 1444 1349 54.96      
9 A' 1442 1347 11.21      
10 A' 1312 1226 148.47      
11 A' 1138 1064 16.32      
12 A' 1027 960 59.65      
13 A' 898 840 7.88      
14 A' 517 483 19.28      
15 A' 277 259 4.97      
16 A" 1054 985 13.54      
17 A" 1004 938 39.36      
18 A" 904 844 67.44      
19 A" 785 733 37.06      
20 A" 377 352 5.17      
21 A" 279 261 4.78      

Unscaled Zero Point Vibrational Energy (zpe) 15198.0 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 14201.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.32662 0.17160 0.11250

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.103 0.000
C2 1.236 0.367 0.000
C3 -1.183 0.428 0.000
O4 1.295 -0.880 0.000
O5 -1.294 -0.895 0.000
H6 -0.361 -1.238 0.000
H7 0.005 2.180 0.000
H8 2.171 0.941 0.000
H9 -2.141 0.933 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9
C11.43851.36192.36862.38022.36871.07722.17712.1472
C21.43852.41971.24842.82692.26422.19151.09773.4236
C31.36192.41972.80251.32781.85802.11673.39321.0826
O42.36861.24842.80252.58921.69473.32112.02143.8851
O52.38022.82691.32782.58920.99393.33813.92162.0149
H62.36872.26421.85801.69470.99393.43783.34122.8074
H71.07722.19152.11673.32113.33813.43782.49502.4815
H82.17711.09773.39322.02143.92163.34122.49504.3116
H92.14723.42361.08263.88512.01492.80742.48154.3116

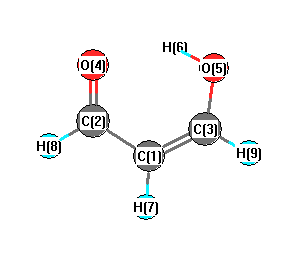
picture of propenalol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O4 123.507 C1 C2 H8 117.645
C1 C3 O5 124.485 C1 C3 H9 122.489
C2 C1 C3 119.525 C2 C1 H7 120.508
C3 C1 H7 119.967 C3 O5 H6 105.413
O4 C2 H8 118.848 O5 C3 H9 113.025
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-31G** Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.404      
2 C 0.414      
3 C 0.341      
4 O -0.636      
5 O -0.621      
6 H 0.434      
7 H 0.161      
8 H 0.135      
9 H 0.176      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 101.805
(<r2>)1/2 10.090