return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H4O2 (propenalol)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-266.427000
Energy at 298.15K-266.432039
HF Energy-265.640149
Nuclear repulsion energy163.997033
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3520 3314 116.48      
2 A' 3300 3106 4.71      
3 A' 3259 3068 17.11      
4 A' 3070 2890 109.48      
5 A' 1759 1656 171.45      
6 A' 1683 1584 294.12      
7 A' 1509 1420 47.53      
8 A' 1449 1364 69.47      
9 A' 1444 1359 10.47      
10 A' 1304 1228 145.62      
11 A' 1135 1069 22.77      
12 A' 1005 946 75.38      
13 A' 899 846 10.04      
14 A' 507 478 20.35      
15 A' 274 258 4.31      
16 A" 1054 992 9.10      
17 A" 992 934 11.32      
18 A" 863 813 101.90      
19 A" 777 731 45.40      
20 A" 377 355 5.38      
21 A" 261 245 4.53      

Unscaled Zero Point Vibrational Energy (zpe) 15219.3 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 14327.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.32707 0.16724 0.11066

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
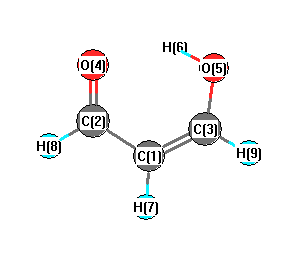
C1 0.000 1.099 0.000
C2 1.250 0.362 0.000
C3 -1.185 0.433 0.000
O4 1.317 -0.878 0.000
O5 -1.321 -0.894 0.000
H6 -0.411 -1.267 0.000
H7 0.010 2.178 0.000
H8 2.180 0.949 0.000
H9 -2.138 0.950 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 O5 H6 H7 H8 H9
C11.45071.35922.37522.39132.40111.07922.18472.1429
C21.45072.43561.24142.86092.32572.19861.09963.4378
C31.35922.43562.82481.33481.86822.11493.40381.0835
O42.37521.24142.82482.63791.77113.32342.01963.9082
O52.39132.86091.33482.63790.98323.34853.95592.0169
H62.40112.32571.86821.77110.98323.47053.40842.8097
H71.07922.19862.11493.32343.34853.47052.49342.4745
H82.18471.09963.40382.01963.95593.40842.49344.3172
H92.14293.43781.08353.90822.01692.80972.47454.3172

picture of propenalol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O4 123.654 C1 C2 H8 117.222
C1 C3 O5 125.154 C1 C3 H9 122.228
C2 C1 C3 120.146 C2 C1 H7 119.984
C3 C1 H7 119.869 C3 O5 H6 106.415
O4 C2 H8 119.124 O5 C3 H9 112.618
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at QCISD/6-31G** Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C -0.399      
2 C 0.412      
3 C 0.335      
4 O -0.620      
5 O -0.619      
6 H 0.423      
7 H 0.161      
8 H 0.132      
9 H 0.175      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 103.144
(<r2>)1/2 10.156