Jump to
S1C2
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
Geometric Data calculated at HSEh1PBE/6-31G**
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -115.823032 |
| Energy at 298.15K | -115.824659 |
| HF Energy | -115.823032 |
| Nuclear repulsion energy | 58.038969 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3329 |
3177 |
0.09 |
|
|
|
| 2 |
A' |
3099 |
2958 |
60.20 |
|
|
|
| 3 |
A' |
1705 |
1627 |
0.25 |
|
|
|
| 4 |
A' |
1280 |
1222 |
10.65 |
|
|
|
| 5 |
A' |
1013 |
967 |
17.96 |
|
|
|
| 6 |
A' |
932 |
890 |
5.63 |
|
|
|
| 7 |
A' |
618 |
589 |
62.69 |
|
|
|
| 8 |
A" |
3282 |
3132 |
1.66 |
|
|
|
| 9 |
A" |
1051 |
1003 |
23.39 |
|
|
|
| 10 |
A" |
998 |
953 |
3.84 |
|
|
|
| 11 |
A" |
883 |
843 |
0.24 |
|
|
|
| 12 |
A" |
733 |
700 |
33.26 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9461.8 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 9029.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.037 |
0.886 |
0.000 |
| H2 |
0.744 |
1.651 |
0.000 |
| C3 |
-0.037 |
-0.417 |
0.656 |
| C4 |
-0.037 |
-0.417 |
-0.656 |
| H5 |
-0.040 |
-0.981 |
1.577 |
| H6 |
-0.040 |
-0.981 |
-1.577 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
| C1 | | 1.0933 | 1.4586 | 1.4586 | 2.4432 | 2.4432 |
H2 | 1.0933 | | 2.3057 | 2.3057 | 3.1664 | 3.1664 | C3 | 1.4586 | 2.3057 | | 1.3123 | 1.0794 | 2.3028 | C4 | 1.4586 | 2.3057 | 1.3123 | | 2.3028 | 1.0794 | H5 | 2.4432 | 3.1664 | 1.0794 | 2.3028 | | 3.1532 | H6 | 2.4432 | 3.1664 | 2.3028 | 1.0794 | 3.1532 | |
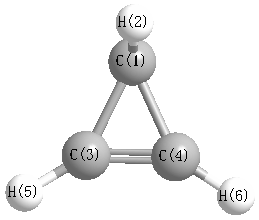 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C4 |
63.266 |
|
C1 |
C3 |
H5 |
148.218 |
| C1 |
C4 |
C3 |
63.266 |
|
C1 |
C4 |
H6 |
148.218 |
| H2 |
C1 |
C3 |
128.687 |
|
H2 |
C1 |
C4 |
128.687 |
| C3 |
C1 |
C4 |
53.468 |
|
C3 |
C4 |
H6 |
148.515 |
| C4 |
C3 |
H5 |
148.515 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.196 |
|
|
|
| 2 |
H |
0.114 |
|
|
|
| 3 |
C |
-0.100 |
|
|
|
| 4 |
C |
-0.100 |
|
|
|
| 5 |
H |
0.141 |
|
|
|
| 6 |
H |
0.141 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.627 |
-1.295 |
0.000 |
1.439 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.709 |
1.176 |
0.000 |
| y |
1.176 |
-17.890 |
0.000 |
| z |
0.000 |
0.000 |
-15.343 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.092 |
1.176 |
0.000 |
| y |
1.176 |
-0.864 |
0.000 |
| z |
0.000 |
0.000 |
2.957 |
|
| Polar |
|---|
| 3z2-r2 | 5.913 |
|---|
| x2-y2 | -0.819 |
|---|
| xy | 1.176 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.223 |
0.351 |
0.000 |
| y |
0.351 |
4.592 |
0.000 |
| z |
0.000 |
0.000 |
4.748 |
<r2> (average value of r
2) Å
2
| <r2> |
32.975 |
| (<r2>)1/2 |
5.742 |
