Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.401948 |
| Energy at 298.15K | -169.405931 |
| HF Energy | -168.906437 |
| Nuclear repulsion energy | 70.608436 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3910 |
3654 |
71.41 |
|
|
|
| 2 |
A' |
3606 |
3369 |
3.82 |
|
|
|
| 3 |
A' |
3140 |
2934 |
57.26 |
|
|
|
| 4 |
A' |
1779 |
1663 |
153.41 |
|
|
|
| 5 |
A' |
1463 |
1367 |
9.09 |
|
|
|
| 6 |
A' |
1354 |
1265 |
164.17 |
|
|
|
| 7 |
A' |
1214 |
1134 |
49.37 |
|
|
|
| 8 |
A' |
1085 |
1014 |
205.55 |
|
|
|
| 9 |
A' |
617 |
577 |
1.28 |
|
|
|
| 10 |
A" |
1051 |
982 |
7.11 |
|
|
|
| 11 |
A" |
850 |
794 |
61.01 |
|
|
|
| 12 |
A" |
380 |
355 |
76.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10224.1 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 9553.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.381 |
0.000 |
| O2 |
-1.007 |
-0.530 |
0.000 |
| N3 |
1.203 |
-0.026 |
0.000 |
| H4 |
-0.332 |
1.421 |
0.000 |
| H5 |
-1.848 |
-0.061 |
0.000 |
| H6 |
1.820 |
0.781 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3579 | 1.2697 | 1.0922 | 1.9004 | 1.8639 |
O2 | 1.3579 | | 2.2666 | 2.0647 | 0.9633 | 3.1167 | N3 | 1.2697 | 2.2666 | | 2.1097 | 3.0514 | 1.0164 | H4 | 1.0922 | 2.0647 | 2.1097 | | 2.1202 | 2.2457 | H5 | 1.9004 | 0.9633 | 3.0514 | 2.1202 | | 3.7641 | H6 | 1.8639 | 3.1167 | 1.0164 | 2.2457 | 3.7641 | |
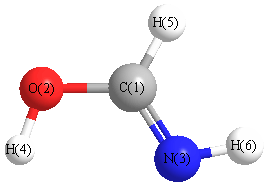 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.722 |
|
C1 |
N3 |
H6 |
108.730 |
| O2 |
C1 |
N3 |
119.185 |
|
O2 |
C1 |
H4 |
114.415 |
| N3 |
C1 |
H4 |
126.400 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.412712 |
| Energy at 298.15K | -169.416884 |
| HF Energy | -168.917771 |
| Nuclear repulsion energy | 70.959563 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3809 |
3559 |
54.26 |
|
|
|
| 2 |
A' |
3616 |
3379 |
4.56 |
|
|
|
| 3 |
A' |
3223 |
3011 |
31.28 |
|
|
|
| 4 |
A' |
1755 |
1640 |
214.40 |
|
|
|
| 5 |
A' |
1433 |
1339 |
20.07 |
|
|
|
| 6 |
A' |
1400 |
1308 |
7.54 |
|
|
|
| 7 |
A' |
1209 |
1130 |
109.23 |
|
|
|
| 8 |
A' |
1094 |
1022 |
172.53 |
|
|
|
| 9 |
A' |
590 |
551 |
52.97 |
|
|
|
| 10 |
A" |
1066 |
996 |
0.09 |
|
|
|
| 11 |
A" |
851 |
795 |
30.24 |
|
|
|
| 12 |
A" |
646 |
603 |
201.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10346.3 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 9667.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.422 |
0.000 |
| O2 |
-1.117 |
-0.335 |
0.000 |
| N3 |
1.162 |
-0.100 |
0.000 |
| H4 |
-0.261 |
1.477 |
0.000 |
| H5 |
-0.806 |
-1.255 |
0.000 |
| H6 |
1.870 |
0.628 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3491 | 1.2732 | 1.0868 | 1.8599 | 1.8809 |
O2 | 1.3491 | | 2.2905 | 2.0041 | 0.9708 | 3.1379 | N3 | 1.2732 | 2.2905 | | 2.1231 | 2.2811 | 1.0156 | H4 | 1.0868 | 2.0041 | 2.1231 | | 2.7853 | 2.2928 | H5 | 1.8599 | 0.9708 | 2.2811 | 2.7853 | | 3.2714 | H6 | 1.8809 | 3.1379 | 1.0156 | 2.2928 | 3.2714 | |
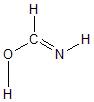 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
105.427 |
|
C1 |
N3 |
H6 |
110.022 |
| O2 |
C1 |
N3 |
121.698 |
|
O2 |
C1 |
H4 |
110.254 |
| N3 |
C1 |
H4 |
128.049 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.407324 |
| Energy at 298.15K | -169.411424 |
| HF Energy | -168.911648 |
| Nuclear repulsion energy | 70.485308 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3842 |
3590 |
32.29 |
|
|
|
| 2 |
A' |
3533 |
3301 |
5.33 |
|
|
|
| 3 |
A' |
3284 |
3069 |
13.22 |
|
|
|
| 4 |
A' |
1747 |
1632 |
210.82 |
|
|
|
| 5 |
A' |
1446 |
1351 |
1.04 |
|
|
|
| 6 |
A' |
1377 |
1286 |
23.57 |
|
|
|
| 7 |
A' |
1135 |
1060 |
274.42 |
|
|
|
| 8 |
A' |
1103 |
1031 |
37.38 |
|
|
|
| 9 |
A' |
593 |
554 |
36.50 |
|
|
|
| 10 |
A" |
1105 |
1032 |
69.66 |
|
|
|
| 11 |
A" |
852 |
796 |
55.51 |
|
|
|
| 12 |
A" |
528 |
494 |
90.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10271.9 cm
-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 9598.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.439 |
0.000 |
| O2 |
-1.108 |
-0.352 |
0.000 |
| N3 |
1.233 |
0.125 |
0.000 |
| H4 |
-0.298 |
1.479 |
0.000 |
| H5 |
-0.829 |
-1.279 |
0.000 |
| H6 |
1.364 |
-0.888 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3617 | 1.2721 | 1.0825 | 1.9077 | 1.9029 |
O2 | 1.3617 | | 2.3893 | 2.0028 | 0.9678 | 2.5297 | N3 | 1.2721 | 2.3893 | | 2.0442 | 2.4949 | 1.0215 | H4 | 1.0825 | 2.0028 | 2.0442 | | 2.8092 | 2.8928 | H5 | 1.9077 | 0.9678 | 2.4949 | 2.8092 | | 2.2281 | H6 | 1.9029 | 2.5297 | 1.0215 | 2.8928 | 2.2281 | |
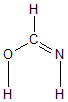 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.774 |
|
C1 |
N3 |
H6 |
111.659 |
| O2 |
C1 |
N3 |
130.199 |
|
O2 |
C1 |
H4 |
109.521 |
| N3 |
C1 |
H4 |
120.280 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
