Jump to
S1C2
S1C3
Energy calculated at QCISD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.411423 |
| Energy at 298.15K | -169.415388 |
| HF Energy | -168.906557 |
| Nuclear repulsion energy | 70.560115 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3923 |
3693 |
58.93 |
|
|
|
| 2 |
A' |
3590 |
3379 |
1.07 |
|
|
|
| 3 |
A' |
3129 |
2945 |
56.57 |
|
|
|
| 4 |
A' |
1804 |
1698 |
159.26 |
|
|
|
| 5 |
A' |
1476 |
1390 |
14.72 |
|
|
|
| 6 |
A' |
1367 |
1287 |
157.07 |
|
|
|
| 7 |
A' |
1226 |
1154 |
64.35 |
|
|
|
| 8 |
A' |
1089 |
1025 |
193.46 |
|
|
|
| 9 |
A' |
617 |
581 |
0.97 |
|
|
|
| 10 |
A" |
1059 |
997 |
8.79 |
|
|
|
| 11 |
A" |
848 |
798 |
55.19 |
|
|
|
| 12 |
A" |
359 |
338 |
78.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10242.9 cm
-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 9642.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.380 |
0.000 |
| O2 |
-1.007 |
-0.535 |
0.000 |
| N3 |
1.203 |
-0.020 |
0.000 |
| H4 |
-0.334 |
1.421 |
0.000 |
| H5 |
-1.850 |
-0.071 |
0.000 |
| H6 |
1.821 |
0.789 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3610 | 1.2679 | 1.0931 | 1.9045 | 1.8661 |
O2 | 1.3610 | | 2.2702 | 2.0685 | 0.9620 | 3.1227 | N3 | 1.2679 | 2.2702 | | 2.1067 | 3.0541 | 1.0171 | H4 | 1.0931 | 2.0685 | 2.1067 | | 2.1272 | 2.2456 | H5 | 1.9045 | 0.9620 | 3.0541 | 2.1272 | | 3.7704 | H6 | 1.8661 | 3.1227 | 1.0171 | 2.2456 | 3.7704 | |
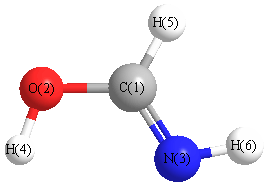 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.926 |
|
C1 |
N3 |
H6 |
109.011 |
| O2 |
C1 |
N3 |
119.391 |
|
O2 |
C1 |
H4 |
114.453 |
| N3 |
C1 |
H4 |
126.156 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at QCISD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.421774 |
| Energy at 298.15K | -169.425940 |
| HF Energy | -168.917941 |
| Nuclear repulsion energy | 70.920972 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3839 |
3614 |
45.70 |
|
|
|
| 2 |
A' |
3600 |
3389 |
1.40 |
|
|
|
| 3 |
A' |
3208 |
3020 |
32.19 |
|
|
|
| 4 |
A' |
1777 |
1672 |
226.16 |
|
|
|
| 5 |
A' |
1447 |
1362 |
28.11 |
|
|
|
| 6 |
A' |
1409 |
1327 |
4.40 |
|
|
|
| 7 |
A' |
1223 |
1152 |
93.92 |
|
|
|
| 8 |
A' |
1100 |
1035 |
182.07 |
|
|
|
| 9 |
A' |
592 |
558 |
51.61 |
|
|
|
| 10 |
A" |
1070 |
1007 |
0.11 |
|
|
|
| 11 |
A" |
851 |
802 |
30.50 |
|
|
|
| 12 |
A" |
626 |
589 |
199.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10370.4 cm
-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 9762.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.421 |
0.000 |
| O2 |
-1.117 |
-0.341 |
0.000 |
| N3 |
1.162 |
-0.094 |
0.000 |
| H4 |
-0.261 |
1.478 |
0.000 |
| H5 |
-0.810 |
-1.260 |
0.000 |
| H6 |
1.869 |
0.636 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3519 | 1.2713 | 1.0879 | 1.8660 | 1.8816 |
O2 | 1.3519 | | 2.2922 | 2.0095 | 0.9686 | 3.1417 | N3 | 1.2713 | 2.2922 | | 2.1201 | 2.2912 | 1.0162 | H4 | 1.0879 | 2.0095 | 2.1201 | | 2.7917 | 2.2904 | H5 | 1.8660 | 0.9686 | 2.2912 | 2.7917 | | 3.2823 | H6 | 1.8816 | 3.1417 | 1.0162 | 2.2904 | 3.2823 | |
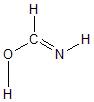 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
105.869 |
|
C1 |
N3 |
H6 |
110.177 |
| O2 |
C1 |
N3 |
121.778 |
|
O2 |
C1 |
H4 |
110.433 |
| N3 |
C1 |
H4 |
127.790 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at QCISD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.416533 |
| Energy at 298.15K | -169.420622 |
| HF Energy | -168.911822 |
| Nuclear repulsion energy | 70.491196 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3867 |
3640 |
26.60 |
|
|
|
| 2 |
A' |
3516 |
3310 |
6.96 |
|
|
|
| 3 |
A' |
3267 |
3075 |
14.51 |
|
|
|
| 4 |
A' |
1769 |
1666 |
224.28 |
|
|
|
| 5 |
A' |
1447 |
1362 |
2.35 |
|
|
|
| 6 |
A' |
1393 |
1311 |
23.48 |
|
|
|
| 7 |
A' |
1153 |
1085 |
241.26 |
|
|
|
| 8 |
A' |
1111 |
1046 |
63.40 |
|
|
|
| 9 |
A' |
596 |
561 |
37.64 |
|
|
|
| 10 |
A" |
1102 |
1038 |
66.90 |
|
|
|
| 11 |
A" |
856 |
806 |
55.68 |
|
|
|
| 12 |
A" |
505 |
475 |
93.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10290.2 cm
-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 9687.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.442 |
0.000 |
| O2 |
-1.106 |
-0.356 |
0.000 |
| N3 |
1.231 |
0.126 |
0.000 |
| H4 |
-0.299 |
1.484 |
0.000 |
| H5 |
-0.828 |
-1.281 |
0.000 |
| H6 |
1.361 |
-0.888 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3639 | 1.2706 | 1.0838 | 1.9114 | 1.9028 |
O2 | 1.3639 | | 2.3860 | 2.0088 | 0.9659 | 2.5236 | N3 | 1.2706 | 2.3860 | | 2.0455 | 2.4932 | 1.0223 | H4 | 1.0838 | 2.0088 | 2.0455 | | 2.8146 | 2.8949 | H5 | 1.9114 | 0.9659 | 2.4932 | 2.8146 | | 2.2234 | H6 | 1.9028 | 2.5236 | 1.0223 | 2.8949 | 2.2234 | |
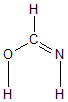 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.055 |
|
C1 |
N3 |
H6 |
111.712 |
| O2 |
C1 |
N3 |
129.789 |
|
O2 |
C1 |
H4 |
109.779 |
| N3 |
C1 |
H4 |
120.431 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
