Jump to
S1C2
S1C3
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.665441 |
| Energy at 298.15K | -169.669447 |
| HF Energy | -169.665441 |
| Nuclear repulsion energy | 70.905112 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3844 |
3663 |
65.15 |
|
|
|
| 2 |
A' |
3560 |
3393 |
2.68 |
|
|
|
| 3 |
A' |
3063 |
2920 |
72.62 |
|
|
|
| 4 |
A' |
1817 |
1732 |
197.74 |
|
|
|
| 5 |
A' |
1462 |
1393 |
19.01 |
|
|
|
| 6 |
A' |
1363 |
1299 |
176.36 |
|
|
|
| 7 |
A' |
1224 |
1166 |
48.46 |
|
|
|
| 8 |
A' |
1090 |
1039 |
207.49 |
|
|
|
| 9 |
A' |
624 |
595 |
1.45 |
|
|
|
| 10 |
A" |
1054 |
1005 |
6.99 |
|
|
|
| 11 |
A" |
841 |
802 |
67.56 |
|
|
|
| 12 |
A" |
399 |
380 |
72.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10170.4 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 9693.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.373 |
0.000 |
| O2 |
-1.002 |
-0.529 |
0.000 |
| N3 |
1.197 |
-0.021 |
0.000 |
| H4 |
-0.339 |
1.420 |
0.000 |
| H5 |
-1.847 |
-0.062 |
0.000 |
| H6 |
1.822 |
0.783 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3482 | 1.2604 | 1.1002 | 1.8972 | 1.8674 |
O2 | 1.3482 | | 2.2576 | 2.0584 | 0.9650 | 3.1139 | N3 | 1.2604 | 2.2576 | | 2.1056 | 3.0445 | 1.0177 | H4 | 1.1002 | 2.0584 | 2.1056 | | 2.1141 | 2.2521 | H5 | 1.8972 | 0.9650 | 3.0445 | 2.1141 | | 3.7645 | H6 | 1.8674 | 3.1139 | 1.0177 | 2.2521 | 3.7645 | |
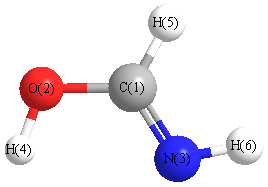 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.081 |
|
C1 |
N3 |
H6 |
109.652 |
| O2 |
C1 |
N3 |
119.838 |
|
O2 |
C1 |
H4 |
114.053 |
| N3 |
C1 |
H4 |
126.109 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.223 |
|
|
|
| 2 |
O |
-0.544 |
|
|
|
| 3 |
N |
-0.557 |
|
|
|
| 4 |
H |
0.140 |
|
|
|
| 5 |
H |
0.425 |
|
|
|
| 6 |
H |
0.312 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.111 |
3.240 |
0.000 |
3.867 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.524 |
1.251 |
0.000 |
| y |
1.251 |
-17.668 |
0.000 |
| z |
0.000 |
0.000 |
-18.506 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.563 |
1.251 |
0.000 |
| y |
1.251 |
-1.653 |
0.000 |
| z |
0.000 |
0.000 |
-2.911 |
|
| Polar |
|---|
| 3z2-r2 | -5.821 |
|---|
| x2-y2 | 4.144 |
|---|
| xy | 1.251 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.759 |
0.016 |
0.000 |
| y |
0.016 |
2.827 |
0.000 |
| z |
0.000 |
0.000 |
1.461 |
<r2> (average value of r
2) Å
2
| <r2> |
40.970 |
| (<r2>)1/2 |
6.401 |
Jump to
S1C1
S1C3
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.676229 |
| Energy at 298.15K | -169.680406 |
| HF Energy | -169.676229 |
| Nuclear repulsion energy | 71.263349 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3728 |
3553 |
44.45 |
|
|
|
| 2 |
A' |
3564 |
3397 |
2.89 |
|
|
|
| 3 |
A' |
3156 |
3008 |
38.83 |
|
|
|
| 4 |
A' |
1787 |
1703 |
265.89 |
|
|
|
| 5 |
A' |
1432 |
1365 |
31.31 |
|
|
|
| 6 |
A' |
1403 |
1337 |
1.54 |
|
|
|
| 7 |
A' |
1219 |
1162 |
121.23 |
|
|
|
| 8 |
A' |
1101 |
1049 |
156.77 |
|
|
|
| 9 |
A' |
593 |
565 |
53.42 |
|
|
|
| 10 |
A" |
1065 |
1015 |
0.00 |
|
|
|
| 11 |
A" |
844 |
804 |
26.91 |
|
|
|
| 12 |
A" |
655 |
624 |
207.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10273.3 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 9791.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.416 |
0.000 |
| O2 |
-1.111 |
-0.331 |
0.000 |
| N3 |
1.155 |
-0.099 |
0.000 |
| H4 |
-0.263 |
1.478 |
0.000 |
| H5 |
-0.808 |
-1.257 |
0.000 |
| H6 |
1.873 |
0.623 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3390 | 1.2646 | 1.0940 | 1.8577 | 1.8839 |
O2 | 1.3390 | | 2.2779 | 1.9983 | 0.9735 | 3.1323 | N3 | 1.2646 | 2.2779 | | 2.1207 | 2.2792 | 1.0174 | H4 | 1.0940 | 1.9983 | 2.1207 | | 2.7885 | 2.3006 | H5 | 1.8577 | 0.9735 | 2.2792 | 2.7885 | | 3.2737 | H6 | 1.8839 | 3.1323 | 1.0174 | 2.3006 | 3.2737 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
105.816 |
|
C1 |
N3 |
H6 |
110.820 |
| O2 |
C1 |
N3 |
122.035 |
|
O2 |
C1 |
H4 |
110.029 |
| N3 |
C1 |
H4 |
127.936 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.251 |
|
|
|
| 2 |
O |
-0.559 |
|
|
|
| 3 |
N |
-0.608 |
|
|
|
| 4 |
H |
0.171 |
|
|
|
| 5 |
H |
0.427 |
|
|
|
| 6 |
H |
0.318 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.076 |
0.977 |
0.000 |
0.980 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.164 |
3.350 |
0.000 |
| y |
3.350 |
-13.863 |
0.000 |
| z |
0.000 |
0.000 |
-18.504 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.981 |
3.350 |
0.000 |
| y |
3.350 |
4.971 |
0.000 |
| z |
0.000 |
0.000 |
-1.990 |
|
| Polar |
|---|
| 3z2-r2 | -3.980 |
|---|
| x2-y2 | -5.301 |
|---|
| xy | 3.350 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.252 |
0.091 |
0.000 |
| y |
0.091 |
3.212 |
0.000 |
| z |
0.000 |
0.000 |
1.454 |
<r2> (average value of r
2) Å
2
| <r2> |
40.306 |
| (<r2>)1/2 |
6.349 |
Jump to
S1C1
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.670482 |
| Energy at 298.15K | -169.674595 |
| HF Energy | -169.670482 |
| Nuclear repulsion energy | 70.814068 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3766 |
3589 |
24.53 |
|
|
|
| 2 |
A' |
3469 |
3307 |
6.58 |
|
|
|
| 3 |
A' |
3222 |
3071 |
14.64 |
|
|
|
| 4 |
A' |
1785 |
1701 |
270.09 |
|
|
|
| 5 |
A' |
1427 |
1360 |
0.83 |
|
|
|
| 6 |
A' |
1382 |
1317 |
24.48 |
|
|
|
| 7 |
A' |
1146 |
1092 |
268.76 |
|
|
|
| 8 |
A' |
1113 |
1061 |
40.94 |
|
|
|
| 9 |
A' |
597 |
569 |
37.13 |
|
|
|
| 10 |
A" |
1089 |
1038 |
68.32 |
|
|
|
| 11 |
A" |
858 |
817 |
62.64 |
|
|
|
| 12 |
A" |
541 |
515 |
86.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10196.9 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 9718.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.435 |
0.000 |
| O2 |
-1.101 |
-0.347 |
0.000 |
| N3 |
1.224 |
0.123 |
0.000 |
| H4 |
-0.298 |
1.483 |
0.000 |
| H5 |
-0.830 |
-1.279 |
0.000 |
| H6 |
1.369 |
-0.891 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3509 | 1.2635 | 1.0895 | 1.9039 | 1.9058 |
O2 | 1.3509 | | 2.3727 | 1.9985 | 0.9701 | 2.5294 | N3 | 1.2635 | 2.3727 | | 2.0417 | 2.4866 | 1.0240 | H4 | 1.0895 | 1.9985 | 2.0417 | | 2.8120 | 2.9009 | H5 | 1.9039 | 0.9701 | 2.4866 | 2.8120 | | 2.2325 | H6 | 1.9058 | 2.5294 | 1.0240 | 2.9009 | 2.2325 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.126 |
|
C1 |
N3 |
H6 |
112.421 |
| O2 |
C1 |
N3 |
130.306 |
|
O2 |
C1 |
H4 |
109.486 |
| N3 |
C1 |
H4 |
120.208 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.220 |
|
|
|
| 2 |
O |
-0.554 |
|
|
|
| 3 |
N |
-0.563 |
|
|
|
| 4 |
H |
0.196 |
|
|
|
| 5 |
H |
0.412 |
|
|
|
| 6 |
H |
0.289 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.007 |
-1.981 |
0.000 |
2.223 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.692 |
-1.882 |
0.000 |
| y |
-1.882 |
-12.789 |
0.000 |
| z |
0.000 |
0.000 |
-18.506 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.044 |
-1.882 |
0.000 |
| y |
-1.882 |
7.810 |
0.000 |
| z |
0.000 |
0.000 |
-0.766 |
|
| Polar |
|---|
| 3z2-r2 | -1.532 |
|---|
| x2-y2 | -9.903 |
|---|
| xy | -1.882 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.030 |
-0.183 |
0.000 |
| y |
-0.183 |
3.411 |
0.000 |
| z |
0.000 |
0.000 |
1.474 |
<r2> (average value of r
2) Å
2
| <r2> |
40.922 |
| (<r2>)1/2 |
6.397 |
