Jump to
S1C2
S1C3
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.690976 |
| Energy at 298.15K | -169.694974 |
| HF Energy | -169.690976 |
| Nuclear repulsion energy | 70.941726 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3903 |
3725 |
67.56 |
|
|
|
| 2 |
A' |
3577 |
3413 |
3.59 |
|
|
|
| 3 |
A' |
3049 |
2910 |
69.99 |
|
|
|
| 4 |
A' |
1813 |
1730 |
200.10 |
|
|
|
| 5 |
A' |
1450 |
1384 |
16.39 |
|
|
|
| 6 |
A' |
1351 |
1289 |
177.02 |
|
|
|
| 7 |
A' |
1210 |
1155 |
44.46 |
|
|
|
| 8 |
A' |
1083 |
1034 |
212.59 |
|
|
|
| 9 |
A' |
624 |
596 |
1.38 |
|
|
|
| 10 |
A" |
1053 |
1005 |
5.78 |
|
|
|
| 11 |
A" |
838 |
800 |
64.94 |
|
|
|
| 12 |
A" |
395 |
377 |
71.87 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10173.2 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 9708.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.373 |
0.000 |
| O2 |
-1.002 |
-0.529 |
0.000 |
| N3 |
1.197 |
-0.020 |
0.000 |
| H4 |
-0.338 |
1.420 |
0.000 |
| H5 |
-1.845 |
-0.065 |
0.000 |
| H6 |
1.821 |
0.782 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3480 | 1.2599 | 1.1000 | 1.8958 | 1.8663 |
O2 | 1.3480 | | 2.2571 | 2.0583 | 0.9616 | 3.1126 | N3 | 1.2599 | 2.2571 | | 2.1048 | 3.0419 | 1.0168 | H4 | 1.1000 | 2.0583 | 2.1048 | | 2.1150 | 2.2510 | H5 | 1.8958 | 0.9616 | 3.0419 | 2.1150 | | 3.7621 | H6 | 1.8663 | 3.1126 | 1.0168 | 2.2510 | 3.7621 | |
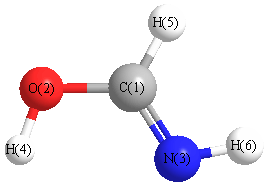 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.196 |
|
C1 |
N3 |
H6 |
109.659 |
| O2 |
C1 |
N3 |
119.838 |
|
O2 |
C1 |
H4 |
114.072 |
| N3 |
C1 |
H4 |
126.090 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.284 |
|
|
|
| 2 |
O |
-0.461 |
|
|
|
| 3 |
N |
-0.507 |
|
|
|
| 4 |
H |
0.100 |
|
|
|
| 5 |
H |
0.334 |
|
|
|
| 6 |
H |
0.251 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.097 |
3.179 |
0.000 |
3.808 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.543 |
1.242 |
0.000 |
| y |
1.242 |
-17.635 |
0.000 |
| z |
0.000 |
0.000 |
-18.431 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.490 |
1.242 |
0.000 |
| y |
1.242 |
-1.648 |
0.000 |
| z |
0.000 |
0.000 |
-2.842 |
|
| Polar |
|---|
| 3z2-r2 | -5.683 |
|---|
| x2-y2 | 4.092 |
|---|
| xy | 1.242 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.802 |
0.022 |
0.000 |
| y |
0.022 |
2.861 |
0.000 |
| z |
0.000 |
0.000 |
1.491 |
<r2> (average value of r
2) Å
2
| <r2> |
40.928 |
| (<r2>)1/2 |
6.398 |
Jump to
S1C1
S1C3
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.701440 |
| Energy at 298.15K | -169.705611 |
| HF Energy | -169.701440 |
| Nuclear repulsion energy | 71.301507 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3792 |
3619 |
45.33 |
|
|
|
| 2 |
A' |
3581 |
3417 |
3.80 |
|
|
|
| 3 |
A' |
3138 |
2995 |
37.64 |
|
|
|
| 4 |
A' |
1782 |
1701 |
263.25 |
|
|
|
| 5 |
A' |
1422 |
1357 |
29.76 |
|
|
|
| 6 |
A' |
1394 |
1330 |
1.90 |
|
|
|
| 7 |
A' |
1210 |
1155 |
125.10 |
|
|
|
| 8 |
A' |
1094 |
1044 |
156.68 |
|
|
|
| 9 |
A' |
593 |
566 |
53.10 |
|
|
|
| 10 |
A" |
1064 |
1015 |
0.04 |
|
|
|
| 11 |
A" |
840 |
801 |
25.48 |
|
|
|
| 12 |
A" |
649 |
620 |
202.23 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10278.9 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 9809.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.416 |
0.000 |
| O2 |
-1.110 |
-0.332 |
0.000 |
| N3 |
1.155 |
-0.098 |
0.000 |
| H4 |
-0.262 |
1.478 |
0.000 |
| H5 |
-0.810 |
-1.254 |
0.000 |
| H6 |
1.871 |
0.623 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3388 | 1.2641 | 1.0941 | 1.8559 | 1.8823 |
O2 | 1.3388 | | 2.2771 | 1.9995 | 0.9697 | 3.1305 | N3 | 1.2641 | 2.2771 | | 2.1195 | 2.2792 | 1.0165 | H4 | 1.0941 | 1.9995 | 2.1195 | | 2.7868 | 2.2978 | H5 | 1.8559 | 0.9697 | 2.2792 | 2.7868 | | 3.2724 | H6 | 1.8823 | 3.1305 | 1.0165 | 2.2978 | 3.2724 | |
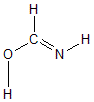 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
105.908 |
|
C1 |
N3 |
H6 |
110.786 |
| O2 |
C1 |
N3 |
122.022 |
|
O2 |
C1 |
H4 |
110.133 |
| N3 |
C1 |
H4 |
127.845 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.309 |
|
|
|
| 2 |
O |
-0.475 |
|
|
|
| 3 |
N |
-0.556 |
|
|
|
| 4 |
H |
0.130 |
|
|
|
| 5 |
H |
0.337 |
|
|
|
| 6 |
H |
0.256 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.058 |
0.954 |
0.000 |
0.956 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.114 |
3.300 |
0.000 |
| y |
3.300 |
-13.881 |
0.000 |
| z |
0.000 |
0.000 |
-18.430 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.958 |
3.300 |
0.000 |
| y |
3.300 |
4.891 |
0.000 |
| z |
0.000 |
0.000 |
-1.933 |
|
| Polar |
|---|
| 3z2-r2 | -3.866 |
|---|
| x2-y2 | -5.233 |
|---|
| xy | 3.300 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.284 |
0.102 |
0.000 |
| y |
0.102 |
3.254 |
0.000 |
| z |
0.000 |
0.000 |
1.484 |
<r2> (average value of r
2) Å
2
| <r2> |
40.262 |
| (<r2>)1/2 |
6.345 |
Jump to
S1C1
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.695849 |
| Energy at 298.15K | -169.699954 |
| HF Energy | -169.695849 |
| Nuclear repulsion energy | 70.868707 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3829 |
3654 |
25.97 |
|
|
|
| 2 |
A' |
3489 |
3330 |
6.13 |
|
|
|
| 3 |
A' |
3200 |
3054 |
14.44 |
|
|
|
| 4 |
A' |
1781 |
1699 |
267.77 |
|
|
|
| 5 |
A' |
1419 |
1354 |
1.00 |
|
|
|
| 6 |
A' |
1369 |
1307 |
25.02 |
|
|
|
| 7 |
A' |
1139 |
1087 |
254.93 |
|
|
|
| 8 |
A' |
1104 |
1054 |
54.21 |
|
|
|
| 9 |
A' |
597 |
570 |
36.58 |
|
|
|
| 10 |
A" |
1086 |
1036 |
65.29 |
|
|
|
| 11 |
A" |
856 |
816 |
62.84 |
|
|
|
| 12 |
A" |
536 |
511 |
84.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10201.6 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 9735.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.435 |
0.000 |
| O2 |
-1.100 |
-0.348 |
0.000 |
| N3 |
1.223 |
0.122 |
0.000 |
| H4 |
-0.298 |
1.484 |
0.000 |
| H5 |
-0.830 |
-1.276 |
0.000 |
| H6 |
1.365 |
-0.891 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3504 | 1.2629 | 1.0899 | 1.9018 | 1.9034 |
O2 | 1.3504 | | 2.3707 | 1.9994 | 0.9664 | 2.5247 | N3 | 1.2629 | 2.3707 | | 2.0417 | 2.4843 | 1.0230 | H4 | 1.0899 | 1.9994 | 2.0417 | | 2.8102 | 2.8992 | H5 | 1.9018 | 0.9664 | 2.4843 | 2.8102 | | 2.2291 | H6 | 1.9034 | 2.5247 | 1.0230 | 2.8992 | 2.2291 | |
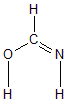 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.222 |
|
C1 |
N3 |
H6 |
112.333 |
| O2 |
C1 |
N3 |
130.202 |
|
O2 |
C1 |
H4 |
109.572 |
| N3 |
C1 |
H4 |
120.226 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.283 |
|
|
|
| 2 |
O |
-0.473 |
|
|
|
| 3 |
N |
-0.514 |
|
|
|
| 4 |
H |
0.155 |
|
|
|
| 5 |
H |
0.321 |
|
|
|
| 6 |
H |
0.228 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.009 |
-1.943 |
0.000 |
2.190 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.575 |
-1.832 |
0.000 |
| y |
-1.832 |
-12.825 |
0.000 |
| z |
0.000 |
0.000 |
-18.434 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.945 |
-1.832 |
0.000 |
| y |
-1.832 |
7.679 |
0.000 |
| z |
0.000 |
0.000 |
-0.733 |
|
| Polar |
|---|
| 3z2-r2 | -1.467 |
|---|
| x2-y2 | -9.749 |
|---|
| xy | -1.832 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.049 |
-0.187 |
0.000 |
| y |
-0.187 |
3.459 |
0.000 |
| z |
0.000 |
0.000 |
1.505 |
<r2> (average value of r
2) Å
2
| <r2> |
40.843 |
| (<r2>)1/2 |
6.391 |
