Jump to
S1C2
S1C3
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.405539 |
| Energy at 298.15K | -169.409504 |
| HF Energy | -168.906985 |
| Nuclear repulsion energy | 70.676431 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3950 |
3704 |
62.66 |
|
|
|
| 2 |
A' |
3602 |
3377 |
1.51 |
|
|
|
| 3 |
A' |
3143 |
2947 |
52.35 |
|
|
|
| 4 |
A' |
1829 |
1714 |
156.12 |
|
|
|
| 5 |
A' |
1489 |
1396 |
14.31 |
|
|
|
| 6 |
A' |
1374 |
1289 |
163.05 |
|
|
|
| 7 |
A' |
1234 |
1157 |
59.18 |
|
|
|
| 8 |
A' |
1098 |
1030 |
193.71 |
|
|
|
| 9 |
A' |
621 |
582 |
0.89 |
|
|
|
| 10 |
A" |
1071 |
1004 |
9.03 |
|
|
|
| 11 |
A" |
856 |
802 |
54.28 |
|
|
|
| 12 |
A" |
349 |
328 |
81.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10308.0 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 9664.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.378 |
0.000 |
| O2 |
-1.005 |
-0.536 |
0.000 |
| N3 |
1.201 |
-0.017 |
0.000 |
| H4 |
-0.333 |
1.419 |
0.000 |
| H5 |
-1.849 |
-0.076 |
0.000 |
| H6 |
1.816 |
0.792 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3589 | 1.2647 | 1.0923 | 1.9039 | 1.8628 |
O2 | 1.3589 | | 2.2668 | 2.0671 | 0.9607 | 3.1186 | N3 | 1.2647 | 2.2668 | | 2.1012 | 3.0507 | 1.0166 | H4 | 1.0923 | 2.0671 | 2.1012 | | 2.1290 | 2.2385 | H5 | 1.9039 | 0.9607 | 3.0507 | 2.1290 | | 3.7666 | H6 | 1.8628 | 3.1186 | 1.0166 | 2.2385 | 3.7666 | |
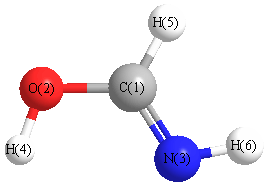 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.118 |
|
C1 |
N3 |
H6 |
109.006 |
| O2 |
C1 |
N3 |
119.498 |
|
O2 |
C1 |
H4 |
114.541 |
| N3 |
C1 |
H4 |
125.961 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.415671 |
| Energy at 298.15K | -169.419844 |
| HF Energy | -168.918408 |
| Nuclear repulsion energy | 71.036839 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3871 |
3630 |
48.65 |
|
|
|
| 2 |
A' |
3615 |
3389 |
2.08 |
|
|
|
| 3 |
A' |
3218 |
3017 |
30.29 |
|
|
|
| 4 |
A' |
1806 |
1693 |
225.32 |
|
|
|
| 5 |
A' |
1461 |
1370 |
24.94 |
|
|
|
| 6 |
A' |
1416 |
1327 |
5.43 |
|
|
|
| 7 |
A' |
1231 |
1154 |
99.75 |
|
|
|
| 8 |
A' |
1108 |
1039 |
186.65 |
|
|
|
| 9 |
A' |
597 |
560 |
51.99 |
|
|
|
| 10 |
A" |
1083 |
1015 |
0.23 |
|
|
|
| 11 |
A" |
860 |
807 |
32.43 |
|
|
|
| 12 |
A" |
620 |
581 |
200.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10442.6 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 9790.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.419 |
0.000 |
| O2 |
-1.114 |
-0.343 |
0.000 |
| N3 |
1.161 |
-0.090 |
0.000 |
| H4 |
-0.261 |
1.475 |
0.000 |
| H5 |
-0.812 |
-1.261 |
0.000 |
| H6 |
1.864 |
0.642 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3502 | 1.2675 | 1.0874 | 1.8668 | 1.8777 |
O2 | 1.3502 | | 2.2890 | 2.0082 | 0.9669 | 3.1375 | N3 | 1.2675 | 2.2890 | | 2.1144 | 2.2946 | 1.0156 | H4 | 1.0874 | 2.0082 | 2.1144 | | 2.7914 | 2.2828 | H5 | 1.8668 | 0.9669 | 2.2946 | 2.7914 | | 3.2847 | H6 | 1.8777 | 3.1375 | 1.0156 | 2.2828 | 3.2847 | |
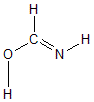 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
106.173 |
|
C1 |
N3 |
H6 |
110.169 |
| O2 |
C1 |
N3 |
121.930 |
|
O2 |
C1 |
H4 |
110.481 |
| N3 |
C1 |
H4 |
127.590 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCD/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.410330 |
| Energy at 298.15K | -169.414424 |
| HF Energy | -168.912325 |
| Nuclear repulsion energy | 70.638573 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3896 |
3653 |
28.16 |
|
|
|
| 2 |
A' |
3532 |
3311 |
6.59 |
|
|
|
| 3 |
A' |
3275 |
3070 |
14.27 |
|
|
|
| 4 |
A' |
1800 |
1687 |
220.39 |
|
|
|
| 5 |
A' |
1455 |
1364 |
3.46 |
|
|
|
| 6 |
A' |
1402 |
1314 |
21.03 |
|
|
|
| 7 |
A' |
1161 |
1088 |
257.61 |
|
|
|
| 8 |
A' |
1122 |
1052 |
51.12 |
|
|
|
| 9 |
A' |
602 |
565 |
38.13 |
|
|
|
| 10 |
A" |
1113 |
1044 |
68.38 |
|
|
|
| 11 |
A" |
867 |
813 |
55.20 |
|
|
|
| 12 |
A" |
496 |
465 |
95.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10359.7 cm
-1
Scaled (by 0.9376) Zero Point Vibrational Energy (zpe) 9713.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.442 |
0.000 |
| O2 |
-1.103 |
-0.357 |
0.000 |
| N3 |
1.227 |
0.127 |
0.000 |
| H4 |
-0.299 |
1.483 |
0.000 |
| H5 |
-0.827 |
-1.281 |
0.000 |
| H6 |
1.358 |
-0.886 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3613 | 1.2669 | 1.0834 | 1.9112 | 1.8992 |
O2 | 1.3613 | | 2.3793 | 2.0074 | 0.9644 | 2.5165 | N3 | 1.2669 | 2.3793 | | 2.0419 | 2.4905 | 1.0215 | H4 | 1.0834 | 2.0074 | 2.0419 | | 2.8140 | 2.8912 | H5 | 1.9112 | 0.9644 | 2.4905 | 2.8140 | | 2.2203 | H6 | 1.8992 | 2.5165 | 1.0215 | 2.8912 | 2.2203 | |
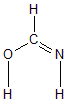 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.334 |
|
C1 |
N3 |
H6 |
111.739 |
| O2 |
C1 |
N3 |
129.689 |
|
O2 |
C1 |
H4 |
109.871 |
| N3 |
C1 |
H4 |
120.440 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
