Jump to
S1C2
S1C3
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.866202 |
| Energy at 298.15K | -169.870187 |
| HF Energy | -169.866202 |
| Nuclear repulsion energy | 70.557061 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3811 |
3653 |
48.61 |
|
|
|
| 2 |
A' |
3506 |
3361 |
0.84 |
|
|
|
| 3 |
A' |
3030 |
2905 |
76.89 |
|
|
|
| 4 |
A' |
1769 |
1696 |
181.87 |
|
|
|
| 5 |
A' |
1437 |
1378 |
14.16 |
|
|
|
| 6 |
A' |
1336 |
1281 |
159.23 |
|
|
|
| 7 |
A' |
1196 |
1147 |
53.00 |
|
|
|
| 8 |
A' |
1065 |
1021 |
214.99 |
|
|
|
| 9 |
A' |
609 |
584 |
1.44 |
|
|
|
| 10 |
A" |
1036 |
993 |
4.95 |
|
|
|
| 11 |
A" |
827 |
793 |
65.65 |
|
|
|
| 12 |
A" |
403 |
386 |
66.15 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10013.1 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 9599.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.379 |
0.000 |
| O2 |
-1.006 |
-0.534 |
0.000 |
| N3 |
1.202 |
-0.020 |
0.000 |
| H4 |
-0.343 |
1.423 |
0.000 |
| H5 |
-1.853 |
-0.066 |
0.000 |
| H6 |
1.827 |
0.787 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3585 | 1.2669 | 1.0989 | 1.9056 | 1.8719 |
O2 | 1.3585 | | 2.2672 | 2.0662 | 0.9677 | 3.1258 | N3 | 1.2669 | 2.2672 | | 2.1146 | 3.0555 | 1.0210 | H4 | 1.0989 | 2.0662 | 2.1146 | | 2.1205 | 2.2613 | H5 | 1.9056 | 0.9677 | 3.0555 | 2.1205 | | 3.7775 | H6 | 1.8719 | 3.1258 | 1.0210 | 2.2613 | 3.7775 | |
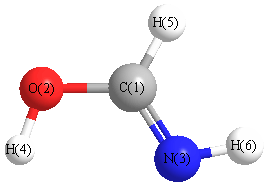 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.849 |
|
C1 |
N3 |
H6 |
109.343 |
| O2 |
C1 |
N3 |
119.394 |
|
O2 |
C1 |
H4 |
114.031 |
| N3 |
C1 |
H4 |
126.575 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.290 |
|
|
|
| 2 |
O |
-0.459 |
|
|
|
| 3 |
N |
-0.508 |
|
|
|
| 4 |
H |
0.099 |
|
|
|
| 5 |
H |
0.331 |
|
|
|
| 6 |
H |
0.247 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.080 |
3.139 |
0.000 |
3.765 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.929 |
0.047 |
0.000 |
| y |
0.047 |
2.929 |
0.000 |
| z |
0.000 |
0.000 |
1.499 |
<r2> (average value of r
2) Å
2
| <r2> |
41.234 |
| (<r2>)1/2 |
6.421 |
Jump to
S1C1
S1C3
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.866202 |
| Energy at 298.15K | -169.870187 |
| HF Energy | -169.866202 |
| Nuclear repulsion energy | 70.557061 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
Geometric Data calculated at TPSSh/6-31G**
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at TPSSh/6-31G**
| | hartrees |
|---|
| Energy at 0K | -169.871530 |
| Energy at 298.15K | -169.875614 |
| HF Energy | -169.871530 |
| Nuclear repulsion energy | 70.430721 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at TPSSh/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3738 |
3584 |
17.26 |
|
|
|
| 2 |
A' |
3423 |
3281 |
9.57 |
|
|
|
| 3 |
A' |
3194 |
3062 |
15.43 |
|
|
|
| 4 |
A' |
1735 |
1663 |
248.96 |
|
|
|
| 5 |
A' |
1421 |
1363 |
0.88 |
|
|
|
| 6 |
A' |
1355 |
1299 |
27.87 |
|
|
|
| 7 |
A' |
1116 |
1070 |
251.90 |
|
|
|
| 8 |
A' |
1087 |
1042 |
49.52 |
|
|
|
| 9 |
A' |
579 |
555 |
35.26 |
|
|
|
| 10 |
A" |
1081 |
1036 |
69.13 |
|
|
|
| 11 |
A" |
840 |
806 |
51.25 |
|
|
|
| 12 |
A" |
537 |
515 |
83.25 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10052.7 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 9637.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at TPSSh/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.437 |
0.000 |
| O2 |
-1.108 |
-0.355 |
0.000 |
| N3 |
1.232 |
0.129 |
0.000 |
| H4 |
-0.305 |
1.482 |
0.000 |
| H5 |
-0.830 |
-1.287 |
0.000 |
| H6 |
1.376 |
-0.887 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3620 | 1.2697 | 1.0882 | 1.9135 | 1.9104 |
O2 | 1.3620 | | 2.3893 | 2.0044 | 0.9723 | 2.5408 | N3 | 1.2697 | 2.3893 | | 2.0475 | 2.5013 | 1.0269 | H4 | 1.0882 | 2.0044 | 2.0475 | | 2.8178 | 2.9055 | H5 | 1.9135 | 0.9723 | 2.5013 | 2.8178 | | 2.2424 | H6 | 1.9104 | 2.5408 | 1.0269 | 2.9055 | 2.2424 | |
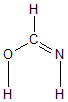 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.971 |
|
C1 |
N3 |
H6 |
112.138 |
| O2 |
C1 |
N3 |
130.391 |
|
O2 |
C1 |
H4 |
109.272 |
| N3 |
C1 |
H4 |
120.337 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at TPSSh/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.286 |
|
|
|
| 2 |
O |
-0.471 |
|
|
|
| 3 |
N |
-0.512 |
|
|
|
| 4 |
H |
0.154 |
|
|
|
| 5 |
H |
0.319 |
|
|
|
| 6 |
H |
0.225 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.986 |
-1.910 |
0.000 |
2.149 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.138 |
-0.164 |
0.000 |
| y |
-0.164 |
3.533 |
0.000 |
| z |
0.000 |
0.000 |
1.510 |
<r2> (average value of r
2) Å
2
| <r2> |
41.236 |
| (<r2>)1/2 |
6.421 |
