return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C10H8 (naphthalene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1Ag
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-384.729245
Energy at 298.15K 
HF Energy-383.367480
Nuclear repulsion energy458.123626
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3278 3063 0.00      
2 Ag 3248 3035 0.00      
3 Ag 1650 1542 0.00      
4 Ag 1523 1423 0.00      
5 Ag 1473 1376 0.00      
6 Ag 1209 1130 0.00      
7 Ag 1071 1000 0.00      
8 Ag 783 731 0.00      
9 Ag 523 489 0.00      
10 Au 888 830 0.00      
11 Au 825 771 0.00      
12 Au 479 448 0.00      
13 Au 182 170 0.00      
14 B1g 897 838 0.00      
15 B1g 718 671 0.00      
16 B1g 369 345 0.00      
17 B1u 3264 3050 43.96      
18 B1u 3244 3031 13.35      
19 B1u 1667 1558 4.03      
20 B1u 1444 1350 5.90      
21 B1u 1310 1224 9.55      
22 B1u 1167 1090 1.53      
23 B1u 815 762 0.00      
24 B1u 361 338 0.93      
25 B2g 919 859 0.00      
26 B2g 834 780 0.00      
27 B2g 440 411 0.00      
28 B2g 114i 107i 0.00      
29 B2u 3277 3062 42.54      
30 B2u 3246 3033 0.00      
31 B2u 1590 1486 6.61      
32 B2u 1498 1400 1.20      
33 B2u 1277 1193 2.21      
34 B2u 1207 1128 0.39      
35 B2u 1059 990 3.72      
36 B2u 632 590 3.62      
37 B3g 3264 3050 0.00      
38 B3g 3243 3030 0.00      
39 B3g 1715 1602 0.00      
40 B3g 1519 1419 0.00      
41 B3g 1286 1202 0.00      
42 B3g 1193 1115 0.00      
43 B3g 945 883 0.00      
44 B3g 515 481 0.00      
45 B3u 899 840 0.08      
46 B3u 772 721 109.06      
47 B3u 450 420 8.87      
48 B3u 168 157 1.79      

Unscaled Zero Point Vibrational Energy (zpe) 32109.9 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 30003.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.10428 0.04123 0.02955

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.240 1.402
C2 0.000 2.429 0.706
C3 0.000 2.429 -0.706
C4 0.000 1.240 -1.402
C5 0.000 -1.240 -1.402
C6 0.000 -2.429 -0.706
C7 0.000 -2.429 0.706
C8 0.000 -1.240 1.402
C9 0.000 0.000 0.715
C10 0.000 0.000 -0.715
H11 0.000 1.238 2.485
H12 0.000 3.369 1.242
H13 0.000 3.369 -1.242
H14 0.000 1.238 -2.485
H15 0.000 -1.238 -2.485
H16 0.000 -3.369 -1.242
H17 0.000 -3.369 1.242
H18 0.000 -1.238 2.485

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 C10 H11 H12 H13 H14 H15 H16 H17 H18
C11.37752.42032.80343.74264.23113.73382.47951.41712.45331.08342.13493.39443.88684.60945.31294.61112.7042
C21.37751.41272.42034.23115.05884.85763.73382.42882.81432.14061.08182.16333.40644.86116.11615.82214.0754
C32.42031.41271.37753.73384.85765.05884.23112.81432.42883.40642.16331.08182.14064.07545.82216.11614.8611
C42.80342.42031.37752.47953.73384.23113.74262.45331.41713.88683.39442.13491.08342.70424.61115.31294.6094
C53.74264.23113.73382.47951.37752.42032.80342.45331.41714.60945.31294.61112.70421.08342.13493.39443.8868
C64.23115.05884.85763.73381.37751.41272.42032.81432.42884.86116.11615.82214.07542.14061.08182.16333.4064
C73.73384.85765.05884.23112.42031.41271.37752.42882.81434.07545.82216.11614.86113.40642.16331.08182.1406
C82.47953.73384.23113.74262.80342.42031.37751.41712.45332.70424.61115.31294.60943.88683.39442.13491.0834
C91.41712.42882.81432.45332.45332.81432.42881.41711.43062.15983.40963.89603.43153.43153.89603.40962.1598
C102.45332.81432.42881.41711.41712.42882.81432.45331.43063.43153.89603.40962.15982.15983.40963.89603.4315
H111.08342.14063.40643.88684.60944.86114.07542.70422.15983.43152.46684.29324.97025.55285.92564.77142.4759
H122.13491.08182.16333.39445.31296.11615.82214.61113.40963.89602.46682.48424.29325.92567.18076.73734.7714
H133.39442.16331.08182.13494.61115.82216.11615.31293.89603.40964.29322.48422.46684.77146.73737.18075.9256
H143.88683.40642.14061.08342.70424.07544.86114.60943.43152.15984.97024.29322.46682.47594.77145.92565.5528
H154.60944.86114.07542.70421.08342.14063.40643.88683.43152.15985.55285.92564.77142.47592.46684.29324.9702
H165.31296.11615.82214.61112.13491.08182.16333.39443.89603.40965.92567.18076.73734.77142.46682.48424.2932
H174.61115.82216.11615.31293.39442.16331.08182.13493.40963.89604.77146.73737.18075.92564.29322.48422.4668
H182.70424.07544.86114.60943.88683.40642.14061.08342.15983.43152.47594.77145.92565.55284.97024.29322.4668

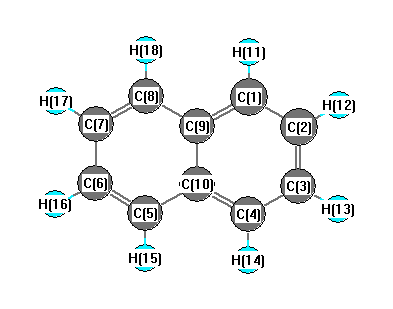
picture of naphthalene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.320 C1 C2 H12 119.995
C1 C9 C8 122.054 C1 C9 C10 118.973
C2 C1 C9 120.707 C2 C1 H11 120.415
C2 C3 C4 120.320 C2 C3 H13 119.685
C3 C2 H12 119.685 C3 C4 C10 120.707
C3 C4 H14 120.415 C4 C3 H13 119.995
C4 C10 C5 122.054 C4 C10 C9 118.973
C5 C6 C7 120.320 C5 C6 H16 119.995
C5 C10 C9 118.973 C6 C5 C10 120.707
C6 C5 H15 120.415 C6 C7 C8 120.320
C6 C7 H17 119.685 C7 C6 H16 119.685
C7 C8 C9 120.707 C7 C8 H18 120.415
C8 C7 H17 119.995 C8 C9 C10 118.973
C9 C1 H11 118.878 C9 C8 H18 118.878
C10 C4 H14 118.878 C10 C5 H15 118.878
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability