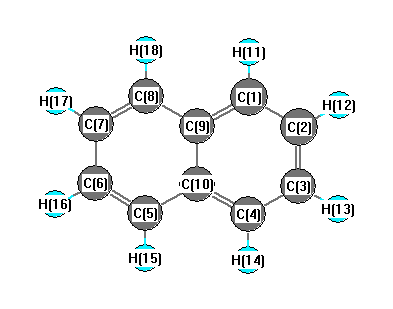
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3237 |
3085 |
0.00 |
|
|
|
| 2 |
Ag |
3210 |
3059 |
0.00 |
|
|
|
| 3 |
Ag |
1659 |
1581 |
0.00 |
|
|
|
| 4 |
Ag |
1514 |
1443 |
0.00 |
|
|
|
| 5 |
Ag |
1453 |
1385 |
0.00 |
|
|
|
| 6 |
Ag |
1195 |
1139 |
0.00 |
|
|
|
| 7 |
Ag |
1067 |
1017 |
0.00 |
|
|
|
| 8 |
Ag |
787 |
750 |
0.00 |
|
|
|
| 9 |
Ag |
522 |
498 |
0.00 |
|
|
|
| 10 |
Au |
994 |
947 |
0.00 |
|
|
|
| 11 |
Au |
855 |
815 |
0.00 |
|
|
|
| 12 |
Au |
637 |
607 |
0.00 |
|
|
|
| 13 |
Au |
190 |
181 |
0.00 |
|
|
|
| 14 |
B1g |
955 |
910 |
0.00 |
|
|
|
| 15 |
B1g |
734 |
700 |
0.00 |
|
|
|
| 16 |
B1g |
397 |
378 |
0.00 |
|
|
|
| 17 |
B1u |
3223 |
3072 |
52.55 |
|
|
|
| 18 |
B1u |
3205 |
3054 |
8.26 |
|
|
|
| 19 |
B1u |
1681 |
1602 |
5.14 |
|
|
|
| 20 |
B1u |
1432 |
1365 |
4.82 |
|
|
|
| 21 |
B1u |
1302 |
1241 |
8.12 |
|
|
|
| 22 |
B1u |
1161 |
1107 |
4.64 |
|
|
|
| 23 |
B1u |
810 |
772 |
0.25 |
|
|
|
| 24 |
B1u |
362 |
345 |
1.25 |
|
|
|
| 25 |
B2g |
1002 |
955 |
0.00 |
|
|
|
| 26 |
B2g |
902 |
860 |
0.00 |
|
|
|
| 27 |
B2g |
786 |
749 |
0.00 |
|
|
|
| 28 |
B2g |
479 |
457 |
0.00 |
|
|
|
| 29 |
B2u |
3236 |
3084 |
44.67 |
|
|
|
| 30 |
B2u |
3207 |
3057 |
0.53 |
|
|
|
| 31 |
B2u |
1584 |
1510 |
8.43 |
|
|
|
| 32 |
B2u |
1433 |
1366 |
1.73 |
|
|
|
| 33 |
B2u |
1257 |
1198 |
1.83 |
|
|
|
| 34 |
B2u |
1186 |
1131 |
0.86 |
|
|
|
| 35 |
B2u |
1058 |
1008 |
5.81 |
|
|
|
| 36 |
B2u |
632 |
603 |
3.68 |
|
|
|
| 37 |
B3g |
3223 |
3072 |
0.00 |
|
|
|
| 38 |
B3g |
3204 |
3053 |
0.00 |
|
|
|
| 39 |
B3g |
1717 |
1636 |
0.00 |
|
|
|
| 40 |
B3g |
1515 |
1444 |
0.00 |
|
|
|
| 41 |
B3g |
1279 |
1219 |
0.00 |
|
|
|
| 42 |
B3g |
1186 |
1130 |
0.00 |
|
|
|
| 43 |
B3g |
949 |
905 |
0.00 |
|
|
|
| 44 |
B3g |
517 |
493 |
0.00 |
|
|
|
| 45 |
B3u |
972 |
927 |
3.93 |
|
|
|
| 46 |
B3u |
807 |
769 |
108.49 |
|
|
|
| 47 |
B3u |
490 |
467 |
15.99 |
|
|
|
| 48 |
B3u |
175 |
167 |
2.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32688.7 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 31155.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.209 |
|
|
|
| 2 |
C |
-0.177 |
|
|
|
| 3 |
C |
-0.177 |
|
|
|
| 4 |
C |
-0.209 |
|
|
|
| 5 |
C |
-0.209 |
|
|
|
| 6 |
C |
-0.177 |
|
|
|
| 7 |
C |
-0.177 |
|
|
|
| 8 |
C |
-0.209 |
|
|
|
| 9 |
C |
0.092 |
|
|
|
| 10 |
C |
0.092 |
|
|
|
| 11 |
H |
0.170 |
|
|
|
| 12 |
H |
0.170 |
|
|
|
| 13 |
H |
0.170 |
|
|
|
| 14 |
H |
0.170 |
|
|
|
| 15 |
H |
0.170 |
|
|
|
| 16 |
H |
0.170 |
|
|
|
| 17 |
H |
0.170 |
|
|
|
| 18 |
H |
0.170 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-62.619 |
0.000 |
0.000 |
| y |
0.000 |
-49.732 |
0.000 |
| z |
0.000 |
0.000 |
-50.111 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-12.698 |
0.000 |
0.000 |
| y |
0.000 |
6.634 |
0.000 |
| z |
0.000 |
0.000 |
6.064 |
|
| Polar |
|---|
| 3z2-r2 | 12.129 |
|---|
| x2-y2 | -12.888 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.885 |
0.000 |
0.000 |
| y |
0.000 |
22.505 |
0.000 |
| z |
0.000 |
0.000 |
16.203 |
<r2> (average value of r
2) Å
2
| <r2> |
359.647 |
| (<r2>)1/2 |
18.964 |