return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (3-Hexyne)

using model chemistry: MP3=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP3=FULL/6-31G**
 hartrees
Energy at 0K-233.907985
Energy at 298.15K-233.917174
HF Energy-232.993578
Nuclear repulsion energy207.390738
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3239 3239 0.00      
2 A 3232 3232 9.90      
3 A 3179 3179 0.00      
4 A 3145 3145 24.13      
5 A 3138 3138 30.74      
6 A 2439 2439 0.00      
7 A 1574 1574 1.04      
8 A 1564 1564 0.00      
9 A 1551 1551 2.18      
10 A 1478 1478 1.66      
11 A 1412 1412 0.02      
12 A 1335 1335 0.00      
13 A 1137 1137 0.00      
14 A 1134 1134 1.15      
15 A 1035 1035 0.01      
16 A 806 806 0.00      
17 A 714 714 0.22      
18 A 434 434 5.97      
19 A 379 379 0.00      
20 A 232 232 0.00      
21 A 91 91 1.28      
22 A 16 16 0.00      
23 B 3239 3239 58.23      
24 B 3231 3231 58.43      
25 B 3178 3178 22.70      
26 B 3145 3145 24.19      
27 B 3138 3138 25.55      
28 B 1573 1573 1.33      
29 B 1564 1564 8.77      
30 B 1549 1549 1.73      
31 B 1477 1477 0.39      
32 B 1416 1416 41.14      
33 B 1338 1338 0.02      
34 B 1196 1196 0.27      
35 B 1156 1156 0.64      
36 B 1122 1122 3.81      
37 B 963 963 0.03      
38 B 811 811 2.10      
39 B 546 546 0.44      
40 B 303 303 3.10      
41 B 207 207 0.13      
42 B 161 161 3.65      

Unscaled Zero Point Vibrational Energy (zpe) 32288.7 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 32288.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3=FULL/6-31G**
ABC
0.31354 0.04700 0.04214

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3=FULL/6-31G**

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.603 -0.572
C2 -0.000 -0.603 -0.572
C3 0.000 2.073 -0.550
C4 -0.000 -2.073 -0.550
C5 -0.000 2.640 0.872
C6 0.000 -2.640 0.872
H7 0.873 2.438 -1.091
H8 -0.873 2.438 -1.091
H9 -0.873 -2.438 -1.091
H10 0.873 -2.438 -1.091
H11 -0.001 3.727 0.849
H12 -0.880 2.306 1.416
H13 0.880 2.307 1.416
H14 0.001 -3.727 0.849
H15 0.880 -2.306 1.416
H16 -0.880 -2.307 1.416

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.20691.46922.67602.49633.55022.09672.09673.20633.20623.43172.76152.76194.55783.63213.6328
C21.20692.67601.46923.55022.49633.20633.20622.09672.09674.55783.63213.63283.43172.76152.7619
C31.46922.67604.14501.53084.92221.09001.09004.62574.62572.16662.16652.16645.96614.87984.8806
C42.67601.46924.14504.92221.53084.62574.62571.09001.09005.96614.87984.88062.16662.16652.1664
C52.49633.55021.53084.92225.27982.15762.15765.51335.51341.08771.08681.08686.36735.05325.0541
C63.55022.49634.92221.53085.27985.51335.51342.15762.15766.36735.05325.05411.08771.08681.0868
H72.09673.20631.09004.62572.15765.51331.74595.17884.87552.48743.06172.51006.52165.36575.6454
H82.09673.20621.09004.62572.15765.51341.74594.87555.17862.48712.51053.06176.52175.64475.3664
H93.20632.09674.62571.09005.51332.15765.17884.87551.74596.52165.36575.64542.48743.06172.5100
H103.20622.09674.62571.09005.51342.15764.87555.17861.74596.52175.64475.36642.48712.51053.0617
H113.43174.55782.16665.96611.08776.36732.48742.48716.52166.52171.76471.76477.45476.12386.1246
H122.76153.63212.16654.87981.08685.05323.06172.51055.36575.64471.76471.75976.12384.93674.6134
H132.76193.63282.16644.88061.08685.05412.51003.06175.64545.36641.76471.75976.12464.61344.9386
H144.55783.43175.96612.16666.36731.08776.52166.52172.48742.48717.45476.12386.12461.76471.7647
H153.63212.76154.87982.16655.05321.08685.36575.64473.06172.51056.12384.93674.61341.76471.7597
H163.63282.76194.88062.16645.05411.08685.64545.36642.51003.06176.12464.61344.93861.76471.7597

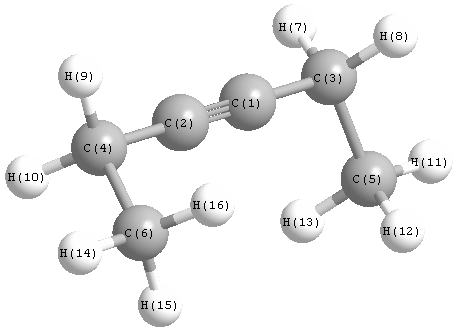
picture of 3-Hexyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 179.142 C1 C3 C5 112.614
C1 C3 H7 109.123 C1 C3 H8 109.124
C2 C1 C3 179.142 C2 C4 C6 112.614
C2 C4 H9 109.123 C2 C4 H10 109.124
C3 C5 H11 110.527 C3 C5 H12 110.571
C3 C5 H13 110.568 C4 C6 H14 110.527
C4 C6 H15 110.571 C4 C6 H16 110.568
C5 C3 H7 109.680 C5 C3 H8 109.682
C6 C4 H9 109.680 C6 C4 H10 109.682
H7 C3 H8 106.425 H9 C4 H10 106.425
H11 C5 H12 108.496 H11 C5 H13 108.495
H12 C5 H13 108.106 H14 C6 H15 108.496
H14 C6 H16 108.495 H15 C6 H15 0.000
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability