return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C9H8 (Indene)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-346.709800
Energy at 298.15K-346.717996
HF Energy-345.475844
Nuclear repulsion energy394.974940
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3306 3089 14.62      
2 A' 3282 3067 3.95      
3 A' 3277 3062 22.04      
4 A' 3264 3050 23.44      
5 A' 3252 3039 1.21      
6 A' 3246 3033 5.92      
7 A' 3115 2911 9.80      
8 A' 1691 1580 1.16      
9 A' 1666 1556 0.12      
10 A' 1629 1522 0.24      
11 A' 1531 1431 8.29      
12 A' 1521 1421 5.83      
13 A' 1498 1400 3.36      
14 A' 1481 1384 4.90      
15 A' 1406 1314 4.45      
16 A' 1346 1258 0.05      
17 A' 1284 1200 1.18      
18 A' 1263 1180 2.42      
19 A' 1210 1131 1.21      
20 A' 1203 1124 0.09      
21 A' 1158 1082 0.23      
22 A' 1109 1036 0.49      
23 A' 1061 992 2.21      
24 A' 991 926 5.65      
25 A' 878 821 2.35      
26 A' 855 799 0.83      
27 A' 749 699 2.13      
28 A' 605 566 1.38      
29 A' 546 510 0.17      
30 A' 386 361 1.11      
31 A" 3164 2956 8.10      
32 A" 1163 1086 1.53      
33 A" 955 893 1.53      
34 A" 919 859 0.27      
35 A" 899 840 0.46      
36 A" 884 826 0.46      
37 A" 850 794 0.16      
38 A" 751 702 78.40      
39 A" 711 664 5.48      
40 A" 539 504 4.17      
41 A" 470 439 0.00      
42 A" 394 368 4.05      
43 A" 369 345 4.10      
44 A" 205 192 2.92      
45 A" 189 177 0.37      

Unscaled Zero Point Vibrational Energy (zpe) 31136.1 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 29093.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.12635 0.05288 0.03753

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -2.280 -0.016 0.000
C2 -1.851 -1.348 0.000
C3 -0.492 -1.661 0.000
C4 -1.355 1.031 0.000
C5 0.000 0.725 0.000
C6 0.432 -0.616 0.000
C7 1.894 -0.623 0.000
C8 2.351 0.648 0.000
C9 1.206 1.624 0.000
H10 -3.339 0.204 0.000
H11 -2.585 -2.143 0.000
H12 -0.165 -2.693 0.000
H13 -1.694 2.060 0.000
H14 2.503 -1.515 0.000
H15 3.389 0.948 0.000
H16 1.230 2.276 0.877
H17 1.230 2.276 -0.877

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17
C11.39892.42941.39762.39742.77804.21784.67873.85291.08192.14893.41162.15745.01215.75014.28304.2830
C21.39891.39502.43042.77892.39813.81484.65254.26392.14971.08192.15723.41154.35715.72074.83704.8370
C32.42941.39502.82722.43561.39512.60203.66283.69783.40362.14751.08273.91032.99804.67584.38574.3857
C41.39762.43042.82721.38932.43073.64573.72592.62902.15003.40463.90991.08324.62244.74443.00003.0000
C52.39742.77892.43561.38931.40852.32412.35241.50473.37973.86073.42162.15723.35863.39602.16552.1655
C62.77802.39811.39512.43071.40851.46152.29782.36983.85983.38162.16133.41802.25713.34433.12573.1257
C74.21783.81482.60203.64572.32411.46151.35062.34935.29814.72962.91994.48001.08062.16793.10063.1006
C84.67874.65253.66283.72592.35242.29781.35061.50415.70795.67064.18284.28452.16911.07982.16262.1626
C93.85294.26393.69782.62901.50472.36982.34931.50414.76245.34454.52952.93303.39642.28471.09331.0933
H101.08192.14973.40362.15003.37973.85985.29815.70794.76242.46554.29752.48056.08966.76905.09335.0933
H112.14891.08192.14753.40463.86073.38164.72965.67065.34452.46552.48124.29685.12576.72565.90355.9035
H123.41162.15721.08273.90993.42162.16132.91994.18284.52954.29752.48124.99302.91585.08745.23535.2353
H132.15743.41153.91031.08322.15723.41804.48004.28452.93302.48054.29684.99305.51325.20313.06023.0602
H145.01214.35712.99804.62243.35862.25711.08062.16913.39646.08965.12572.91585.51322.61764.09474.0947
H155.75015.72074.67584.74443.39603.34432.16791.07982.28476.76906.72565.08745.20312.61762.68252.6825
H164.28304.83704.38573.00002.16553.12573.10062.16261.09335.09335.90355.23533.06024.09472.68251.7541
H174.28304.83704.38573.00002.16553.12573.10062.16261.09335.09335.90355.23533.06024.09472.68251.7541

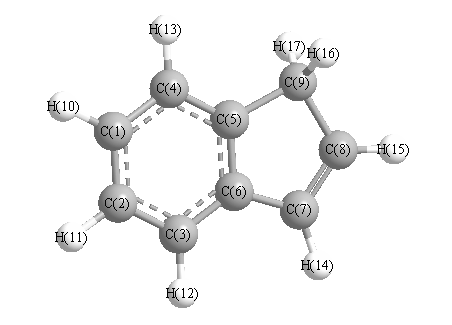
picture of Indene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.817 C1 C2 H11 119.491
C1 C4 C5 118.690 C1 C4 H13 120.310
C2 C1 C4 120.705 C2 C1 H10 119.578
C2 C3 C6 118.527 C2 C3 H12 120.551
C3 C2 H11 119.692 C3 C6 C5 120.626
C3 C6 C7 131.236 C4 C1 H10 119.717
C4 C5 C6 120.635 C4 C5 C9 130.537
C5 C4 H13 121.001 C5 C6 C7 108.138
C5 C9 C8 102.858 C5 C9 H16 111.951
C5 C9 H17 111.951 C6 C3 H12 120.923
C6 C5 C9 108.828 C6 C7 C8 109.528
C6 C7 H14 124.539 C7 C8 C9 110.647
C7 C8 H15 125.878 C8 C7 H14 125.933
C8 C9 H16 111.758 C8 C9 H17 111.758
C9 C8 H15 123.475 H16 C9 H17 106.682
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability