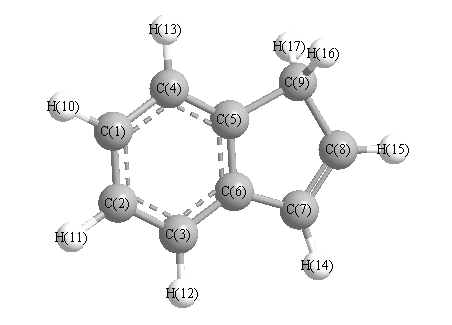
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3265 |
3112 |
15.06 |
|
|
|
| 2 |
A' |
3241 |
3089 |
4.92 |
|
|
|
| 3 |
A' |
3235 |
3083 |
23.12 |
|
|
|
| 4 |
A' |
3222 |
3071 |
28.01 |
|
|
|
| 5 |
A' |
3211 |
3060 |
2.77 |
|
|
|
| 6 |
A' |
3204 |
3054 |
4.06 |
|
|
|
| 7 |
A' |
3060 |
2917 |
11.32 |
|
|
|
| 8 |
A' |
1696 |
1616 |
2.57 |
|
|
|
| 9 |
A' |
1680 |
1601 |
0.40 |
|
|
|
| 10 |
A' |
1645 |
1568 |
0.31 |
|
|
|
| 11 |
A' |
1522 |
1451 |
11.25 |
|
|
|
| 12 |
A' |
1517 |
1446 |
5.81 |
|
|
|
| 13 |
A' |
1459 |
1391 |
9.59 |
|
|
|
| 14 |
A' |
1428 |
1361 |
1.30 |
|
|
|
| 15 |
A' |
1376 |
1311 |
3.90 |
|
|
|
| 16 |
A' |
1336 |
1274 |
0.14 |
|
|
|
| 17 |
A' |
1273 |
1213 |
2.20 |
|
|
|
| 18 |
A' |
1252 |
1193 |
2.76 |
|
|
|
| 19 |
A' |
1200 |
1144 |
1.16 |
|
|
|
| 20 |
A' |
1190 |
1134 |
0.03 |
|
|
|
| 21 |
A' |
1147 |
1093 |
0.48 |
|
|
|
| 22 |
A' |
1099 |
1048 |
1.20 |
|
|
|
| 23 |
A' |
1063 |
1013 |
3.46 |
|
|
|
| 24 |
A' |
982 |
936 |
9.37 |
|
|
|
| 25 |
A' |
877 |
836 |
2.80 |
|
|
|
| 26 |
A' |
850 |
810 |
1.45 |
|
|
|
| 27 |
A' |
749 |
714 |
2.87 |
|
|
|
| 28 |
A' |
603 |
575 |
1.65 |
|
|
|
| 29 |
A' |
545 |
520 |
0.17 |
|
|
|
| 30 |
A' |
389 |
370 |
1.35 |
|
|
|
| 31 |
A" |
3094 |
2949 |
8.22 |
|
|
|
| 32 |
A" |
1157 |
1103 |
1.74 |
|
|
|
| 33 |
A" |
989 |
943 |
0.13 |
|
|
|
| 34 |
A" |
966 |
921 |
1.04 |
|
|
|
| 35 |
A" |
962 |
916 |
1.87 |
|
|
|
| 36 |
A" |
938 |
894 |
3.88 |
|
|
|
| 37 |
A" |
879 |
838 |
0.36 |
|
|
|
| 38 |
A" |
791 |
754 |
53.46 |
|
|
|
| 39 |
A" |
739 |
704 |
16.73 |
|
|
|
| 40 |
A" |
712 |
679 |
14.56 |
|
|
|
| 41 |
A" |
565 |
538 |
4.36 |
|
|
|
| 42 |
A" |
428 |
408 |
6.83 |
|
|
|
| 43 |
A" |
396 |
377 |
4.93 |
|
|
|
| 44 |
A" |
213 |
203 |
3.46 |
|
|
|
| 45 |
A" |
198 |
188 |
0.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31170.8 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 29708.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.179 |
|
|
|
| 2 |
C |
-0.175 |
|
|
|
| 3 |
C |
-0.219 |
|
|
|
| 4 |
C |
-0.213 |
|
|
|
| 5 |
C |
0.071 |
|
|
|
| 6 |
C |
0.082 |
|
|
|
| 7 |
C |
-0.157 |
|
|
|
| 8 |
C |
-0.183 |
|
|
|
| 9 |
C |
-0.435 |
|
|
|
| 10 |
H |
0.165 |
|
|
|
| 11 |
H |
0.166 |
|
|
|
| 12 |
H |
0.166 |
|
|
|
| 13 |
H |
0.164 |
|
|
|
| 14 |
H |
0.170 |
|
|
|
| 15 |
H |
0.171 |
|
|
|
| 16 |
H |
0.204 |
|
|
|
| 17 |
H |
0.204 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.425 |
0.553 |
0.000 |
0.698 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.760 |
0.610 |
0.000 |
| y |
0.610 |
-45.610 |
0.000 |
| z |
0.000 |
0.000 |
-55.912 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.001 |
0.610 |
0.000 |
| y |
0.610 |
5.726 |
0.000 |
| z |
0.000 |
0.000 |
-9.726 |
|
| Polar |
|---|
| 3z2-r2 | -19.453 |
|---|
| x2-y2 | -1.150 |
|---|
| xy | 0.610 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
17.639 |
1.644 |
0.000 |
| y |
1.644 |
15.122 |
0.000 |
| z |
0.000 |
0.000 |
5.325 |
<r2> (average value of r
2) Å
2
| <r2> |
292.084 |
| (<r2>)1/2 |
17.090 |