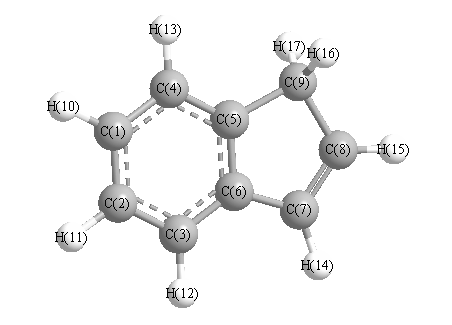
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3258 |
3110 |
14.04 |
|
|
|
| 2 |
A' |
3234 |
3086 |
4.51 |
|
|
|
| 3 |
A' |
3230 |
3082 |
22.34 |
|
|
|
| 4 |
A' |
3217 |
3070 |
26.40 |
|
|
|
| 5 |
A' |
3206 |
3059 |
2.05 |
|
|
|
| 6 |
A' |
3199 |
3053 |
4.45 |
|
|
|
| 7 |
A' |
3054 |
2915 |
11.66 |
|
|
|
| 8 |
A' |
1691 |
1614 |
2.50 |
|
|
|
| 9 |
A' |
1675 |
1599 |
0.41 |
|
|
|
| 10 |
A' |
1639 |
1565 |
0.34 |
|
|
|
| 11 |
A' |
1516 |
1447 |
11.37 |
|
|
|
| 12 |
A' |
1511 |
1442 |
5.63 |
|
|
|
| 13 |
A' |
1439 |
1374 |
9.30 |
|
|
|
| 14 |
A' |
1423 |
1358 |
1.23 |
|
|
|
| 15 |
A' |
1371 |
1309 |
4.36 |
|
|
|
| 16 |
A' |
1329 |
1268 |
0.09 |
|
|
|
| 17 |
A' |
1267 |
1209 |
1.93 |
|
|
|
| 18 |
A' |
1245 |
1188 |
3.09 |
|
|
|
| 19 |
A' |
1194 |
1139 |
1.00 |
|
|
|
| 20 |
A' |
1183 |
1129 |
0.02 |
|
|
|
| 21 |
A' |
1142 |
1090 |
0.49 |
|
|
|
| 22 |
A' |
1095 |
1045 |
1.20 |
|
|
|
| 23 |
A' |
1059 |
1011 |
3.52 |
|
|
|
| 24 |
A' |
977 |
932 |
9.35 |
|
|
|
| 25 |
A' |
876 |
836 |
2.87 |
|
|
|
| 26 |
A' |
848 |
810 |
1.49 |
|
|
|
| 27 |
A' |
748 |
714 |
2.81 |
|
|
|
| 28 |
A' |
604 |
576 |
1.64 |
|
|
|
| 29 |
A' |
545 |
520 |
0.16 |
|
|
|
| 30 |
A' |
389 |
371 |
1.33 |
|
|
|
| 31 |
A" |
3089 |
2948 |
8.11 |
|
|
|
| 32 |
A" |
1146 |
1094 |
1.70 |
|
|
|
| 33 |
A" |
991 |
945 |
0.09 |
|
|
|
| 34 |
A" |
967 |
923 |
1.90 |
|
|
|
| 35 |
A" |
958 |
915 |
0.50 |
|
|
|
| 36 |
A" |
936 |
894 |
4.45 |
|
|
|
| 37 |
A" |
877 |
837 |
0.22 |
|
|
|
| 38 |
A" |
790 |
754 |
50.56 |
|
|
|
| 39 |
A" |
737 |
703 |
15.76 |
|
|
|
| 40 |
A" |
712 |
679 |
13.23 |
|
|
|
| 41 |
A" |
565 |
539 |
4.07 |
|
|
|
| 42 |
A" |
427 |
408 |
6.48 |
|
|
|
| 43 |
A" |
395 |
377 |
5.01 |
|
|
|
| 44 |
A" |
213 |
203 |
3.44 |
|
|
|
| 45 |
A" |
198 |
189 |
0.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31084.1 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 29663.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.132 |
|
|
|
| 2 |
C |
-0.128 |
|
|
|
| 3 |
C |
-0.156 |
|
|
|
| 4 |
C |
-0.149 |
|
|
|
| 5 |
C |
0.032 |
|
|
|
| 6 |
C |
0.059 |
|
|
|
| 7 |
C |
-0.100 |
|
|
|
| 8 |
C |
-0.144 |
|
|
|
| 9 |
C |
-0.321 |
|
|
|
| 10 |
H |
0.118 |
|
|
|
| 11 |
H |
0.119 |
|
|
|
| 12 |
H |
0.119 |
|
|
|
| 13 |
H |
0.116 |
|
|
|
| 14 |
H |
0.122 |
|
|
|
| 15 |
H |
0.124 |
|
|
|
| 16 |
H |
0.161 |
|
|
|
| 17 |
H |
0.161 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.436 |
0.566 |
0.000 |
0.715 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.912 |
0.609 |
0.000 |
| y |
0.609 |
-45.702 |
0.000 |
| z |
0.000 |
0.000 |
-55.801 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.839 |
0.609 |
0.000 |
| y |
0.609 |
5.654 |
0.000 |
| z |
0.000 |
0.000 |
-9.494 |
|
| Polar |
|---|
| 3z2-r2 | -18.987 |
|---|
| x2-y2 | -1.210 |
|---|
| xy | 0.609 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
17.805 |
1.651 |
0.000 |
| y |
1.651 |
15.246 |
0.000 |
| z |
0.000 |
0.000 |
5.400 |
<r2> (average value of r
2) Å
2
| <r2> |
291.992 |
| (<r2>)1/2 |
17.088 |