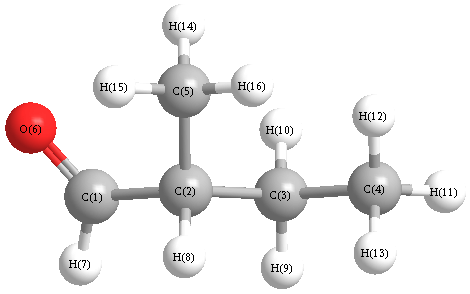
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3176 |
3027 |
15.18 |
|
|
|
| 2 |
A |
3165 |
3017 |
19.96 |
|
|
|
| 3 |
A |
3157 |
3009 |
29.47 |
|
|
|
| 4 |
A |
3149 |
3001 |
32.63 |
|
|
|
| 5 |
A |
3096 |
2951 |
18.94 |
|
|
|
| 6 |
A |
3087 |
2942 |
13.18 |
|
|
|
| 7 |
A |
3072 |
2928 |
22.42 |
|
|
|
| 8 |
A |
3054 |
2911 |
28.61 |
|
|
|
| 9 |
A |
3012 |
2871 |
16.94 |
|
|
|
| 10 |
A |
2911 |
2775 |
123.10 |
|
|
|
| 11 |
A |
1867 |
1780 |
166.95 |
|
|
|
| 12 |
A |
1536 |
1464 |
4.29 |
|
|
|
| 13 |
A |
1529 |
1457 |
10.72 |
|
|
|
| 14 |
A |
1523 |
1452 |
6.53 |
|
|
|
| 15 |
A |
1519 |
1448 |
9.60 |
|
|
|
| 16 |
A |
1509 |
1438 |
0.69 |
|
|
|
| 17 |
A |
1451 |
1383 |
9.65 |
|
|
|
| 18 |
A |
1441 |
1373 |
3.66 |
|
|
|
| 19 |
A |
1428 |
1361 |
4.42 |
|
|
|
| 20 |
A |
1395 |
1329 |
4.46 |
|
|
|
| 21 |
A |
1378 |
1313 |
1.78 |
|
|
|
| 22 |
A |
1324 |
1262 |
2.05 |
|
|
|
| 23 |
A |
1287 |
1226 |
0.37 |
|
|
|
| 24 |
A |
1208 |
1151 |
2.08 |
|
|
|
| 25 |
A |
1185 |
1129 |
3.66 |
|
|
|
| 26 |
A |
1135 |
1082 |
2.96 |
|
|
|
| 27 |
A |
1068 |
1018 |
0.04 |
|
|
|
| 28 |
A |
1032 |
984 |
13.31 |
|
|
|
| 29 |
A |
987 |
941 |
8.77 |
|
|
|
| 30 |
A |
949 |
904 |
9.57 |
|
|
|
| 31 |
A |
923 |
880 |
12.79 |
|
|
|
| 32 |
A |
806 |
768 |
6.49 |
|
|
|
| 33 |
A |
788 |
751 |
3.15 |
|
|
|
| 34 |
A |
663 |
632 |
7.54 |
|
|
|
| 35 |
A |
406 |
387 |
0.94 |
|
|
|
| 36 |
A |
391 |
373 |
0.70 |
|
|
|
| 37 |
A |
290 |
277 |
5.37 |
|
|
|
| 38 |
A |
267 |
254 |
0.41 |
|
|
|
| 39 |
A |
231 |
220 |
2.73 |
|
|
|
| 40 |
A |
194 |
185 |
0.54 |
|
|
|
| 41 |
A |
94 |
90 |
2.34 |
|
|
|
| 42 |
A |
75 |
71 |
3.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31377.2 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 29905.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.246 |
|
|
|
| 2 |
C |
-0.226 |
|
|
|
| 3 |
C |
-0.331 |
|
|
|
| 4 |
C |
-0.523 |
|
|
|
| 5 |
C |
-0.519 |
|
|
|
| 6 |
O |
-0.387 |
|
|
|
| 7 |
H |
0.126 |
|
|
|
| 8 |
H |
0.183 |
|
|
|
| 9 |
H |
0.170 |
|
|
|
| 10 |
H |
0.179 |
|
|
|
| 11 |
H |
0.180 |
|
|
|
| 12 |
H |
0.180 |
|
|
|
| 13 |
H |
0.173 |
|
|
|
| 14 |
H |
0.183 |
|
|
|
| 15 |
H |
0.193 |
|
|
|
| 16 |
H |
0.173 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.562 |
-0.458 |
0.767 |
2.713 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.383 |
-0.031 |
-1.702 |
| y |
-0.031 |
-36.748 |
-0.278 |
| z |
-1.702 |
-0.278 |
-36.977 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.521 |
-0.031 |
-1.702 |
| y |
-0.031 |
4.432 |
-0.278 |
| z |
-1.702 |
-0.278 |
4.089 |
|
| Polar |
|---|
| 3z2-r2 | 8.178 |
|---|
| x2-y2 | -8.635 |
|---|
| xy | -0.031 |
|---|
| xz | -1.702 |
|---|
| yz | -0.278 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.433 |
-0.235 |
0.273 |
| y |
-0.235 |
8.081 |
-0.244 |
| z |
0.273 |
-0.244 |
6.816 |
<r2> (average value of r
2) Å
2
| <r2> |
203.115 |
| (<r2>)1/2 |
14.252 |