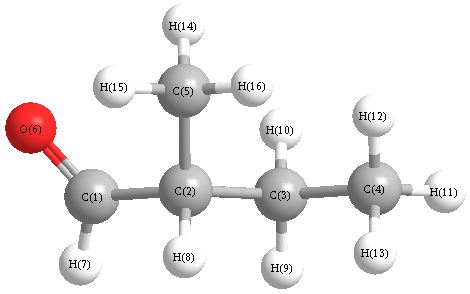
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3172 |
3027 |
14.38 |
|
|
|
| 2 |
A |
3160 |
3016 |
20.40 |
|
|
|
| 3 |
A |
3153 |
3009 |
28.32 |
|
|
|
| 4 |
A |
3145 |
3001 |
32.42 |
|
|
|
| 5 |
A |
3092 |
2951 |
19.04 |
|
|
|
| 6 |
A |
3078 |
2938 |
13.86 |
|
|
|
| 7 |
A |
3065 |
2925 |
23.13 |
|
|
|
| 8 |
A |
3047 |
2908 |
28.17 |
|
|
|
| 9 |
A |
3005 |
2868 |
16.40 |
|
|
|
| 10 |
A |
2895 |
2763 |
122.38 |
|
|
|
| 11 |
A |
1863 |
1778 |
166.20 |
|
|
|
| 12 |
A |
1518 |
1449 |
4.04 |
|
|
|
| 13 |
A |
1511 |
1442 |
9.89 |
|
|
|
| 14 |
A |
1506 |
1437 |
6.13 |
|
|
|
| 15 |
A |
1501 |
1432 |
8.85 |
|
|
|
| 16 |
A |
1491 |
1423 |
0.95 |
|
|
|
| 17 |
A |
1440 |
1374 |
8.38 |
|
|
|
| 18 |
A |
1426 |
1361 |
3.82 |
|
|
|
| 19 |
A |
1415 |
1351 |
4.89 |
|
|
|
| 20 |
A |
1382 |
1319 |
4.03 |
|
|
|
| 21 |
A |
1367 |
1304 |
1.98 |
|
|
|
| 22 |
A |
1313 |
1253 |
1.98 |
|
|
|
| 23 |
A |
1276 |
1218 |
0.32 |
|
|
|
| 24 |
A |
1200 |
1145 |
2.09 |
|
|
|
| 25 |
A |
1179 |
1125 |
3.81 |
|
|
|
| 26 |
A |
1129 |
1078 |
2.48 |
|
|
|
| 27 |
A |
1062 |
1013 |
0.03 |
|
|
|
| 28 |
A |
1026 |
979 |
13.94 |
|
|
|
| 29 |
A |
981 |
936 |
8.50 |
|
|
|
| 30 |
A |
945 |
902 |
9.92 |
|
|
|
| 31 |
A |
918 |
876 |
12.34 |
|
|
|
| 32 |
A |
803 |
766 |
6.89 |
|
|
|
| 33 |
A |
784 |
748 |
3.21 |
|
|
|
| 34 |
A |
659 |
629 |
7.30 |
|
|
|
| 35 |
A |
406 |
387 |
0.92 |
|
|
|
| 36 |
A |
392 |
374 |
0.75 |
|
|
|
| 37 |
A |
288 |
275 |
5.59 |
|
|
|
| 38 |
A |
266 |
254 |
0.41 |
|
|
|
| 39 |
A |
230 |
219 |
2.66 |
|
|
|
| 40 |
A |
193 |
184 |
0.55 |
|
|
|
| 41 |
A |
94 |
90 |
2.16 |
|
|
|
| 42 |
A |
75 |
72 |
3.52 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31224.9 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 29797.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.293 |
|
|
|
| 2 |
C |
-0.195 |
|
|
|
| 3 |
C |
-0.246 |
|
|
|
| 4 |
C |
-0.391 |
|
|
|
| 5 |
C |
-0.377 |
|
|
|
| 6 |
O |
-0.397 |
|
|
|
| 7 |
H |
0.084 |
|
|
|
| 8 |
H |
0.143 |
|
|
|
| 9 |
H |
0.127 |
|
|
|
| 10 |
H |
0.137 |
|
|
|
| 11 |
H |
0.135 |
|
|
|
| 12 |
H |
0.137 |
|
|
|
| 13 |
H |
0.131 |
|
|
|
| 14 |
H |
0.141 |
|
|
|
| 15 |
H |
0.149 |
|
|
|
| 16 |
H |
0.127 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.572 |
-0.456 |
0.732 |
2.712 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-45.379 |
-0.009 |
-1.620 |
| y |
-0.009 |
-36.728 |
-0.262 |
| z |
-1.620 |
-0.262 |
-36.990 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.520 |
-0.009 |
-1.620 |
| y |
-0.009 |
4.456 |
-0.262 |
| z |
-1.620 |
-0.262 |
4.064 |
|
| Polar |
|---|
| 3z2-r2 | 8.128 |
|---|
| x2-y2 | -8.651 |
|---|
| xy | -0.009 |
|---|
| xz | -1.620 |
|---|
| yz | -0.262 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.582 |
-0.242 |
0.259 |
| y |
-0.242 |
8.219 |
-0.232 |
| z |
0.259 |
-0.232 |
6.900 |
<r2> (average value of r
2) Å
2
| <r2> |
203.217 |
| (<r2>)1/2 |
14.255 |