return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10O (3-Pentanone)

using model chemistry: QCISD/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD/6-31G**
 hartrees
Energy at 0K-271.000614
Energy at 298.15K 
HF Energy-270.047662
Nuclear repulsion energy238.887428
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3201 3014 34.25      
2 A1 3120 2937 1.46      
3 A1 3098 2916 29.55      
4 A1 1816 1710 87.75      
5 A1 1556 1465 5.38      
6 A1 1527 1438 8.57      
7 A1 1474 1387 7.20      
8 A1 1406 1323 3.73      
9 A1 1151 1083 1.28      
10 A1 1048 987 2.09      
11 A1 805 758 0.73      
12 A1 417 392 0.40      
13 A1 199 188 0.43      
14 A2 3211 3023 0.00      
15 A2 3127 2944 0.00      
16 A2 1553 1462 0.00      
17 A2 1305 1228 0.00      
18 A2 1034 974 0.00      
19 A2 740 696 0.00      
20 A2 226 213 0.00      
21 A2 25 24 0.00      
22 B1 3211 3023 49.28      
23 B1 3137 2953 24.85      
24 B1 1553 1462 8.43      
25 B1 1341 1263 1.50      
26 B1 1178 1109 0.06      
27 B1 840 790 6.69      
28 B1 468 440 0.16      
29 B1 207 195 0.32      
30 B1 64 60 0.20      
31 B2 3201 3013 16.61      
32 B2 3120 2937 42.44      
33 B2 3090 2909 8.26      
34 B2 1556 1465 8.16      
35 B2 1517 1428 1.18      
36 B2 1474 1387 0.79      
37 B2 1446 1361 26.66      
38 B2 1180 1111 59.42      
39 B2 1050 988 9.42      
40 B2 998 940 12.72      
41 B2 636 599 6.89      
42 B2 319 300 12.95      

Unscaled Zero Point Vibrational Energy (zpe) 31811.4 cm-1
Scaled (by 0.9414) Zero Point Vibrational Energy (zpe) 29947.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD/6-31G**
ABC
0.29638 0.06527 0.05567

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD/6-31G**

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
O1 0.000 0.000 1.285
C2 0.000 0.000 0.061
C3 0.000 1.295 -0.739
C4 0.000 -1.295 -0.739
C5 0.000 2.541 0.138
C6 0.000 -2.541 0.138
H7 0.872 1.280 -1.401
H8 -0.872 1.280 -1.401
H9 -0.872 -1.280 -1.401
H10 0.872 -1.280 -1.401
H11 0.000 3.442 -0.477
H12 -0.878 2.560 0.781
H13 0.878 2.560 0.781
H14 0.000 -3.442 -0.477
H15 0.878 -2.560 0.781
H16 -0.878 -2.560 0.781

Atom - Atom Distances (Å)
  O1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
O11.22342.40232.40232.78802.78803.10043.10043.10043.10043.86632.75242.75243.86632.75242.7524
C21.22341.52211.52212.54242.54242.13002.13002.13002.13003.48352.80022.80023.48352.80022.8002
C32.40231.52212.58981.52393.93511.09511.09512.79762.79762.16262.16382.16384.74374.23554.2355
C42.40231.52212.58983.93511.52392.79762.79761.09511.09514.74374.23554.23552.16262.16382.1638
C52.78802.54241.52393.93515.08252.17292.17294.21054.21051.09031.08911.08916.01445.21575.2157
C62.78802.54243.93511.52395.08254.21054.21052.17292.17296.01445.21575.21571.09031.08911.0891
H73.10042.13001.09512.79762.17294.21051.74343.09652.55912.50783.07672.53054.88914.41634.7504
H83.10042.13001.09512.79762.17294.21051.74342.55913.09652.50782.53053.07674.88914.75044.4163
H93.10042.13002.79761.09514.21052.17293.09652.55911.74344.88914.41634.75042.50783.07672.5305
H103.10042.13002.79761.09514.21052.17292.55913.09651.74344.88914.75044.41632.50782.53053.0767
H113.86633.48352.16264.74371.09036.01442.50782.50784.88914.88911.77021.77026.88326.19436.1943
H122.75242.80022.16384.23551.08915.21573.07672.53054.41634.75041.77021.75686.19435.41215.1190
H132.75242.80022.16384.23551.08915.21572.53053.07674.75044.41631.77021.75686.19435.11905.4121
H143.86633.48354.74372.16266.01441.09034.88914.88912.50782.50786.88326.19436.19431.77021.7702
H152.75242.80024.23552.16385.21571.08914.41634.75043.07672.53056.19435.41215.11901.77021.7568
H162.75242.80024.23552.16385.21571.08914.75044.41632.53053.07676.19435.11905.41211.77021.7568

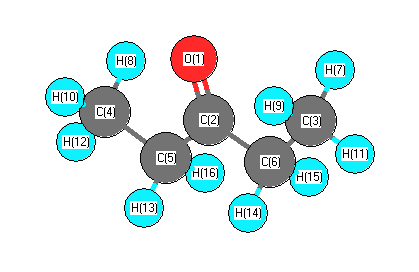
picture of 3-Pentanone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
O1 C2 C3 121.712 O1 C2 C4 121.712
C2 C3 C5 113.164 C2 C3 H7 107.829
C2 C3 H8 107.829 C2 C4 C6 113.164
C2 C4 H9 107.829 C2 C4 H10 107.829
C3 C2 C4 116.575 C3 C5 H11 110.541
C3 C5 H12 110.712 C3 C5 H13 110.712
C4 C6 H14 110.541 C4 C6 H15 110.712
C4 C6 H16 110.712 C5 C3 H7 111.076
C5 C3 H8 111.076 C6 C4 H9 111.076
C6 C4 H10 111.076 H7 C3 H8 105.499
H9 C4 H10 105.499 H11 C5 H12 108.630
H11 C5 H13 108.630 H12 C5 H13 107.525
H14 C6 H15 108.630 H14 C6 H16 108.630
H15 C6 H16 107.525
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability