Jump to
S1C2
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -342.962588 |
| Energy at 298.15K | -342.967466 |
| HF Energy | -342.962588 |
| Nuclear repulsion energy | 272.220637 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3326 |
3170 |
0.20 |
142.01 |
0.13 |
0.23 |
| 2 |
A' |
3305 |
3150 |
0.11 |
56.06 |
0.38 |
0.55 |
| 3 |
A' |
3293 |
3138 |
1.02 |
78.14 |
0.68 |
0.81 |
| 4 |
A' |
2955 |
2816 |
110.37 |
123.40 |
0.32 |
0.49 |
| 5 |
A' |
1831 |
1745 |
312.99 |
124.25 |
0.32 |
0.49 |
| 6 |
A' |
1644 |
1567 |
7.89 |
8.00 |
0.45 |
0.62 |
| 7 |
A' |
1542 |
1470 |
51.30 |
95.05 |
0.33 |
0.50 |
| 8 |
A' |
1466 |
1397 |
40.18 |
39.83 |
0.42 |
0.59 |
| 9 |
A' |
1417 |
1350 |
0.26 |
13.38 |
0.11 |
0.21 |
| 10 |
A' |
1339 |
1276 |
45.29 |
6.62 |
0.34 |
0.50 |
| 11 |
A' |
1272 |
1212 |
3.91 |
9.98 |
0.50 |
0.66 |
| 12 |
A' |
1228 |
1171 |
6.76 |
3.18 |
0.12 |
0.22 |
| 13 |
A' |
1134 |
1081 |
11.02 |
10.45 |
0.35 |
0.52 |
| 14 |
A' |
1058 |
1008 |
41.40 |
3.51 |
0.36 |
0.53 |
| 15 |
A' |
968 |
923 |
18.44 |
5.91 |
0.12 |
0.21 |
| 16 |
A' |
903 |
860 |
11.16 |
7.17 |
0.74 |
0.85 |
| 17 |
A' |
776 |
740 |
64.00 |
1.51 |
0.67 |
0.80 |
| 18 |
A' |
506 |
482 |
1.12 |
7.69 |
0.33 |
0.49 |
| 19 |
A' |
199 |
190 |
5.55 |
0.92 |
0.67 |
0.80 |
| 20 |
A" |
1016 |
968 |
0.00 |
6.13 |
0.75 |
0.86 |
| 21 |
A" |
900 |
858 |
1.17 |
1.16 |
0.75 |
0.86 |
| 22 |
A" |
845 |
806 |
5.79 |
0.24 |
0.75 |
0.86 |
| 23 |
A" |
785 |
748 |
59.13 |
0.68 |
0.75 |
0.86 |
| 24 |
A" |
662 |
631 |
1.75 |
3.10 |
0.75 |
0.86 |
| 25 |
A" |
613 |
584 |
7.24 |
0.19 |
0.75 |
0.86 |
| 26 |
A" |
294 |
280 |
11.47 |
0.62 |
0.75 |
0.86 |
| 27 |
A" |
138 |
132 |
1.40 |
1.72 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17705.9 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 16875.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.504 |
-0.894 |
0.000 |
| C2 |
-0.544 |
-1.734 |
0.000 |
| C3 |
-1.726 |
-1.051 |
0.000 |
| C4 |
0.000 |
0.368 |
0.000 |
| C5 |
-1.371 |
0.323 |
0.000 |
| C6 |
0.914 |
1.497 |
0.000 |
| O7 |
2.124 |
1.417 |
0.000 |
| H8 |
-0.301 |
-2.786 |
0.000 |
| H9 |
-2.718 |
-1.479 |
0.000 |
| H10 |
-2.036 |
1.176 |
0.000 |
| H11 |
0.391 |
2.478 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3423 | 2.2350 | 1.3592 | 2.2348 | 2.4262 | 2.8226 | 2.0563 | 3.2745 | 3.2763 | 3.3736 |
C2 | 1.3423 | | 1.3652 | 2.1711 | 2.2164 | 3.5444 | 4.1284 | 1.0803 | 2.1894 | 3.2699 | 4.3137 | C3 | 2.2350 | 1.3652 | | 2.2342 | 1.4184 | 3.6688 | 4.5728 | 2.2455 | 1.0808 | 2.2481 | 4.1145 | C4 | 1.3592 | 2.1711 | 2.2342 | | 1.3716 | 1.4525 | 2.3688 | 3.1690 | 3.2863 | 2.1902 | 2.1452 | C5 | 2.2348 | 2.2164 | 1.4184 | 1.3716 | | 2.5691 | 3.6621 | 3.2879 | 2.2495 | 1.0818 | 2.7835 | C6 | 2.4262 | 3.5444 | 3.6688 | 1.4525 | 2.5691 | | 1.2124 | 4.4525 | 4.6956 | 2.9673 | 1.1113 | O7 | 2.8226 | 4.1284 | 4.5728 | 2.3688 | 3.6621 | 1.2124 | | 4.8529 | 5.6420 | 4.1667 | 2.0316 | H8 | 2.0563 | 1.0803 | 2.2455 | 3.1690 | 3.2879 | 4.4525 | 4.8529 | | 2.7481 | 4.3254 | 5.3093 | H9 | 3.2745 | 2.1894 | 1.0808 | 3.2863 | 2.2495 | 4.6956 | 5.6420 | 2.7481 | | 2.7410 | 5.0319 | H10 | 3.2763 | 3.2699 | 2.2481 | 2.1902 | 1.0818 | 2.9673 | 4.1667 | 4.3254 | 2.7410 | | 2.7539 | H11 | 3.3736 | 4.3137 | 4.1145 | 2.1452 | 2.7835 | 1.1113 | 2.0316 | 5.3093 | 5.0319 | 2.7539 | |
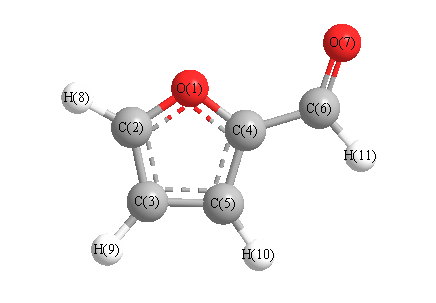 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.268 |
|
O1 |
C2 |
H8 |
115.740 |
| O1 |
C4 |
C5 |
109.847 |
|
O1 |
C4 |
C6 |
119.248 |
| C2 |
O1 |
C4 |
106.962 |
|
C2 |
C3 |
C5 |
105.524 |
| C2 |
C3 |
H9 |
126.648 |
|
C3 |
C2 |
H8 |
132.991 |
| C3 |
C5 |
C4 |
106.398 |
|
C3 |
C5 |
H10 |
127.576 |
| C4 |
C5 |
H10 |
126.026 |
|
C4 |
C6 |
O7 |
125.220 |
| C4 |
C6 |
H11 |
112.922 |
|
C5 |
C3 |
H9 |
127.827 |
| C5 |
C4 |
C6 |
130.905 |
|
O7 |
C6 |
H11 |
121.857 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.413 |
|
|
|
| 2 |
C |
0.089 |
|
|
|
| 3 |
C |
-0.238 |
|
|
|
| 4 |
C |
0.264 |
|
|
|
| 5 |
C |
-0.213 |
|
|
|
| 6 |
C |
0.179 |
|
|
|
| 7 |
O |
-0.408 |
|
|
|
| 8 |
H |
0.207 |
|
|
|
| 9 |
H |
0.195 |
|
|
|
| 10 |
H |
0.200 |
|
|
|
| 11 |
H |
0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.964 |
-1.151 |
0.000 |
4.128 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.053 |
-2.773 |
0.000 |
| y |
-2.773 |
-33.055 |
0.000 |
| z |
0.000 |
0.000 |
-40.469 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.291 |
-2.773 |
0.000 |
| y |
-2.773 |
8.706 |
0.000 |
| z |
0.000 |
0.000 |
-2.415 |
|
| Polar |
|---|
| 3z2-r2 | -4.829 |
|---|
| x2-y2 | -9.998 |
|---|
| xy | -2.773 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.441 |
1.901 |
0.000 |
| y |
1.901 |
10.236 |
0.000 |
| z |
0.000 |
0.000 |
3.331 |
<r2> (average value of r
2) Å
2
| <r2> |
189.572 |
| (<r2>)1/2 |
13.769 |
Jump to
S1C1
Energy calculated at PBE1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -342.964118 |
| Energy at 298.15K | -342.968979 |
| HF Energy | -342.964118 |
| Nuclear repulsion energy | 271.391533 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3326 |
3170 |
0.36 |
152.86 |
0.13 |
0.23 |
| 2 |
A' |
3314 |
3159 |
0.47 |
28.48 |
0.35 |
0.52 |
| 3 |
A' |
3298 |
3143 |
0.73 |
86.85 |
0.72 |
0.83 |
| 4 |
A' |
2975 |
2835 |
104.13 |
133.83 |
0.32 |
0.49 |
| 5 |
A' |
1825 |
1739 |
261.39 |
96.48 |
0.35 |
0.51 |
| 6 |
A' |
1651 |
1574 |
54.03 |
22.07 |
0.36 |
0.53 |
| 7 |
A' |
1534 |
1462 |
117.63 |
160.50 |
0.31 |
0.48 |
| 8 |
A' |
1475 |
1406 |
2.88 |
11.00 |
0.71 |
0.83 |
| 9 |
A' |
1425 |
1358 |
13.64 |
20.67 |
0.39 |
0.56 |
| 10 |
A' |
1290 |
1229 |
29.43 |
4.93 |
0.31 |
0.47 |
| 11 |
A' |
1277 |
1217 |
0.44 |
4.11 |
0.53 |
0.69 |
| 12 |
A' |
1212 |
1156 |
12.88 |
2.80 |
0.13 |
0.23 |
| 13 |
A' |
1139 |
1086 |
23.11 |
13.79 |
0.32 |
0.48 |
| 14 |
A' |
1050 |
1001 |
39.26 |
1.83 |
0.22 |
0.36 |
| 15 |
A' |
982 |
936 |
6.69 |
6.27 |
0.16 |
0.28 |
| 16 |
A' |
902 |
860 |
7.45 |
6.25 |
0.75 |
0.86 |
| 17 |
A' |
762 |
727 |
72.09 |
2.84 |
0.61 |
0.76 |
| 18 |
A' |
503 |
479 |
1.07 |
6.23 |
0.37 |
0.54 |
| 19 |
A' |
202 |
192 |
6.68 |
0.30 |
0.42 |
0.59 |
| 20 |
A" |
1024 |
976 |
0.10 |
7.43 |
0.75 |
0.86 |
| 21 |
A" |
908 |
866 |
1.60 |
1.52 |
0.75 |
0.86 |
| 22 |
A" |
852 |
812 |
7.70 |
0.45 |
0.75 |
0.86 |
| 23 |
A" |
783 |
746 |
54.50 |
0.68 |
0.75 |
0.86 |
| 24 |
A" |
652 |
621 |
1.75 |
1.45 |
0.75 |
0.86 |
| 25 |
A" |
614 |
585 |
10.11 |
0.49 |
0.75 |
0.86 |
| 26 |
A" |
254 |
242 |
12.51 |
1.63 |
0.75 |
0.86 |
| 27 |
A" |
162 |
155 |
2.63 |
1.29 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17694.7 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 16864.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBE1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.250 |
-0.258 |
0.000 |
| C2 |
1.094 |
-1.593 |
0.000 |
| C3 |
-0.229 |
-1.932 |
0.000 |
| C4 |
0.000 |
0.287 |
0.000 |
| C5 |
-0.942 |
-0.707 |
0.000 |
| C6 |
-0.096 |
1.737 |
0.000 |
| O7 |
-1.151 |
2.338 |
0.000 |
| H8 |
2.005 |
-2.173 |
0.000 |
| H9 |
-0.635 |
-2.933 |
0.000 |
| H10 |
-2.012 |
-0.553 |
0.000 |
| H11 |
0.882 |
2.261 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3432 | 2.2330 | 1.3642 | 2.2379 | 2.4069 | 3.5366 | 2.0584 | 3.2723 | 3.2757 | 2.5462 |
C2 | 1.3432 | | 1.3659 | 2.1754 | 2.2212 | 3.5359 | 4.5269 | 1.0802 | 2.1881 | 3.2759 | 3.8595 | C3 | 2.2330 | 1.3659 | | 2.2308 | 1.4177 | 3.6709 | 4.3683 | 2.2470 | 1.0810 | 2.2540 | 4.3374 | C4 | 1.3642 | 2.1754 | 2.2308 | | 1.3698 | 1.4526 | 2.3516 | 3.1744 | 3.2828 | 2.1808 | 2.1619 | C5 | 2.2379 | 2.2212 | 1.4177 | 1.3698 | | 2.5860 | 3.0520 | 3.2924 | 2.2480 | 1.0808 | 3.4838 | C6 | 2.4069 | 3.5359 | 3.6709 | 1.4526 | 2.5860 | | 1.2143 | 4.4391 | 4.7013 | 2.9862 | 1.1099 | O7 | 3.5366 | 4.5269 | 4.3683 | 2.3516 | 3.0520 | 1.2143 | | 5.5059 | 5.2968 | 3.0171 | 2.0347 | H8 | 2.0584 | 1.0802 | 2.2470 | 3.1744 | 3.2924 | 4.4391 | 5.5059 | | 2.7472 | 4.3318 | 4.5744 | H9 | 3.2723 | 2.1881 | 1.0810 | 3.2828 | 2.2480 | 4.7013 | 5.2968 | 2.7472 | | 2.7500 | 5.4115 | H10 | 3.2757 | 3.2759 | 2.2540 | 2.1808 | 1.0808 | 2.9862 | 3.0171 | 4.3318 | 2.7500 | | 4.0373 | H11 | 2.5462 | 3.8595 | 4.3374 | 2.1619 | 3.4838 | 1.1099 | 2.0347 | 4.5744 | 5.4115 | 4.0373 | |
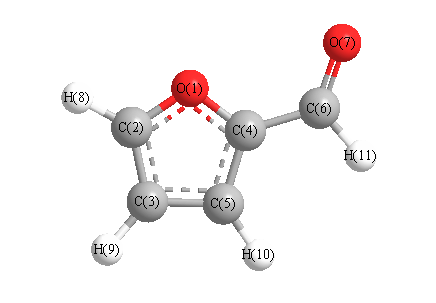 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.027 |
|
O1 |
C2 |
H8 |
115.866 |
| O1 |
C4 |
C5 |
109.880 |
|
O1 |
C4 |
C6 |
117.376 |
| C2 |
O1 |
C4 |
106.933 |
|
C2 |
C3 |
C5 |
105.855 |
| C2 |
C3 |
H9 |
126.432 |
|
C3 |
C2 |
H8 |
133.107 |
| C3 |
C5 |
C4 |
106.305 |
|
C3 |
C5 |
H10 |
128.380 |
| C4 |
C5 |
H10 |
125.315 |
|
C4 |
C6 |
O7 |
123.470 |
| C4 |
C6 |
H11 |
114.394 |
|
C5 |
C3 |
H9 |
127.713 |
| C5 |
C4 |
C6 |
132.744 |
|
O7 |
C6 |
H11 |
122.136 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.433 |
|
|
|
| 2 |
C |
0.083 |
|
|
|
| 3 |
C |
-0.236 |
|
|
|
| 4 |
C |
0.256 |
|
|
|
| 5 |
C |
-0.193 |
|
|
|
| 6 |
C |
0.175 |
|
|
|
| 7 |
O |
-0.412 |
|
|
|
| 8 |
H |
0.206 |
|
|
|
| 9 |
H |
0.195 |
|
|
|
| 10 |
H |
0.212 |
|
|
|
| 11 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.512 |
-3.238 |
0.000 |
3.574 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.894 |
3.765 |
0.000 |
| y |
3.765 |
-39.084 |
0.000 |
| z |
0.000 |
0.000 |
-40.436 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.866 |
3.765 |
0.000 |
| y |
3.765 |
0.081 |
0.000 |
| z |
0.000 |
0.000 |
-1.947 |
|
| Polar |
|---|
| 3z2-r2 | -3.894 |
|---|
| x2-y2 | 1.190 |
|---|
| xy | 3.765 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.595 |
-1.133 |
0.000 |
| y |
-1.133 |
12.042 |
0.000 |
| z |
0.000 |
0.000 |
3.330 |
<r2> (average value of r
2) Å
2
| <r2> |
191.897 |
| (<r2>)1/2 |
13.853 |
