Jump to
S1C2
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -342.997961 |
| Energy at 298.15K | -343.002837 |
| HF Energy | -342.997961 |
| Nuclear repulsion energy | 272.228833 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3317 |
3166 |
0.10 |
150.11 |
0.12 |
0.21 |
| 2 |
A' |
3297 |
3147 |
0.12 |
50.83 |
0.44 |
0.61 |
| 3 |
A' |
3286 |
3136 |
1.02 |
80.68 |
0.67 |
0.80 |
| 4 |
A' |
2940 |
2805 |
108.54 |
125.03 |
0.32 |
0.48 |
| 5 |
A' |
1825 |
1742 |
311.51 |
123.31 |
0.32 |
0.49 |
| 6 |
A' |
1641 |
1566 |
7.97 |
7.69 |
0.45 |
0.62 |
| 7 |
A' |
1539 |
1469 |
51.39 |
97.43 |
0.33 |
0.50 |
| 8 |
A' |
1461 |
1395 |
39.33 |
38.02 |
0.43 |
0.60 |
| 9 |
A' |
1411 |
1346 |
0.24 |
13.38 |
0.12 |
0.21 |
| 10 |
A' |
1335 |
1274 |
45.65 |
6.89 |
0.34 |
0.51 |
| 11 |
A' |
1266 |
1208 |
3.27 |
9.92 |
0.49 |
0.66 |
| 12 |
A' |
1224 |
1168 |
6.57 |
3.21 |
0.12 |
0.22 |
| 13 |
A' |
1130 |
1078 |
10.66 |
10.79 |
0.34 |
0.51 |
| 14 |
A' |
1054 |
1006 |
41.51 |
3.56 |
0.36 |
0.53 |
| 15 |
A' |
966 |
922 |
18.74 |
6.05 |
0.12 |
0.22 |
| 16 |
A' |
902 |
861 |
10.92 |
7.08 |
0.74 |
0.85 |
| 17 |
A' |
775 |
739 |
64.23 |
1.52 |
0.66 |
0.79 |
| 18 |
A' |
506 |
483 |
1.13 |
7.83 |
0.32 |
0.49 |
| 19 |
A' |
199 |
190 |
5.58 |
0.94 |
0.67 |
0.80 |
| 20 |
A" |
1015 |
969 |
0.00 |
6.06 |
0.75 |
0.86 |
| 21 |
A" |
904 |
862 |
1.02 |
1.16 |
0.75 |
0.86 |
| 22 |
A" |
848 |
809 |
5.11 |
0.31 |
0.75 |
0.86 |
| 23 |
A" |
785 |
750 |
57.18 |
0.80 |
0.75 |
0.86 |
| 24 |
A" |
661 |
631 |
1.67 |
3.12 |
0.75 |
0.86 |
| 25 |
A" |
613 |
585 |
7.00 |
0.20 |
0.75 |
0.86 |
| 26 |
A" |
294 |
281 |
11.35 |
0.61 |
0.75 |
0.86 |
| 27 |
A" |
138 |
132 |
1.40 |
1.75 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17666.5 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 16859.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.505 |
-0.894 |
0.000 |
| C2 |
-0.542 |
-1.735 |
0.000 |
| C3 |
-1.724 |
-1.052 |
0.000 |
| C4 |
0.000 |
0.368 |
0.000 |
| C5 |
-1.371 |
0.321 |
0.000 |
| C6 |
0.911 |
1.499 |
0.000 |
| O7 |
2.122 |
1.421 |
0.000 |
| H8 |
-0.301 |
-2.787 |
0.000 |
| H9 |
-2.715 |
-1.482 |
0.000 |
| H10 |
-2.035 |
1.174 |
0.000 |
| H11 |
0.384 |
2.477 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3430 | 2.2349 | 1.3596 | 2.2349 | 2.4264 | 2.8227 | 2.0582 | 3.2734 | 3.2754 | 3.3727 |
C2 | 1.3430 | | 1.3652 | 2.1719 | 2.2168 | 3.5448 | 4.1290 | 1.0796 | 2.1880 | 3.2696 | 4.3122 | C3 | 2.2349 | 1.3652 | | 2.2340 | 1.4181 | 3.6677 | 4.5720 | 2.2438 | 1.0799 | 2.2477 | 4.1109 | C4 | 1.3596 | 2.1719 | 2.2340 | | 1.3715 | 1.4517 | 2.3681 | 3.1702 | 3.2853 | 2.1888 | 2.1430 | C5 | 2.2349 | 2.2168 | 1.4181 | 1.3715 | | 2.5679 | 3.6613 | 3.2873 | 2.2486 | 1.0810 | 2.7797 | C6 | 2.4264 | 3.5448 | 3.6677 | 1.4517 | 2.5679 | | 1.2127 | 4.4541 | 4.6937 | 2.9645 | 1.1113 | O7 | 2.8227 | 4.1290 | 4.5720 | 2.3681 | 3.6613 | 1.2127 | | 4.8554 | 5.6403 | 4.1642 | 2.0333 | H8 | 2.0582 | 1.0796 | 2.2438 | 3.1702 | 3.2873 | 4.4541 | 4.8554 | | 2.7443 | 4.3240 | 5.3086 | H9 | 3.2734 | 2.1880 | 1.0799 | 3.2853 | 2.2486 | 4.6937 | 5.6403 | 2.7443 | | 2.7408 | 5.0272 | H10 | 3.2754 | 3.2696 | 2.2477 | 2.1888 | 1.0810 | 2.9645 | 4.1642 | 4.3240 | 2.7408 | | 2.7481 | H11 | 3.3727 | 4.3122 | 4.1109 | 2.1430 | 2.7797 | 1.1113 | 2.0333 | 5.3086 | 5.0272 | 2.7481 | |
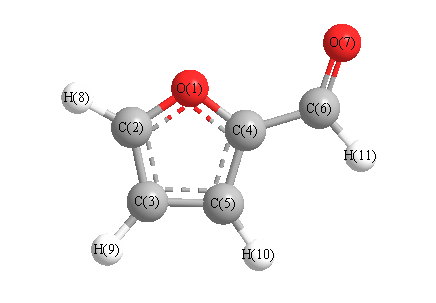 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.225 |
|
O1 |
C2 |
H8 |
115.910 |
| O1 |
C4 |
C5 |
109.836 |
|
O1 |
C4 |
C6 |
119.294 |
| C2 |
O1 |
C4 |
106.962 |
|
C2 |
C3 |
C5 |
105.571 |
| C2 |
C3 |
H9 |
126.585 |
|
C3 |
C2 |
H8 |
132.865 |
| C3 |
C5 |
C4 |
106.406 |
|
C3 |
C5 |
H10 |
127.640 |
| C4 |
C5 |
H10 |
125.954 |
|
C4 |
C6 |
O7 |
125.199 |
| C4 |
C6 |
H11 |
112.792 |
|
C5 |
C3 |
H9 |
127.844 |
| C5 |
C4 |
C6 |
130.870 |
|
O7 |
C6 |
H11 |
122.009 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.418 |
|
|
|
| 2 |
C |
0.138 |
|
|
|
| 3 |
C |
-0.187 |
|
|
|
| 4 |
C |
0.244 |
|
|
|
| 5 |
C |
-0.154 |
|
|
|
| 6 |
C |
0.239 |
|
|
|
| 7 |
O |
-0.417 |
|
|
|
| 8 |
H |
0.161 |
|
|
|
| 9 |
H |
0.147 |
|
|
|
| 10 |
H |
0.152 |
|
|
|
| 11 |
H |
0.095 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.963 |
-1.160 |
0.000 |
4.130 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-43.093 |
-2.781 |
0.000 |
| y |
-2.781 |
-33.113 |
0.000 |
| z |
0.000 |
0.000 |
-40.393 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.340 |
-2.781 |
0.000 |
| y |
-2.781 |
8.630 |
0.000 |
| z |
0.000 |
0.000 |
-2.290 |
|
| Polar |
|---|
| 3z2-r2 | -4.580 |
|---|
| x2-y2 | -9.980 |
|---|
| xy | -2.781 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.498 |
1.912 |
0.000 |
| y |
1.912 |
10.341 |
0.000 |
| z |
0.000 |
0.000 |
3.369 |
<r2> (average value of r
2) Å
2
| <r2> |
189.544 |
| (<r2>)1/2 |
13.767 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-31G**
| | hartrees |
|---|
| Energy at 0K | -342.999490 |
| Energy at 298.15K | -343.004350 |
| HF Energy | -342.999490 |
| Nuclear repulsion energy | 271.405446 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3318 |
3166 |
0.28 |
161.47 |
0.12 |
0.21 |
| 2 |
A' |
3307 |
3156 |
0.66 |
24.12 |
0.46 |
0.63 |
| 3 |
A' |
3291 |
3141 |
0.72 |
88.04 |
0.73 |
0.84 |
| 4 |
A' |
2960 |
2824 |
102.17 |
135.99 |
0.32 |
0.49 |
| 5 |
A' |
1820 |
1736 |
259.62 |
95.88 |
0.35 |
0.51 |
| 6 |
A' |
1647 |
1572 |
54.60 |
21.39 |
0.36 |
0.53 |
| 7 |
A' |
1530 |
1460 |
116.72 |
161.78 |
0.31 |
0.48 |
| 8 |
A' |
1470 |
1403 |
2.36 |
10.35 |
0.72 |
0.84 |
| 9 |
A' |
1419 |
1354 |
13.92 |
20.71 |
0.39 |
0.56 |
| 10 |
A' |
1286 |
1227 |
28.99 |
4.96 |
0.30 |
0.46 |
| 11 |
A' |
1270 |
1212 |
0.48 |
4.02 |
0.55 |
0.71 |
| 12 |
A' |
1208 |
1153 |
12.66 |
2.87 |
0.12 |
0.22 |
| 13 |
A' |
1135 |
1083 |
22.86 |
14.18 |
0.31 |
0.48 |
| 14 |
A' |
1046 |
998 |
39.89 |
1.76 |
0.22 |
0.36 |
| 15 |
A' |
980 |
935 |
6.72 |
6.46 |
0.17 |
0.29 |
| 16 |
A' |
902 |
860 |
7.26 |
6.19 |
0.75 |
0.86 |
| 17 |
A' |
762 |
727 |
72.21 |
2.92 |
0.60 |
0.75 |
| 18 |
A' |
503 |
480 |
1.10 |
6.33 |
0.37 |
0.54 |
| 19 |
A' |
201 |
192 |
6.77 |
0.29 |
0.41 |
0.58 |
| 20 |
A" |
1024 |
977 |
0.08 |
7.34 |
0.75 |
0.86 |
| 21 |
A" |
911 |
870 |
1.43 |
1.50 |
0.75 |
0.86 |
| 22 |
A" |
854 |
815 |
6.91 |
0.56 |
0.75 |
0.86 |
| 23 |
A" |
784 |
748 |
52.80 |
0.79 |
0.75 |
0.86 |
| 24 |
A" |
651 |
622 |
1.66 |
1.46 |
0.75 |
0.86 |
| 25 |
A" |
614 |
586 |
9.91 |
0.50 |
0.75 |
0.86 |
| 26 |
A" |
255 |
243 |
12.39 |
1.64 |
0.75 |
0.86 |
| 27 |
A" |
163 |
155 |
2.63 |
1.33 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17655.2 cm
-1
Scaled (by 0.9543) Zero Point Vibrational Energy (zpe) 16848.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-31G**
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.250 |
-0.259 |
0.000 |
| C2 |
1.093 |
-1.594 |
0.000 |
| C3 |
-0.230 |
-1.931 |
0.000 |
| C4 |
0.000 |
0.288 |
0.000 |
| C5 |
-0.943 |
-0.706 |
0.000 |
| C6 |
-0.094 |
1.737 |
0.000 |
| O7 |
-1.149 |
2.338 |
0.000 |
| H8 |
2.002 |
-2.178 |
0.000 |
| H9 |
-0.636 |
-2.932 |
0.000 |
| H10 |
-2.012 |
-0.551 |
0.000 |
| H11 |
0.886 |
2.258 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3439 | 2.2329 | 1.3646 | 2.2380 | 2.4065 | 3.5363 | 2.0603 | 3.2712 | 3.2748 | 2.5433 |
C2 | 1.3439 | | 1.3658 | 2.1764 | 2.2216 | 3.5361 | 4.5270 | 1.0795 | 2.1867 | 3.2756 | 3.8573 | C3 | 2.2329 | 1.3658 | | 2.2306 | 1.4173 | 3.6700 | 4.3670 | 2.2453 | 1.0801 | 2.2535 | 4.3347 | C4 | 1.3646 | 2.1764 | 2.2306 | | 1.3697 | 1.4518 | 2.3507 | 3.1757 | 3.2818 | 2.1794 | 2.1598 | C5 | 2.2380 | 2.2216 | 1.4173 | 1.3697 | | 2.5855 | 3.0511 | 3.2918 | 2.2470 | 1.0799 | 3.4822 | C6 | 2.4065 | 3.5361 | 3.6700 | 1.4518 | 2.5855 | | 1.2145 | 4.4401 | 4.6996 | 2.9845 | 1.1100 | O7 | 3.5363 | 4.5270 | 4.3670 | 2.3507 | 3.0511 | 1.2145 | | 5.5066 | 5.2949 | 3.0149 | 2.0366 | H8 | 2.0603 | 1.0795 | 2.2453 | 3.1757 | 3.2918 | 4.4401 | 5.5066 | | 2.7435 | 4.3304 | 4.5737 | H9 | 3.2712 | 2.1867 | 1.0801 | 3.2818 | 2.2470 | 4.6996 | 5.2949 | 2.7435 | | 2.7497 | 5.4080 | H10 | 3.2748 | 3.2756 | 2.2535 | 2.1794 | 1.0799 | 2.9845 | 3.0149 | 4.3304 | 2.7497 | | 4.0350 | H11 | 2.5433 | 3.8573 | 4.3347 | 2.1598 | 3.4822 | 1.1100 | 2.0366 | 4.5737 | 5.4080 | 4.0350 | |
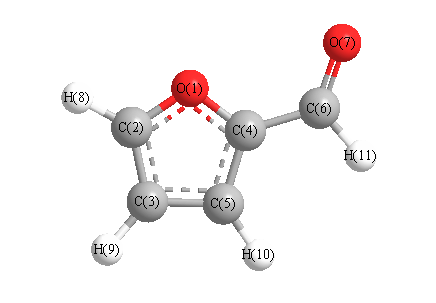 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
110.977 |
|
O1 |
C2 |
H8 |
116.036 |
| O1 |
C4 |
C5 |
109.866 |
|
O1 |
C4 |
C6 |
117.364 |
| C2 |
O1 |
C4 |
106.937 |
|
C2 |
C3 |
C5 |
105.907 |
| C2 |
C3 |
H9 |
126.366 |
|
C3 |
C2 |
H8 |
132.987 |
| C3 |
C5 |
C4 |
106.312 |
|
C3 |
C5 |
H10 |
128.442 |
| C4 |
C5 |
H10 |
125.246 |
|
C4 |
C6 |
O7 |
123.428 |
| C4 |
C6 |
H11 |
114.274 |
|
C5 |
C3 |
H9 |
127.726 |
| C5 |
C4 |
C6 |
132.769 |
|
O7 |
C6 |
H11 |
122.297 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.438 |
|
|
|
| 2 |
C |
0.133 |
|
|
|
| 3 |
C |
-0.186 |
|
|
|
| 4 |
C |
0.234 |
|
|
|
| 5 |
C |
-0.134 |
|
|
|
| 6 |
C |
0.236 |
|
|
|
| 7 |
O |
-0.421 |
|
|
|
| 8 |
H |
0.159 |
|
|
|
| 9 |
H |
0.147 |
|
|
|
| 10 |
H |
0.165 |
|
|
|
| 11 |
H |
0.105 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.506 |
-3.240 |
0.000 |
3.573 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.943 |
3.760 |
0.000 |
| y |
3.760 |
-39.129 |
0.000 |
| z |
0.000 |
0.000 |
-40.362 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.803 |
3.760 |
0.000 |
| y |
3.760 |
0.024 |
0.000 |
| z |
0.000 |
0.000 |
-1.826 |
|
| Polar |
|---|
| 3z2-r2 | -3.653 |
|---|
| x2-y2 | 1.186 |
|---|
| xy | 3.760 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.661 |
-1.140 |
0.000 |
| y |
-1.140 |
12.139 |
0.000 |
| z |
0.000 |
0.000 |
3.369 |
<r2> (average value of r
2) Å
2
| <r2> |
191.854 |
| (<r2>)1/2 |
13.851 |
