return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3COC6H5 (acetophenone)

using model chemistry: MP2=FULL/6-31G**

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31G**
 hartrees
Energy at 0K-383.768500
Energy at 298.15K-383.776665
HF Energy-382.487448
Nuclear repulsion energy401.382343
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G**
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3290 3074 6.22      
2 A' 3285 3069 10.68      
3 A' 3277 3062 14.82      
4 A' 3267 3053 4.78      
5 A' 3266 3051 10.79      
6 A' 3257 3044 0.20      
7 A' 3135 2929 1.96      
8 A' 1763 1648 85.77      
9 A' 1682 1572 9.73      
10 A' 1661 1552 7.29      
11 A' 1557 1455 1.67      
12 A' 1532 1432 8.19      
13 A' 1511 1412 17.79      
14 A' 1477 1380 2.42      
15 A' 1435 1341 50.40      
16 A' 1365 1275 7.08      
17 A' 1327 1240 154.18      
18 A' 1229 1148 6.76      
19 A' 1211 1132 0.49      
20 A' 1131 1057 4.78      
21 A' 1120 1047 0.84      
22 A' 1065 996 3.43      
23 A' 1022 955 0.65      
24 A' 987 922 20.28      
25 A' 752 703 1.83      
26 A' 630 589 0.66      
27 A' 598 559 25.42      
28 A' 468 438 0.48      
29 A' 371 346 0.73      
30 A' 224 209 4.63      
31 A" 3226 3015 6.43      
32 A" 1537 1436 9.06      
33 A" 1062 992 1.02      
34 A" 939 877 0.43      
35 A" 926 865 0.27      
36 A" 900 841 0.72      
37 A" 855 799 0.28      
38 A" 746 698 60.21      
39 A" 603 563 12.11      
40 A" 512 478 0.75      
41 A" 411 384 0.05      
42 A" 394 368 0.00      
43 A" 182 170 0.22      
44 A" 152 142 0.01      
45 A" 53 50 3.02      

Unscaled Zero Point Vibrational Energy (zpe) 30697.6 cm-1
Scaled (by 0.9344) Zero Point Vibrational Energy (zpe) 28683.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G**
ABC
0.12294 0.04068 0.03074

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G**

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.655 2.199 0.000
C2 0.232 1.688 0.000
C3 0.000 0.212 0.000
O4 -0.717 2.472 0.000
C5 1.050 -0.714 0.000
C6 -1.323 -0.247 0.000
C7 0.777 -2.081 0.000
C8 -1.592 -1.611 0.000
C9 -0.542 -2.532 0.000
H10 1.631 3.284 0.000
H11 2.190 1.846 0.879
H12 2.190 1.846 -0.879
H13 2.078 -0.379 0.000
H14 -2.119 0.485 0.000
H15 1.592 -2.792 0.000
H16 -2.616 -1.959 0.000
H17 -0.752 -3.593 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 O4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17
C11.51102.58592.38672.97543.85354.36905.00545.21591.08481.08871.08872.61244.14504.99175.96036.2719
C21.51101.49481.23062.53812.48293.80843.76984.29052.12132.15202.15202.77122.64194.68224.62765.3721
C32.58591.49482.37101.40031.40052.42072.41982.79653.47782.87052.87052.16032.13683.39963.39923.8781
O42.38671.23062.37103.64332.78544.79144.17535.00652.48363.10083.10083.99202.43255.74794.82066.0649
C52.97542.53811.40033.64332.41931.39372.79022.41654.03982.93692.93691.08103.38882.14733.87193.3961
C63.85352.48291.40052.78542.41932.78851.39022.41484.60314.18284.18283.40391.08083.87012.14533.3948
C74.36903.80842.42074.79141.39372.78852.41521.39425.43204.26474.26472.14243.86911.08163.39532.1500
C85.00543.76982.41984.17532.79021.39022.41521.39645.85995.19845.19843.87122.16053.39591.08172.1531
C95.21594.29052.79655.00652.41652.41481.39421.39646.20805.23455.23453.39133.40362.14992.15151.0817
H101.08482.12133.47782.48364.03984.60315.43205.85996.20801.77601.77603.68964.67946.07596.74667.2774
H111.08872.15202.87053.10082.93694.18284.26475.19845.23451.77601.75892.39454.60414.75826.19256.2453
H121.08872.15202.87053.10082.93694.18284.26475.19845.23451.77601.75892.39454.60414.75826.19256.2453
H132.61242.77122.16033.99201.08103.40392.14243.87123.39133.68962.39452.39454.28522.46204.95294.2821
H144.14502.64192.13682.43253.38881.08083.86912.16053.40364.67944.60414.60414.28524.95072.49334.3008
H154.99174.68223.39965.74792.14733.87011.08163.39592.14996.07594.75824.75822.46204.95074.28962.4762
H165.96034.62763.39924.82063.87192.14533.39531.08172.15156.74666.19256.19254.95292.49334.28962.4794
H176.27195.37213.87816.06493.39613.39482.15002.15311.08177.27746.24536.24534.28214.30082.47622.4794

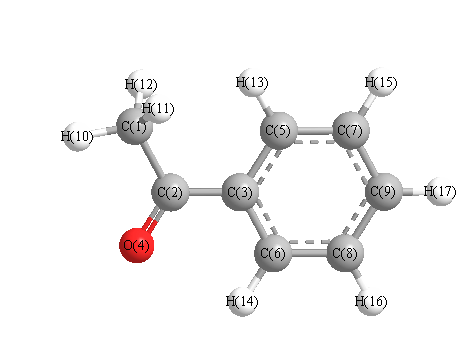
picture of acetophenone state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 118.699 C1 C2 O4 120.700
C2 C1 H10 108.496 C2 C1 H11 110.690
C2 C1 H12 110.690 C2 C3 C5 122.456
C2 C3 C6 118.048 C3 C2 O4 120.601
C3 C5 C7 120.086 C3 C5 H13 120.520
C3 C6 C8 120.246 C3 C6 H14 118.323
C5 C3 C6 119.496 C5 C7 C9 120.178
C5 C7 H15 119.809 C6 C8 C9 120.124
C6 C8 H16 119.913 C7 C5 H13 119.394
C7 C9 C8 119.870 C7 C9 H17 120.013
C8 C6 H14 121.432 C8 C9 H17 120.117
C9 C7 H15 120.013 C9 C8 H16 119.963
H10 C1 H11 109.598 H10 C1 H12 109.598
H11 C1 H12 107.757
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability