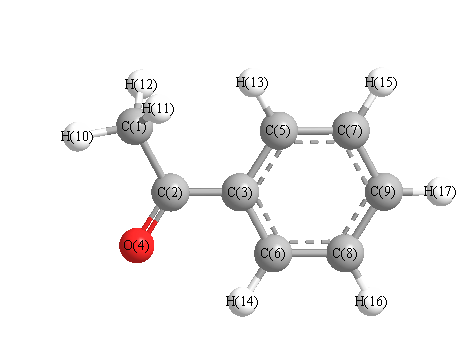
Vibrational Frequencies calculated at PBE1PBE/6-31G**
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3249 |
3096 |
6.48 |
|
|
|
| 2 |
A' |
3240 |
3088 |
12.36 |
|
|
|
| 3 |
A' |
3231 |
3080 |
19.56 |
|
|
|
| 4 |
A' |
3222 |
3071 |
5.33 |
|
|
|
| 5 |
A' |
3213 |
3062 |
0.10 |
|
|
|
| 6 |
A' |
3203 |
3053 |
8.57 |
|
|
|
| 7 |
A' |
3077 |
2933 |
1.90 |
|
|
|
| 8 |
A' |
1821 |
1735 |
168.66 |
|
|
|
| 9 |
A' |
1685 |
1606 |
20.21 |
|
|
|
| 10 |
A' |
1666 |
1588 |
7.43 |
|
|
|
| 11 |
A' |
1548 |
1475 |
1.70 |
|
|
|
| 12 |
A' |
1503 |
1433 |
20.56 |
|
|
|
| 13 |
A' |
1494 |
1424 |
8.93 |
|
|
|
| 14 |
A' |
1409 |
1343 |
72.00 |
|
|
|
| 15 |
A' |
1397 |
1331 |
4.00 |
|
|
|
| 16 |
A' |
1351 |
1288 |
3.53 |
|
|
|
| 17 |
A' |
1310 |
1248 |
174.07 |
|
|
|
| 18 |
A' |
1213 |
1156 |
13.07 |
|
|
|
| 19 |
A' |
1195 |
1139 |
0.56 |
|
|
|
| 20 |
A' |
1123 |
1070 |
7.10 |
|
|
|
| 21 |
A' |
1109 |
1057 |
1.33 |
|
|
|
| 22 |
A' |
1063 |
1014 |
5.16 |
|
|
|
| 23 |
A' |
1023 |
975 |
0.74 |
|
|
|
| 24 |
A' |
975 |
930 |
21.57 |
|
|
|
| 25 |
A' |
751 |
716 |
0.87 |
|
|
|
| 26 |
A' |
629 |
600 |
0.99 |
|
|
|
| 27 |
A' |
598 |
570 |
26.31 |
|
|
|
| 28 |
A' |
468 |
446 |
0.78 |
|
|
|
| 29 |
A' |
367 |
350 |
0.84 |
|
|
|
| 30 |
A' |
216 |
206 |
4.91 |
|
|
|
| 31 |
A" |
3149 |
3001 |
6.08 |
|
|
|
| 32 |
A" |
1503 |
1433 |
11.90 |
|
|
|
| 33 |
A" |
1056 |
1007 |
1.71 |
|
|
|
| 34 |
A" |
1017 |
969 |
0.43 |
|
|
|
| 35 |
A" |
994 |
947 |
0.38 |
|
|
|
| 36 |
A" |
951 |
906 |
2.51 |
|
|
|
| 37 |
A" |
872 |
831 |
0.19 |
|
|
|
| 38 |
A" |
780 |
744 |
35.53 |
|
|
|
| 39 |
A" |
710 |
676 |
31.15 |
|
|
|
| 40 |
A" |
604 |
576 |
9.77 |
|
|
|
| 41 |
A" |
430 |
410 |
0.06 |
|
|
|
| 42 |
A" |
415 |
395 |
0.01 |
|
|
|
| 43 |
A" |
171 |
163 |
0.35 |
|
|
|
| 44 |
A" |
154 |
147 |
0.02 |
|
|
|
| 45 |
A" |
65 |
62 |
3.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30609.4 cm
-1
Scaled (by 0.9531) Zero Point Vibrational Energy (zpe) 29173.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-31G**
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.629 |
|
|
|
| 2 |
C |
0.428 |
|
|
|
| 3 |
C |
0.018 |
|
|
|
| 4 |
O |
-0.452 |
|
|
|
| 5 |
C |
-0.202 |
|
|
|
| 6 |
C |
-0.179 |
|
|
|
| 7 |
C |
-0.175 |
|
|
|
| 8 |
C |
-0.173 |
|
|
|
| 9 |
C |
-0.159 |
|
|
|
| 10 |
H |
0.213 |
|
|
|
| 11 |
H |
0.200 |
|
|
|
| 12 |
H |
0.200 |
|
|
|
| 13 |
H |
0.174 |
|
|
|
| 14 |
H |
0.205 |
|
|
|
| 15 |
H |
0.176 |
|
|
|
| 16 |
H |
0.179 |
|
|
|
| 17 |
H |
0.178 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.853 |
-2.354 |
0.000 |
2.995 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.543 |
5.399 |
0.000 |
| y |
5.399 |
-52.515 |
0.000 |
| z |
0.000 |
0.000 |
-54.329 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.879 |
5.399 |
0.000 |
| y |
5.399 |
-2.079 |
0.000 |
| z |
0.000 |
0.000 |
-4.800 |
|
| Polar |
|---|
| 3z2-r2 | -9.600 |
|---|
| x2-y2 | 5.972 |
|---|
| xy | 5.399 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.571 |
0.601 |
0.000 |
| y |
0.601 |
16.632 |
0.000 |
| z |
0.000 |
0.000 |
5.299 |
<r2> (average value of r
2) Å
2
| <r2> |
343.909 |
| (<r2>)1/2 |
18.545 |