return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CHO (benzaldehyde)

using model chemistry: ROMP2/TZVP

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at ROMP2/TZVP
 hartrees
Energy at 0K-344.680389
Energy at 298.15K 
HF Energy-343.536883
Nuclear repulsion energy321.158314
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROMP2/TZVP
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3248 3248        
2 A' 3241 3241        
3 A' 3232 3232        
4 A' 3222 3222        
5 A' 3210 3210        
6 A' 2965 2965        
7 A' 1743 1743        
8 A' 1651 1651        
9 A' 1635 1635        
10 A' 1530 1530        
11 A' 1494 1494        
12 A' 1457 1457        
13 A' 1433 1433        
14 A' 1344 1344        
15 A' 1239 1239        
16 A' 1202 1202        
17 A' 1195 1195        
18 A' 1109 1109        
19 A' 1048 1048        
20 A' 1004 1004        
21 A' 839 839        
22 A' 661 661        
23 A' 624 624        
24 A' 439 439        
25 A' 220 220        
26 A" 968 968        
27 A" 776 776        
28 A" 725 725        
29 A" 685 685        
30 A" 621 621        
31 A" 610 610        
32 A" 148 148        
33 A" 124i 124i        
34 A" 336i 336i        
35 A" 416i 416i        
36 A" 1811i 1811i        

Unscaled Zero Point Vibrational Energy (zpe) 21414.9 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 21414.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at ROMP2/TZVP
ABC
0.17535 0.05218 0.04021

See section I.F.4 to change rotational constant units
Geometric Data calculated at ROMP2/TZVP

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.569 0.000
C2 -1.043 -0.362 0.000
C3 -0.750 -1.720 0.000
C4 0.578 -2.152 0.000
C5 1.617 -1.224 0.000
C6 1.328 0.138 0.000
C7 -0.286 2.021 0.000
O8 -1.403 2.508 0.000
H9 0.610 2.670 0.000
H10 -2.066 -0.005 0.000
H11 -1.553 -2.447 0.000
H12 0.801 -3.212 0.000
H13 2.646 -1.561 0.000
H14 2.130 0.869 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 O8 H9 H10 H11 H12 H13 H14
C11.39832.40812.78172.41391.39641.48042.39362.18812.14453.39173.86483.39672.1510
C21.39831.38892.41532.79632.42402.50062.89233.45381.08322.14593.39493.87923.4039
C32.40811.38891.39662.41822.78753.76934.27754.59552.16131.08312.15253.39933.8725
C42.78172.41531.39661.39332.41014.26195.06374.82243.40602.15091.08322.15043.3969
C52.41392.79632.41821.39331.39203.76184.80054.02173.87933.39732.14931.08292.1550
C61.39642.42402.78752.41011.39202.48063.61632.63183.39763.87063.39152.15041.0850
C71.48042.50063.76934.26193.76182.48061.21831.10692.69704.64385.34514.62962.6768
O82.39362.89234.27755.06374.80053.61631.21832.02022.59904.95666.13005.74053.8946
H92.18813.45384.59554.82244.02172.63181.10692.02023.78435.55515.88534.69562.3562
H102.14451.08322.16133.40603.87933.39762.69702.59903.78432.49484.30174.96224.2863
H113.39172.14591.08312.15093.39733.87064.64384.95665.55512.49482.47514.29074.9556
H123.86483.39492.15251.08322.14933.39155.34516.13005.88534.30172.47512.47534.2924
H133.39673.87923.39932.15041.08292.15044.62965.74054.69564.96224.29072.47532.4851
H142.15103.40393.87253.39692.15501.08502.67683.89462.35624.28634.95564.29242.4851

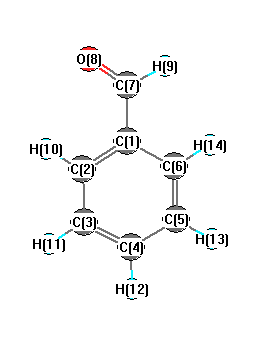
picture of benzaldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.533 C1 C2 H10 119.035
C1 C6 C5 119.924 C1 C6 H14 119.662
C1 C7 O8 124.696 C1 C7 H9 114.735
C2 C1 C6 120.303 C2 C1 C7 120.577
C2 C3 C4 120.248 C2 C3 H11 119.968
C3 C2 H10 121.433 C3 C4 C5 120.163
C3 C4 H12 119.932 C4 C3 H11 119.784
C4 C5 C6 119.829 C4 C5 H13 120.030
C5 C4 H12 119.905 C5 C6 H14 120.414
C6 C1 C7 119.119 C6 C5 H13 120.141
O8 C7 H9 120.569
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROMP2/TZVP Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.204      
2 C -0.190      
3 C -0.068      
4 C -0.109      
5 C -0.095      
6 C -0.190      
7 C 0.121      
8 O -0.392      
9 H 0.061      
10 H 0.155      
11 H 0.126      
12 H 0.126      
13 H 0.123      
14 H 0.130      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 262.361
(<r2>)1/2 16.198