Jump to
S1C2
Vibrational Frequencies calculated at MP3/TZVP
Geometric Data calculated at MP3/TZVP
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP3/TZVP
| | hartrees |
|---|
| Energy at 0K | -208.651138 |
| Energy at 298.15K | |
| HF Energy | -207.960135 |
| Nuclear repulsion energy | 120.136636 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/TZVP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3951 |
3695 |
82.08 |
|
|
|
| 2 |
A' |
3236 |
3026 |
12.85 |
|
|
|
| 3 |
A' |
3222 |
3014 |
1.50 |
|
|
|
| 4 |
A' |
3100 |
2899 |
13.37 |
|
|
|
| 5 |
A' |
1791 |
1675 |
5.06 |
|
|
|
| 6 |
A' |
1518 |
1419 |
14.71 |
|
|
|
| 7 |
A' |
1447 |
1353 |
13.03 |
|
|
|
| 8 |
A' |
1420 |
1328 |
36.64 |
|
|
|
| 9 |
A' |
1388 |
1298 |
42.54 |
|
|
|
| 10 |
A' |
1175 |
1099 |
8.27 |
|
|
|
| 11 |
A' |
980 |
916 |
109.50 |
|
|
|
| 12 |
A' |
935 |
875 |
0.03 |
|
|
|
| 13 |
A' |
690 |
646 |
11.70 |
|
|
|
| 14 |
A' |
328 |
306 |
1.64 |
|
|
|
| 15 |
A" |
3160 |
2956 |
14.19 |
|
|
|
| 16 |
A" |
1509 |
1411 |
10.01 |
|
|
|
| 17 |
A" |
1086 |
1015 |
2.19 |
|
|
|
| 18 |
A" |
876 |
820 |
11.23 |
|
|
|
| 19 |
A" |
479 |
448 |
28.20 |
|
|
|
| 20 |
A" |
380 |
355 |
111.76 |
|
|
|
| 21 |
A" |
69i |
64i |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16301.2 cm
-1
Scaled (by 0.9352) Zero Point Vibrational Energy (zpe) 15244.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/TZVP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.447 |
0.476 |
0.000 |
| C2 |
0.000 |
0.859 |
0.000 |
| N3 |
1.016 |
0.094 |
0.000 |
| O4 |
0.657 |
-1.260 |
0.000 |
| H5 |
1.511 |
-1.692 |
0.000 |
| H6 |
-1.577 |
-0.601 |
0.000 |
| H7 |
-1.937 |
0.896 |
0.880 |
| H8 |
-1.937 |
0.896 |
-0.880 |
| H9 |
0.258 |
1.911 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4973 | 2.4925 | 2.7278 | 3.6678 | 1.0847 | 1.0908 | 1.0908 | 2.2293 |
C2 | 1.4973 | | 1.2716 | 2.2187 | 2.9654 | 2.1491 | 2.1277 | 2.1277 | 1.0832 | N3 | 2.4925 | 1.2716 | | 1.4008 | 1.8539 | 2.6840 | 3.1835 | 3.1835 | 1.9686 | O4 | 2.7278 | 2.2187 | 1.4008 | | 0.9573 | 2.3287 | 3.4856 | 3.4856 | 3.1961 | H5 | 3.6678 | 2.9654 | 1.8539 | 0.9573 | | 3.2748 | 4.4001 | 4.4001 | 3.8151 | H6 | 1.0847 | 2.1491 | 2.6840 | 2.3287 | 3.2748 | | 1.7732 | 1.7732 | 3.1111 | H7 | 1.0908 | 2.1277 | 3.1835 | 3.4856 | 4.4001 | 1.7732 | | 1.7592 | 2.5738 | H8 | 1.0908 | 2.1277 | 3.1835 | 3.4856 | 4.4001 | 1.7732 | 1.7592 | | 2.5738 | H9 | 2.2293 | 1.0832 | 1.9686 | 3.1961 | 3.8151 | 3.1111 | 2.5738 | 2.5738 | |
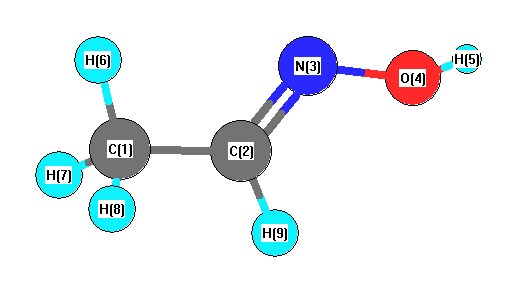 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.171 |
|
C1 |
C2 |
H9 |
118.637 |
| C2 |
C1 |
H6 |
111.685 |
|
C2 |
C1 |
H7 |
109.589 |
| C2 |
C1 |
H8 |
109.589 |
|
C2 |
N3 |
O4 |
112.149 |
| N3 |
C2 |
H9 |
113.192 |
|
N3 |
O4 |
H5 |
102.015 |
| H6 |
C1 |
H7 |
109.195 |
|
H6 |
C1 |
H8 |
109.195 |
| H7 |
C1 |
H8 |
107.491 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
