return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(C2H5)2 (diethylamine)

using model chemistry: MP3/TZVP

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP3/TZVP
 hartrees
Energy at 0K-213.236749
Energy at 298.15K-213.249497
HF Energy-212.387536
Nuclear repulsion energy188.791351
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/TZVP
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3574 3342 0.18      
2 A' 3180 2974 46.54      
3 A' 3161 2956 72.29      
4 A' 3111 2910 45.43      
5 A' 3087 2887 14.14      
6 A' 3005 2810 157.38      
7 A' 1562 1461 0.92      
8 A' 1532 1432 3.30      
9 A' 1518 1420 11.27      
10 A' 1473 1378 2.23      
11 A' 1433 1340 0.89      
12 A' 1351 1263 6.19      
13 A' 1265 1183 1.22      
14 A' 1200 1122 14.10      
15 A' 1090 1020 6.09      
16 A' 927 867 8.75      
17 A' 849 794 7.62      
18 A' 808 755 68.16      
19 A' 434 405 0.31      
20 A' 258 241 1.18      
21 A' 189 176 0.62      
22 A' 112 104 1.48      
23 A" 3180 2974 10.22      
24 A" 3160 2956 17.93      
25 A" 3109 2907 0.86      
26 A" 3087 2887 30.53      
27 A" 3002 2807 8.59      
28 A" 1550 1450 12.10      
29 A" 1538 1438 13.11      
30 A" 1528 1429 10.50      
31 A" 1512 1414 12.36      
32 A" 1448 1354 10.49      
33 A" 1393 1303 20.05      
34 A" 1310 1225 2.62      
35 A" 1197 1119 41.53      
36 A" 1142 1068 8.11      
37 A" 1090 1019 0.93      
38 A" 965 902 0.66      
39 A" 826 773 0.42      
40 A" 433 405 0.46      
41 A" 257 240 0.54      
42 A" 97 91 1.12      

Unscaled Zero Point Vibrational Energy (zpe) 33469.5 cm-1
Scaled (by 0.9352) Zero Point Vibrational Energy (zpe) 31300.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/TZVP
ABC
0.59666 0.07024 0.06625

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/TZVP

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.017 -0.288 0.000
C2 0.017 0.516 1.215
C3 0.017 0.516 -1.215
C4 0.017 -0.368 2.452
C5 0.017 -0.368 -2.452
H6 -0.801 -0.885 0.000
H7 -0.836 1.210 1.253
H8 0.922 1.128 1.203
H9 -0.836 1.210 -1.253
H10 0.922 1.128 -1.203
H11 0.056 0.236 3.359
H12 -0.888 -0.977 2.494
H13 0.877 -1.037 2.437
H14 0.056 0.236 -3.359
H15 -0.888 -0.977 -2.494
H16 0.877 -1.037 -2.437

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
N11.45701.45702.45382.45381.01272.13182.06672.13182.06673.39992.74162.69023.39992.74162.6902
C21.45702.42941.52133.77232.02681.10081.09232.70202.65272.16282.16492.15504.58244.09964.0600
C31.45702.42943.77231.52132.02682.70202.65271.10081.09234.58244.09964.06002.16282.16492.1550
C42.45381.52133.77234.90492.63652.15832.14934.11734.05191.09021.09191.08925.84295.06585.0090
C52.45383.77231.52134.90492.63654.11734.05192.15832.14935.84295.06585.00901.09021.09191.0892
H61.01272.02682.02682.63652.63652.44132.90992.44132.90993.64332.49762.96273.64332.49762.9627
H72.13181.10082.70202.15834.11732.44131.76112.50643.02162.48592.51513.06354.79764.33934.6476
H82.06671.09232.65272.14934.05192.90991.76113.02162.40552.48903.06212.49224.72814.62354.2348
H92.13182.70201.10084.11732.15832.44132.50643.02161.76114.79764.33934.64762.48592.51513.0635
H102.06672.65271.09234.05192.14932.90993.02162.40551.76114.72814.62354.23482.48903.06212.4922
H113.39992.16284.58241.09025.84293.64332.48592.48904.79764.72811.76361.77336.71806.05185.9903
H122.74162.16494.09961.09195.06582.49762.51513.06214.33934.62351.76361.76756.05184.98885.2380
H132.69022.15504.06001.08925.00902.96273.06352.49224.64764.23481.77331.76755.99035.23804.8735
H143.39994.58242.16285.84291.09023.64334.79764.72812.48592.48906.71806.05185.99031.76361.7733
H152.74164.09962.16495.06581.09192.49764.33934.62352.51513.06216.05184.98885.23801.76361.7675
H162.69024.06002.15505.00901.08922.96274.64764.23483.06352.49225.99035.23804.87351.77331.7675

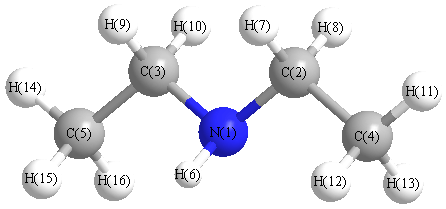
picture of diethylamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 110.936 N1 C2 H7 112.162
N1 C2 H8 107.469 N1 C3 C5 110.936
N1 C3 H9 112.162 N1 C3 H10 107.469
C2 N1 C3 112.967 C2 N1 H6 108.976
C2 C4 H11 110.750 C2 C4 H12 110.808
C2 C4 H13 110.182 C3 N1 H6 108.976
C3 C5 H14 110.750 C3 C5 H15 110.808
C3 C5 H16 110.182 C4 C2 H7 109.759
C4 C2 H8 109.549 C5 C3 H8 151.112
C5 C3 H10 109.549 H7 C2 H8 106.832
H9 C3 H10 106.832 H11 C4 H12 107.845
H11 C4 H13 108.912 H12 C4 H13 108.265
H14 C5 H15 107.845 H14 C5 H16 108.912
H15 C5 H16 108.265
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability